Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -637.920583 |
| Energy at 298.15K | -637.925522 |
| HF Energy | -637.088848 |
| Nuclear repulsion energy | 157.552255 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3128 |
2993 |
13.75 |
|
|
|
| 2 |
A' |
3098 |
2965 |
13.83 |
|
|
|
| 3 |
A' |
1548 |
1482 |
2.65 |
|
|
|
| 4 |
A' |
1526 |
1461 |
5.19 |
|
|
|
| 5 |
A' |
1449 |
1386 |
4.82 |
|
|
|
| 6 |
A' |
1310 |
1253 |
7.46 |
|
|
|
| 7 |
A' |
1115 |
1067 |
82.48 |
|
|
|
| 8 |
A' |
1092 |
1045 |
47.45 |
|
|
|
| 9 |
A' |
812 |
777 |
38.60 |
|
|
|
| 10 |
A' |
393 |
376 |
2.74 |
|
|
|
| 11 |
A' |
247 |
236 |
10.90 |
|
|
|
| 12 |
A" |
3194 |
3057 |
5.48 |
|
|
|
| 13 |
A" |
3153 |
3017 |
9.85 |
|
|
|
| 14 |
A" |
1327 |
1270 |
0.03 |
|
|
|
| 15 |
A" |
1244 |
1190 |
1.65 |
|
|
|
| 16 |
A" |
1084 |
1038 |
1.41 |
|
|
|
| 17 |
A" |
806 |
771 |
0.46 |
|
|
|
| 18 |
A" |
131 |
125 |
9.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13327.8 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 12754.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.622 |
0.000 |
| C2 |
0.981 |
-0.530 |
0.000 |
| Cl3 |
-1.659 |
-0.038 |
0.000 |
| F4 |
2.262 |
-0.010 |
0.000 |
| H5 |
0.116 |
1.235 |
0.889 |
| H6 |
0.116 |
1.235 |
-0.889 |
| H7 |
0.859 |
-1.146 |
0.890 |
| H8 |
0.859 |
-1.146 |
-0.890 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5136 | 1.7849 | 2.3483 | 1.0858 | 1.0858 | 2.1578 | 2.1578 |
C2 | 1.5136 | | 2.6854 | 1.3822 | 2.1574 | 2.1574 | 1.0892 | 1.0892 | Cl3 | 1.7849 | 2.6854 | | 3.9204 | 2.3577 | 2.3577 | 2.8910 | 2.8910 | F4 | 2.3483 | 1.3822 | 3.9204 | | 2.6348 | 2.6348 | 2.0130 | 2.0130 | H5 | 1.0858 | 2.1574 | 2.3577 | 2.6348 | | 1.7773 | 2.4943 | 3.0635 | H6 | 1.0858 | 2.1574 | 2.3577 | 2.6348 | 1.7773 | | 3.0635 | 2.4943 | H7 | 2.1578 | 1.0892 | 2.8910 | 2.0130 | 2.4943 | 3.0635 | | 1.7799 | H8 | 2.1578 | 1.0892 | 2.8910 | 2.0130 | 3.0635 | 2.4943 | 1.7799 | |
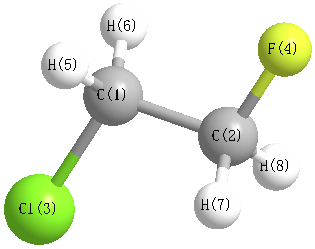 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.290 |
|
C1 |
C2 |
H7 |
110.951 |
| C1 |
C2 |
H8 |
110.951 |
|
C2 |
C1 |
Cl3 |
108.731 |
| C2 |
C1 |
H5 |
111.124 |
|
C2 |
C1 |
H6 |
111.124 |
| Cl3 |
C1 |
H5 |
107.944 |
|
Cl3 |
C1 |
H6 |
107.944 |
| F4 |
C2 |
H7 |
108.493 |
|
F4 |
C2 |
H8 |
108.493 |
| H5 |
C1 |
H6 |
109.859 |
|
H7 |
C2 |
H8 |
109.586 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -637.919657 |
| Energy at 298.15K | |
| HF Energy | -637.087636 |
| Nuclear repulsion energy | 161.514376 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.088 |
0.842 |
-0.297 |
| C2 |
1.196 |
0.410 |
0.365 |
| Cl3 |
-1.426 |
-0.284 |
0.065 |
| F4 |
1.661 |
-0.754 |
-0.205 |
| H5 |
-0.381 |
1.823 |
0.069 |
| H6 |
0.028 |
0.873 |
-1.377 |
| H7 |
1.954 |
1.182 |
0.219 |
| H8 |
1.050 |
0.234 |
1.430 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5075 | 1.7857 | 2.3699 | 1.0874 | 1.0865 | 2.1341 | 2.1554 |
C2 | 1.5075 | | 2.7294 | 1.3774 | 2.1373 | 2.1472 | 1.0921 | 1.0896 | Cl3 | 1.7857 | 2.7294 | | 3.1351 | 2.3524 | 2.3515 | 3.6885 | 2.8751 | F4 | 2.3699 | 1.3774 | 3.1351 | | 3.2995 | 2.5861 | 2.0042 | 2.0064 | H5 | 1.0874 | 2.1373 | 2.3524 | 3.2995 | | 1.7778 | 2.4258 | 2.5339 | H6 | 1.0865 | 2.1472 | 2.3515 | 2.5861 | 1.7778 | | 2.5209 | 3.0544 | H7 | 2.1341 | 1.0921 | 3.6885 | 2.0042 | 2.4258 | 2.5209 | | 1.7840 | H8 | 2.1554 | 1.0896 | 2.8751 | 2.0064 | 2.5339 | 3.0544 | 1.7840 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.382 |
|
C1 |
C2 |
H7 |
109.316 |
| C1 |
C2 |
H8 |
111.159 |
|
C2 |
C1 |
Cl3 |
111.667 |
| C2 |
C1 |
H5 |
109.847 |
|
C2 |
C1 |
H6 |
110.692 |
| Cl3 |
C1 |
H5 |
107.415 |
|
Cl3 |
C1 |
H6 |
107.398 |
| F4 |
C2 |
H7 |
107.944 |
|
F4 |
C2 |
H8 |
108.270 |
| H5 |
C1 |
H6 |
109.732 |
|
H7 |
C2 |
H8 |
109.706 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
