return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (Methyl glyoxal)

using model chemistry: CCD/aug-cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/aug-cc-pVTZ
 hartrees
Energy at 0K-266.707901
Energy at 298.15K-266.712072
HF Energy-265.742705
Nuclear repulsion energy160.390719
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3190 3053 3.86      
2 A' 3075 2942 0.17      
3 A' 3018 2888 51.64      
4 A' 1851 1771 6.95      
5 A' 1839 1759 189.93      
6 A' 1488 1424 14.30      
7 A' 1428 1367 27.64      
8 A' 1387 1327 3.00      
9 A' 1291 1235 17.71      
10 A' 1036 991 3.78      
11 A' 809 774 11.05      
12 A' 586 561 12.00      
13 A' 492 471 29.06      
14 A' 250 239 16.55      
15 A" 3145 3009 2.38      
16 A" 1494 1430 8.96      
17 A" 1101 1054 3.01      
18 A" 934 893 0.05      
19 A" 474 454 0.78      
20 A" 136 130 11.62      
21 A" 105 101 13.61      

Unscaled Zero Point Vibrational Energy (zpe) 14563.4 cm-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 13937.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/aug-cc-pVTZ
ABC
0.30546 0.14898 0.10201

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/aug-cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.841 -0.726 0.000
C2 0.000 0.551 0.000
C3 1.491 0.391 0.000
O4 -0.357 -1.825 0.000
O5 -0.584 1.606 0.000
H6 -1.928 -0.544 0.000
H7 1.964 1.369 0.000
H8 1.798 -0.182 0.875
H9 1.798 -0.182 -0.875

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.52882.58601.20142.34631.10133.50102.83342.8334
C21.52881.49952.40271.20662.21682.12762.12992.1299
C32.58601.49952.88592.40473.54421.08601.09051.0905
O41.20142.40272.88593.43922.02673.94852.84812.8481
O52.34631.20662.40473.43922.53572.55923.10493.1049
H61.10132.21683.54422.02672.53574.33633.84423.8442
H73.50102.12761.08603.94852.55924.33631.78861.7886
H82.83342.12991.09052.84813.10493.84421.78861.7506
H92.83342.12991.09052.84813.10493.84421.78861.7506

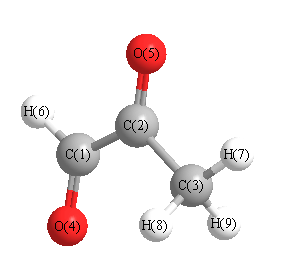
picture of Methyl glyoxal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 117.285 C1 C2 O5 117.656
C2 C1 O4 122.845 C2 C1 H6 113.898
C2 C3 H7 109.718 C2 C3 H8 109.631
C2 C3 H9 109.631 C3 C2 O5 125.059
O4 C1 H6 123.256 H7 C3 H8 110.522
H7 C3 H9 110.522 H8 C3 H9 106.769
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability