Jump to
S1C2
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1071.811721 |
| Energy at 298.15K | |
| HF Energy | -1070.834171 |
| Nuclear repulsion energy | 267.092284 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.098 |
-0.028 |
0.519 |
| C2 |
-0.723 |
-1.273 |
0.188 |
| H3 |
0.223 |
0.050 |
1.595 |
| Cl4 |
1.724 |
-0.273 |
-0.167 |
| Cl5 |
-0.654 |
1.450 |
-0.058 |
| O6 |
-1.807 |
-1.255 |
-0.311 |
| H7 |
-0.210 |
-2.208 |
0.467 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5270 | 1.0864 | 1.7812 | 1.7556 | 2.4127 | 2.2016 |
C2 | 1.5270 | | 2.1499 | 2.6666 | 2.7350 | 1.1940 | 1.1014 | H3 | 1.0864 | 2.1499 | | 2.3367 | 2.3365 | 3.0749 | 2.5605 | Cl4 | 1.7812 | 2.6666 | 2.3367 | | 2.9382 | 3.6673 | 2.8078 | Cl5 | 1.7556 | 2.7350 | 2.3365 | 2.9382 | | 2.9512 | 3.7214 | O6 | 2.4127 | 1.1940 | 3.0749 | 3.6673 | 2.9512 | | 2.0155 | H7 | 2.2016 | 1.1014 | 2.5605 | 2.8078 | 3.7214 | 2.0155 | |
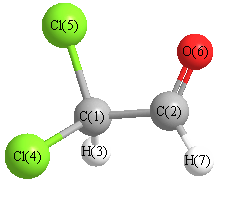 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.461 |
|
C1 |
C2 |
H7 |
112.773 |
| C2 |
C1 |
H3 |
109.543 |
|
C2 |
C1 |
Cl4 |
107.175 |
| C2 |
C1 |
Cl5 |
112.670 |
|
H3 |
C1 |
Cl4 |
106.626 |
| H3 |
C1 |
Cl5 |
108.286 |
|
Cl4 |
C1 |
Cl5 |
112.351 |
| O6 |
C2 |
H7 |
122.765 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
