All results from a given calculation for C5H10O (2-methylbutyraldehyde)
using model chemistry: CCD/aug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C1 |
1A |
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -271.271135 |
| Energy at 298.15K | |
| HF Energy | -270.120002 |
| Nuclear repulsion energy | 240.616000 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.372 |
-0.691 |
0.221 |
| C2 |
-0.085 |
0.077 |
0.406 |
| C3 |
1.043 |
-0.689 |
-0.305 |
| C4 |
2.436 |
-0.147 |
0.008 |
| C5 |
-0.221 |
1.524 |
-0.049 |
| O6 |
-2.382 |
-0.248 |
-0.261 |
| H7 |
-1.334 |
-1.745 |
0.558 |
| H8 |
0.125 |
0.043 |
1.483 |
| H9 |
0.996 |
-1.742 |
-0.014 |
| H10 |
0.865 |
-0.654 |
-1.384 |
| H11 |
3.204 |
-0.755 |
-0.471 |
| H12 |
2.557 |
0.877 |
-0.346 |
| H13 |
2.622 |
-0.156 |
1.083 |
| H14 |
-0.383 |
1.571 |
-1.127 |
| H15 |
-1.074 |
2.002 |
0.432 |
| H16 |
0.674 |
2.095 |
0.193 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5109 | 2.4718 | 3.8525 | 2.5111 | 1.2036 | 1.1070 | 2.0915 | 2.6017 | 2.7530 | 4.6281 | 4.2679 | 4.1214 | 2.8127 | 2.7176 | 3.4571 |
C2 | 1.5109 | | 1.5373 | 2.5613 | 1.5227 | 2.4142 | 2.2146 | 1.0982 | 2.1575 | 2.1536 | 3.5032 | 2.8601 | 2.8000 | 2.1606 | 2.1640 | 2.1660 | C3 | 2.4718 | 1.5373 | | 1.5265 | 2.5617 | 3.4537 | 2.7405 | 2.1388 | 1.0939 | 1.0940 | 2.1677 | 2.1779 | 2.1690 | 2.7961 | 3.5020 | 2.8516 | C4 | 3.8525 | 2.5613 | 1.5265 | | 3.1395 | 4.8262 | 4.1312 | 2.7481 | 2.1489 | 2.1588 | 1.0903 | 1.0898 | 1.0919 | 3.4907 | 4.1370 | 2.8568 | C5 | 2.5111 | 1.5227 | 2.5617 | 3.1395 | | 2.8024 | 3.5061 | 2.1588 | 3.4863 | 2.7765 | 4.1357 | 2.8680 | 3.4914 | 1.0910 | 1.0891 | 1.0892 | O6 | 1.2036 | 2.4142 | 3.4537 | 4.8262 | 2.8024 | | 2.0027 | 3.0679 | 3.7023 | 3.4593 | 5.6124 | 5.0657 | 5.1825 | 2.8376 | 2.6928 | 3.8776 | H7 | 1.1070 | 2.2146 | 2.7405 | 4.1312 | 3.5061 | 2.0027 | | 2.4859 | 2.3993 | 3.1295 | 4.7568 | 4.7777 | 4.2955 | 3.8390 | 3.7576 | 4.3486 | H8 | 2.0915 | 1.0982 | 2.1388 | 2.7481 | 2.1588 | 3.0679 | 2.4859 | | 2.4872 | 3.0415 | 3.7326 | 3.1550 | 2.5367 | 3.0666 | 2.5256 | 2.4854 | H9 | 2.6017 | 2.1575 | 1.0939 | 2.1489 | 3.4863 | 3.7023 | 2.3993 | 2.4872 | | 1.7540 | 2.4607 | 3.0667 | 2.5228 | 3.7577 | 4.3014 | 3.8561 | H10 | 2.7530 | 2.1536 | 1.0940 | 2.1588 | 2.7765 | 3.4593 | 3.1295 | 3.0415 | 1.7540 | | 2.5123 | 2.5065 | 3.0697 | 2.5647 | 3.7565 | 3.1748 | H11 | 4.6281 | 3.5032 | 2.1677 | 1.0903 | 4.1357 | 5.6124 | 4.7568 | 3.7326 | 2.4607 | 2.5123 | | 1.7599 | 1.7647 | 4.3252 | 5.1685 | 3.8677 | H12 | 4.2679 | 2.8601 | 2.1779 | 1.0898 | 2.8680 | 5.0657 | 4.7777 | 3.1550 | 3.0667 | 2.5065 | 1.7599 | | 1.7647 | 3.1200 | 3.8799 | 2.3058 | H13 | 4.1214 | 2.8000 | 2.1690 | 1.0919 | 3.4914 | 5.1825 | 4.2955 | 2.5367 | 2.5228 | 3.0697 | 1.7647 | 1.7647 | | 4.1109 | 4.3290 | 3.1068 | H14 | 2.8127 | 2.1606 | 2.7961 | 3.4907 | 1.0910 | 2.8376 | 3.8390 | 3.0666 | 3.7577 | 2.5647 | 4.3252 | 3.1200 | 4.1109 | | 1.7584 | 1.7701 | H15 | 2.7176 | 2.1640 | 3.5020 | 4.1370 | 1.0891 | 2.6928 | 3.7576 | 2.5256 | 4.3014 | 3.7565 | 5.1685 | 3.8799 | 4.3290 | 1.7584 | | 1.7673 | H16 | 3.4571 | 2.1660 | 2.8516 | 2.8568 | 1.0892 | 3.8776 | 4.3486 | 2.4854 | 3.8561 | 3.1748 | 3.8677 | 2.3058 | 3.1068 | 1.7701 | 1.7673 | |
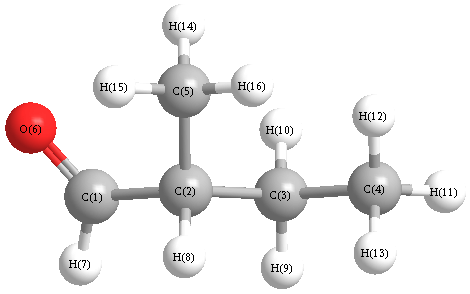 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
108.367 |
|
C1 |
C2 |
C5 |
111.732 |
| C1 |
C2 |
H8 |
105.471 |
|
C2 |
C1 |
O6 |
125.210 |
| C2 |
C1 |
H7 |
114.670 |
|
C2 |
C3 |
C4 |
113.437 |
| C2 |
C3 |
H9 |
109.001 |
|
C2 |
C3 |
H10 |
108.690 |
| C2 |
C5 |
H14 |
110.421 |
|
C2 |
C5 |
H15 |
110.801 |
| C2 |
C5 |
H16 |
110.962 |
|
C3 |
C2 |
C5 |
113.683 |
| C3 |
C2 |
H8 |
107.321 |
|
C3 |
C4 |
H11 |
110.762 |
| C3 |
C4 |
H12 |
111.606 |
|
C3 |
C4 |
H13 |
110.762 |
| C4 |
C3 |
H9 |
109.062 |
|
C4 |
C3 |
H10 |
109.834 |
| C5 |
C2 |
H8 |
109.852 |
|
O6 |
C1 |
H7 |
120.116 |
| H9 |
C3 |
H10 |
106.588 |
|
H11 |
C4 |
H12 |
107.660 |
| H11 |
C4 |
H13 |
107.931 |
|
H12 |
C4 |
H13 |
107.970 |
| H14 |
C5 |
H15 |
107.529 |
|
H14 |
C5 |
H16 |
108.568 |
| H15 |
C5 |
H16 |
108.453 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
