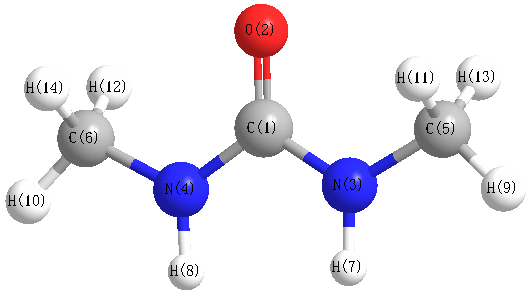
Vibrational Frequencies calculated at B3PW91/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3635 |
3508 |
22.92 |
|
|
|
| 2 |
A |
3149 |
3039 |
6.04 |
|
|
|
| 3 |
A |
3097 |
2988 |
7.33 |
|
|
|
| 4 |
A |
3019 |
2913 |
3.47 |
|
|
|
| 5 |
A |
1766 |
1705 |
280.02 |
|
|
|
| 6 |
A |
1527 |
1474 |
24.26 |
|
|
|
| 7 |
A |
1489 |
1437 |
0.19 |
|
|
|
| 8 |
A |
1476 |
1424 |
2.93 |
|
|
|
| 9 |
A |
1437 |
1386 |
11.42 |
|
|
|
| 10 |
A |
1201 |
1160 |
0.54 |
|
|
|
| 11 |
A |
1181 |
1140 |
3.16 |
|
|
|
| 12 |
A |
1142 |
1102 |
5.35 |
|
|
|
| 13 |
A |
924 |
892 |
3.80 |
|
|
|
| 14 |
A |
514 |
496 |
24.30 |
|
|
|
| 15 |
A |
379 |
366 |
48.46 |
|
|
|
| 16 |
A |
223 |
215 |
4.17 |
|
|
|
| 17 |
A |
174 |
168 |
0.00 |
|
|
|
| 18 |
A |
70 |
68 |
0.13 |
|
|
|
| 19 |
B |
3630 |
3504 |
7.59 |
|
|
|
| 20 |
B |
3149 |
3039 |
2.09 |
|
|
|
| 21 |
B |
3097 |
2989 |
47.88 |
|
|
|
| 22 |
B |
3017 |
2912 |
127.70 |
|
|
|
| 23 |
B |
1569 |
1514 |
378.97 |
|
|
|
| 24 |
B |
1503 |
1450 |
6.21 |
|
|
|
| 25 |
B |
1480 |
1429 |
72.18 |
|
|
|
| 26 |
B |
1440 |
1390 |
7.23 |
|
|
|
| 27 |
B |
1263 |
1219 |
220.58 |
|
|
|
| 28 |
B |
1151 |
1111 |
23.17 |
|
|
|
| 29 |
B |
1137 |
1098 |
5.95 |
|
|
|
| 30 |
B |
1054 |
1018 |
0.91 |
|
|
|
| 31 |
B |
774 |
747 |
9.79 |
|
|
|
| 32 |
B |
720 |
695 |
18.19 |
|
|
|
| 33 |
B |
475 |
458 |
118.22 |
|
|
|
| 34 |
B |
322 |
311 |
42.38 |
|
|
|
| 35 |
B |
119 |
115 |
11.81 |
|
|
|
| 36 |
B |
73 |
70 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26187.2 cm
-1
Scaled (by 0.9651) Zero Point Vibrational Energy (zpe) 25273.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.433 |
|
|
|
| 2 |
O |
-0.692 |
|
|
|
| 3 |
N |
-0.035 |
|
|
|
| 4 |
N |
-0.035 |
|
|
|
| 5 |
C |
-0.645 |
|
|
|
| 6 |
C |
-0.645 |
|
|
|
| 7 |
H |
0.023 |
|
|
|
| 8 |
H |
0.023 |
|
|
|
| 9 |
H |
0.254 |
|
|
|
| 10 |
H |
0.254 |
|
|
|
| 11 |
H |
0.321 |
|
|
|
| 12 |
H |
0.321 |
|
|
|
| 13 |
H |
0.212 |
|
|
|
| 14 |
H |
0.212 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.773 |
3.773 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.822 |
-1.462 |
0.000 |
| y |
-1.462 |
-30.474 |
0.000 |
| z |
0.000 |
0.000 |
-37.358 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.906 |
-1.462 |
0.000 |
| y |
-1.462 |
7.116 |
0.000 |
| z |
0.000 |
0.000 |
-3.210 |
|
| Polar |
|---|
| 3z2-r2 | -6.420 |
|---|
| x2-y2 | -7.348 |
|---|
| xy | -1.462 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.846 |
-0.434 |
0.000 |
| y |
-0.434 |
11.423 |
0.000 |
| z |
0.000 |
0.000 |
8.691 |
<r2> (average value of r
2) Å
2
| <r2> |
192.249 |
| (<r2>)1/2 |
13.865 |