Jump to
S1C2
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -341.925115 |
| Energy at 298.15K | |
| HF Energy | -340.658373 |
| Nuclear repulsion energy | 245.073583 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.867 |
| O2 |
0.000 |
0.000 |
2.053 |
| O3 |
0.000 |
1.136 |
0.058 |
| O4 |
0.000 |
-1.136 |
0.058 |
| C5 |
0.000 |
0.775 |
-1.289 |
| C6 |
0.000 |
-0.775 |
-1.289 |
| H7 |
-0.889 |
1.184 |
-1.772 |
| H8 |
0.889 |
1.184 |
-1.772 |
| H9 |
0.889 |
-1.184 |
-1.772 |
| H10 |
-0.889 |
-1.184 |
-1.772 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
O4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.1862 | 1.3945 | 1.3945 | 2.2914 | 2.2914 | 3.0261 | 3.0261 | 3.0261 | 3.0261 |
O2 | 1.1862 | | 2.2960 | 2.2960 | 3.4312 | 3.4312 | 4.1018 | 4.1018 | 4.1018 | 4.1018 | O3 | 1.3945 | 2.2960 | | 2.2711 | 1.3945 | 2.3376 | 2.0348 | 2.0348 | 3.0854 | 3.0854 | O4 | 1.3945 | 2.2960 | 2.2711 | | 2.3376 | 1.3945 | 3.0854 | 3.0854 | 2.0348 | 2.0348 | C5 | 2.2914 | 3.4312 | 1.3945 | 2.3376 | | 1.5498 | 1.0914 | 1.0914 | 2.2050 | 2.2050 | C6 | 2.2914 | 3.4312 | 2.3376 | 1.3945 | 1.5498 | | 2.2050 | 2.2050 | 1.0914 | 1.0914 | H7 | 3.0261 | 4.1018 | 2.0348 | 3.0854 | 1.0914 | 2.2050 | | 1.7785 | 2.9620 | 2.3687 | H8 | 3.0261 | 4.1018 | 2.0348 | 3.0854 | 1.0914 | 2.2050 | 1.7785 | | 2.3687 | 2.9620 | H9 | 3.0261 | 4.1018 | 3.0854 | 2.0348 | 2.2050 | 1.0914 | 2.9620 | 2.3687 | | 1.7785 | H10 | 3.0261 | 4.1018 | 3.0854 | 2.0348 | 2.2050 | 1.0914 | 2.3687 | 2.9620 | 1.7785 | |
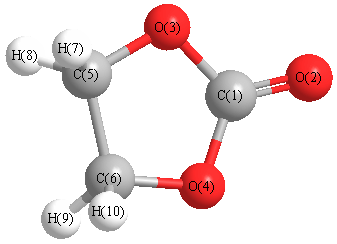 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O3 |
C5 |
110.489 |
|
C1 |
O4 |
C6 |
110.489 |
| O2 |
C1 |
O3 |
125.479 |
|
O2 |
C1 |
O4 |
125.479 |
| O3 |
C1 |
O4 |
109.042 |
|
O3 |
C5 |
C6 |
104.990 |
| O3 |
C5 |
H7 |
109.268 |
|
O3 |
C5 |
H8 |
109.268 |
| O4 |
C6 |
C5 |
104.990 |
|
O4 |
C6 |
H9 |
109.268 |
| O4 |
C6 |
H10 |
109.268 |
|
C5 |
C6 |
H9 |
112.034 |
| C5 |
C6 |
H10 |
112.034 |
|
C6 |
C5 |
H7 |
112.034 |
| C6 |
C5 |
H8 |
112.034 |
|
H7 |
C5 |
H8 |
109.127 |
| H9 |
C6 |
H10 |
109.127 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -341.922703 |
| Energy at 298.15K | |
| HF Energy | -340.652768 |
| Nuclear repulsion energy | 243.285107 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is C2
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
