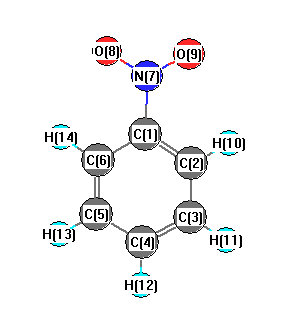
Vibrational Frequencies calculated at B1B95/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3248 |
3113 |
6.34 |
|
|
|
| 2 |
A1 |
3225 |
3092 |
6.31 |
|
|
|
| 3 |
A1 |
3203 |
3070 |
0.74 |
|
|
|
| 4 |
A1 |
1655 |
1586 |
2.76 |
|
|
|
| 5 |
A1 |
1514 |
1452 |
8.82 |
|
|
|
| 6 |
A1 |
1429 |
1370 |
271.01 |
|
|
|
| 7 |
A1 |
1191 |
1142 |
0.06 |
|
|
|
| 8 |
A1 |
1129 |
1082 |
30.58 |
|
|
|
| 9 |
A1 |
1044 |
1001 |
11.60 |
|
|
|
| 10 |
A1 |
1023 |
980 |
0.07 |
|
|
|
| 11 |
A1 |
883 |
846 |
34.23 |
|
|
|
| 12 |
A1 |
693 |
665 |
8.94 |
|
|
|
| 13 |
A1 |
392 |
376 |
1.69 |
|
|
|
| 14 |
A2 |
1009 |
967 |
0.00 |
|
|
|
| 15 |
A2 |
867 |
831 |
0.00 |
|
|
|
| 16 |
A2 |
414 |
397 |
0.00 |
|
|
|
| 17 |
A2 |
56 |
54 |
0.00 |
|
|
|
| 18 |
B1 |
1023 |
980 |
0.06 |
|
|
|
| 19 |
B1 |
974 |
934 |
2.77 |
|
|
|
| 20 |
B1 |
832 |
797 |
5.63 |
|
|
|
| 21 |
B1 |
732 |
702 |
82.06 |
|
|
|
| 22 |
B1 |
686 |
658 |
5.88 |
|
|
|
| 23 |
B1 |
445 |
426 |
0.58 |
|
|
|
| 24 |
B1 |
171 |
164 |
0.90 |
|
|
|
| 25 |
B2 |
3248 |
3113 |
0.50 |
|
|
|
| 26 |
B2 |
3217 |
3083 |
8.44 |
|
|
|
| 27 |
B2 |
1690 |
1620 |
89.34 |
|
|
|
| 28 |
B2 |
1641 |
1573 |
207.68 |
|
|
|
| 29 |
B2 |
1500 |
1438 |
0.44 |
|
|
|
| 30 |
B2 |
1379 |
1322 |
12.17 |
|
|
|
| 31 |
B2 |
1325 |
1270 |
0.19 |
|
|
|
| 32 |
B2 |
1180 |
1131 |
0.91 |
|
|
|
| 33 |
B2 |
1088 |
1042 |
8.07 |
|
|
|
| 34 |
B2 |
614 |
589 |
0.06 |
|
|
|
| 35 |
B2 |
529 |
507 |
1.28 |
|
|
|
| 36 |
B2 |
245 |
235 |
0.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22746.8 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 21802.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.559 |
|
|
|
| 2 |
C |
-0.746 |
|
|
|
| 3 |
C |
-0.459 |
|
|
|
| 4 |
C |
-0.516 |
|
|
|
| 5 |
C |
-0.459 |
|
|
|
| 6 |
C |
-0.746 |
|
|
|
| 7 |
N |
0.777 |
|
|
|
| 8 |
O |
-0.653 |
|
|
|
| 9 |
O |
-0.653 |
|
|
|
| 10 |
H |
0.722 |
|
|
|
| 11 |
H |
0.442 |
|
|
|
| 12 |
H |
0.571 |
|
|
|
| 13 |
H |
0.442 |
|
|
|
| 14 |
H |
0.722 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.583 |
4.583 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-52.514 |
0.000 |
0.000 |
| y |
0.000 |
-47.455 |
0.000 |
| z |
0.000 |
0.000 |
-54.708 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.433 |
0.000 |
0.000 |
| y |
0.000 |
6.156 |
0.000 |
| z |
0.000 |
0.000 |
-4.723 |
|
| Polar |
|---|
| 3z2-r2 | -9.446 |
|---|
| x2-y2 | -5.059 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.196 |
0.000 |
0.000 |
| y |
0.000 |
14.114 |
0.000 |
| z |
0.000 |
0.000 |
16.773 |
<r2> (average value of r
2) Å
2
| <r2> |
309.123 |
| (<r2>)1/2 |
17.582 |