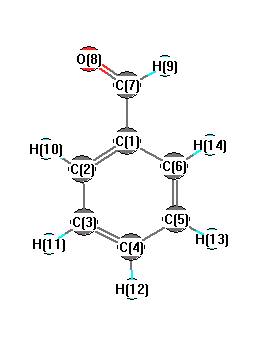
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3256 |
3076 |
6.87 |
245.52 |
0.11 |
0.19 |
| 2 |
A' |
3251 |
3070 |
4.55 |
19.66 |
0.42 |
0.60 |
| 3 |
A' |
3242 |
3062 |
4.48 |
75.81 |
0.73 |
0.85 |
| 4 |
A' |
3231 |
3052 |
0.53 |
65.35 |
0.72 |
0.84 |
| 5 |
A' |
3219 |
3040 |
2.37 |
43.94 |
0.39 |
0.57 |
| 6 |
A' |
2961 |
2797 |
103.52 |
110.66 |
0.27 |
0.43 |
| 7 |
A' |
1769 |
1671 |
144.58 |
77.79 |
0.32 |
0.48 |
| 8 |
A' |
1646 |
1554 |
11.83 |
47.98 |
0.54 |
0.70 |
| 9 |
A' |
1633 |
1542 |
7.39 |
8.76 |
0.56 |
0.71 |
| 10 |
A' |
1523 |
1438 |
0.47 |
1.65 |
0.25 |
0.40 |
| 11 |
A' |
1486 |
1404 |
9.61 |
1.21 |
0.11 |
0.20 |
| 12 |
A' |
1475 |
1393 |
3.63 |
1.58 |
0.73 |
0.85 |
| 13 |
A' |
1429 |
1350 |
11.47 |
2.70 |
0.66 |
0.80 |
| 14 |
A' |
1332 |
1258 |
14.12 |
1.71 |
0.44 |
0.61 |
| 15 |
A' |
1245 |
1176 |
53.69 |
24.86 |
0.23 |
0.38 |
| 16 |
A' |
1196 |
1130 |
8.91 |
6.00 |
0.30 |
0.46 |
| 17 |
A' |
1190 |
1123 |
1.81 |
6.33 |
0.69 |
0.82 |
| 18 |
A' |
1104 |
1042 |
3.99 |
0.71 |
0.25 |
0.40 |
| 19 |
A' |
1048 |
990 |
1.45 |
11.14 |
0.09 |
0.17 |
| 20 |
A' |
1007 |
952 |
0.42 |
21.55 |
0.09 |
0.16 |
| 21 |
A' |
841 |
794 |
31.72 |
9.21 |
0.17 |
0.28 |
| 22 |
A' |
655 |
618 |
17.64 |
3.49 |
0.32 |
0.48 |
| 23 |
A' |
614 |
580 |
0.67 |
4.47 |
0.75 |
0.86 |
| 24 |
A' |
437 |
412 |
0.30 |
4.57 |
0.32 |
0.49 |
| 25 |
A' |
217 |
205 |
7.37 |
0.53 |
0.50 |
0.67 |
| 26 |
A" |
1040 |
982 |
0.32 |
4.82 |
0.75 |
0.86 |
| 27 |
A" |
985 |
930 |
0.14 |
0.04 |
0.75 |
0.86 |
| 28 |
A" |
962 |
909 |
0.13 |
0.00 |
0.75 |
0.86 |
| 29 |
A" |
928 |
876 |
1.19 |
0.88 |
0.75 |
0.86 |
| 30 |
A" |
873 |
825 |
0.03 |
3.19 |
0.75 |
0.86 |
| 31 |
A" |
761 |
719 |
55.18 |
0.97 |
0.75 |
0.86 |
| 32 |
A" |
684 |
646 |
5.45 |
0.02 |
0.75 |
0.86 |
| 33 |
A" |
461 |
435 |
4.58 |
0.26 |
0.75 |
0.86 |
| 34 |
A" |
411 |
388 |
0.08 |
0.01 |
0.75 |
0.86 |
| 35 |
A" |
236 |
223 |
6.15 |
1.02 |
0.75 |
0.86 |
| 36 |
A" |
119 |
112 |
3.84 |
2.07 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 24230.6 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 22885.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.