return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5OCH3 (Anisole)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-345.905907
Energy at 298.15K-345.914872
HF Energy-344.616978
Nuclear repulsion energy346.520965
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3271 3089 2.47      
2 A' 3258 3077 7.75      
3 A' 3250 3070 6.60      
4 A' 3236 3056 6.21      
5 A' 3228 3049 1.36      
6 A' 3217 3039 15.17      
7 A' 3075 2904 37.05      
8 A' 1654 1562 63.23      
9 A' 1635 1544 16.57      
10 A' 1543 1458 1.53      
11 A' 1536 1451 110.52      
12 A' 1497 1414 29.40      
13 A' 1487 1405 0.41      
14 A' 1465 1383 9.41      
15 A' 1337 1263 0.89      
16 A' 1310 1238 210.21      
17 A' 1226 1158 7.02      
18 A' 1204 1137 9.28      
19 A' 1183 1118 1.08      
20 A' 1109 1047 11.78      
21 A' 1097 1036 36.56      
22 A' 1050 991 1.53      
23 A' 1008 952 0.53      
24 A' 801 756 14.41      
25 A' 614 580 0.37      
26 A' 556 525 4.60      
27 A' 445 421 0.74      
28 A' 268 253 2.32      
29 A" 3156 2981 26.83      
30 A" 1524 1440 6.31      
31 A" 1197 1131 0.72      
32 A" 958 905 0.08      
33 A" 937 885 0.18      
34 A" 890 841 4.58      
35 A" 840 793 0.07      
36 A" 768 725 61.76      
37 A" 678 641 6.65      
38 A" 518 489 4.59      
39 A" 418 395 0.01      
40 A" 301 284 0.16      
41 A" 220 208 0.52      
42 A" 90 85 4.10      

Unscaled Zero Point Vibrational Energy (zpe) 29525.8 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 27887.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.16810 0.05277 0.04047

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.537 0.000
C2 0.929 -0.507 0.000
C3 0.477 -1.829 0.000
C4 -0.883 -2.120 0.000
C5 -1.805 -1.071 0.000
C6 -1.369 0.247 0.000
O7 0.317 1.860 0.000
C8 1.695 2.173 0.000
H9 1.992 -0.310 0.000
H10 1.205 -2.632 0.000
H11 -1.223 -3.148 0.000
H12 -2.868 -1.280 0.000
H13 -2.067 1.076 0.000
H14 1.756 3.260 0.000
H15 2.197 1.783 0.891
H16 2.197 1.783 -0.891

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 C8 H9 H10 H11 H12 H13 H14 H15 H16
C11.39702.41312.79922.41671.39901.36082.35662.16483.39023.88233.39512.13603.24062.67852.6785
C21.39701.39732.42582.79132.41842.44442.78751.08122.14333.40703.87543.38833.85652.76512.7651
C32.41311.39731.39062.40442.77823.69224.18352.14521.08422.15193.39043.86145.24714.09874.0987
C42.79922.42581.39061.39662.41664.15665.00783.39712.14991.08312.15633.40835.99225.05095.0509
C52.41672.79132.40441.39661.38843.61824.77243.87243.39082.15721.08412.16305.60684.99524.9952
C61.39902.41842.77822.41661.38842.33303.61923.40703.86243.39862.14021.08364.34083.98333.9833
O71.36082.44443.69224.15663.61822.33301.41342.74144.57905.23954.47272.50952.00782.08152.0815
C82.35662.78754.18355.00784.77243.61921.41342.50134.83046.06925.72303.91891.08831.09441.0944
H92.16481.08122.14523.39713.87243.40702.74142.50132.45164.28854.95654.28933.57802.28462.2846
H103.39022.14331.08422.14993.39083.86244.57904.83042.45162.48254.29254.94555.91784.61244.6124
H113.88233.40702.15191.08312.15723.39865.23956.06924.28852.48252.48994.30787.06686.06696.0669
H123.39513.87543.39042.15631.08412.14024.47275.72304.95654.29252.48992.48856.48055.98605.9860
H132.13603.38833.86143.40832.16301.08362.50953.91894.28934.94554.30782.48854.40284.41264.4126
H143.24063.85655.24715.99225.60684.34082.00781.08833.57805.91787.06686.48054.40281.78001.7800
H152.67852.76514.09875.05094.99523.98332.08151.09442.28464.61246.06695.98604.41261.78001.7825
H162.67852.76514.09875.05094.99523.98332.08151.09442.28464.61246.06695.98604.41261.78001.7825

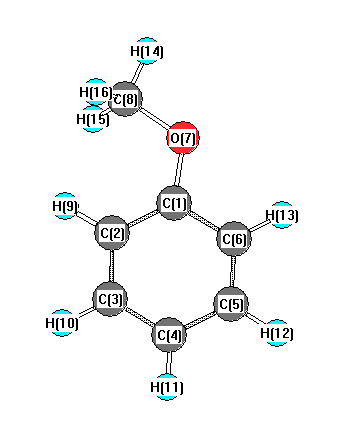
picture of Anisole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.444 C1 C2 H9 121.217
C1 C6 C5 120.226 C1 C6 H13 118.170
C1 O7 C8 116.296 C2 C1 C6 119.756
C2 C1 O7 124.834 C2 C3 C4 120.948
C2 C3 H10 118.935 C3 C2 H9 119.339
C3 C4 C5 119.237 C3 C4 H11 120.385
C4 C3 H10 120.116 C4 C5 C6 120.389
C4 C5 H12 120.215 C5 C4 H11 120.378
C5 C6 H13 121.604 C6 C1 O7 115.410
C6 C5 H12 119.395 O7 C8 H14 106.024
O7 C8 H15 111.567 O7 C8 H16 111.567
H14 C8 H15 109.279 H14 C8 H16 109.279
H15 C8 H16 109.050
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2/6-31G(2df,p) Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.461      
2 C -0.298      
3 C -0.135      
4 C -0.177      
5 C -0.135      
6 C -0.222      
7 O -0.478      
8 C -0.203      
9 H 0.151      
10 H 0.148      
11 H 0.143      
12 H 0.149      
13 H 0.156      
14 H 0.160      
15 H 0.140      
16 H 0.140      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 272.164
(<r2>)1/2 16.497