return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,4-dichlorobenzene)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1AG
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-1149.784875
Energy at 298.15K-1149.789050
HF Energy-1148.531069
Nuclear repulsion energy449.949819
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3266 3085 0.00      
2 Ag 1618 1529 0.00      
3 Ag 1201 1135 0.00      
4 Ag 1133 1071 0.00      
5 Ag 761 719 0.00      
6 Ag 332 314 0.00      
7 Au 938 886 0.00      
8 Au 412 389 0.00      
9 B1g 836 790 0.00      
10 B1u 3252 3071 0.04      
11 B1u 1505 1421 101.28      
12 B1u 1126 1063 90.67      
13 B1u 1018 961 52.53      
14 B1u 568 537 22.80      
15 B2g 940 888 0.00      
16 B2g 676 638 0.00      
17 B2g 300 283 0.00      
18 B2u 3265 3084 0.12      
19 B2u 1468 1387 0.71      
20 B2u 1407 1329 6.75      
21 B2u 1124 1061 5.44      
22 B2u 223 210 1.06      
23 B3g 3253 3072 0.00      
24 B3g 1618 1528 0.00      
25 B3g 1309 1236 0.00      
26 B3g 627 592 0.00      
27 B3g 354 334 0.00      
28 B3u 834 788 47.35      
29 B3u 494 466 11.82      
30 B3u 103 97 0.64      

Unscaled Zero Point Vibrational Energy (zpe) 17980.4 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 16982.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.18894 0.02239 0.02002

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.380
C2 0.000 0.000 -1.380
C3 0.000 1.213 0.696
C4 0.000 -1.213 0.696
C5 0.000 -1.213 -0.696
C6 0.000 1.213 -0.696
Cl7 0.000 0.000 3.113
Cl8 0.000 0.000 -3.113
H9 0.000 2.146 1.247
H10 0.000 -2.146 1.247
H11 0.000 -2.146 -1.247
H12 0.000 2.146 -1.247

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C12.76101.39301.39302.40522.40521.73234.49332.14972.14973.39233.3923
C22.76102.40522.40521.39301.39304.49331.73233.39233.39232.14972.1497
C31.39302.40522.42672.79781.39252.70413.99761.08283.40373.88052.1554
C41.39302.40522.42671.39252.79782.70413.99763.40371.08282.15543.8805
C52.40521.39302.79781.39252.42673.99762.70413.88052.15541.08283.4037
C62.40521.39301.39252.79782.42673.99762.70412.15543.88053.40371.0828
Cl71.73234.49332.70412.70413.99763.99766.22562.84322.84324.85924.8592
Cl84.49331.73233.99763.99762.70412.70416.22564.85924.85922.84322.8432
H92.14973.39231.08283.40373.88052.15542.84324.85924.29104.96332.4942
H102.14973.39233.40371.08282.15543.88052.84324.85924.29102.49424.9633
H113.39232.14973.88052.15541.08283.40374.85922.84324.96332.49424.2910
H123.39232.14972.15543.88053.40371.08284.85922.84322.49424.96334.2910

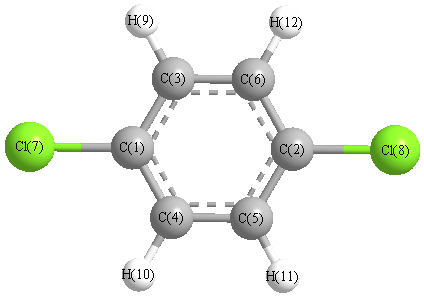
picture of 1,4-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.420 C1 C3 H9 119.999
C1 C4 C5 119.420 C1 C4 H10 119.999
C2 C5 C4 119.420 C2 C5 H11 119.999
C2 C6 C3 119.420 C2 C6 H12 119.999
C3 C1 C4 121.160 C3 C1 Cl7 119.420
C3 C6 H12 120.581 C4 C1 Cl7 119.420
C4 C5 H11 120.581 C5 C2 C6 121.160
C5 C2 Cl8 119.420 C5 C4 H10 120.581
C6 C2 Cl8 119.420 C6 C3 H9 120.581
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability