Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -5218.848942 |
| Energy at 298.15K | -5218.859222 |
| HF Energy | -5218.214416 |
| Nuclear repulsion energy | 423.374397 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3154 |
2979 |
0.00 |
|
|
|
| 2 |
Ag |
1504 |
1421 |
0.00 |
|
|
|
| 3 |
Ag |
1314 |
1241 |
0.00 |
|
|
|
| 4 |
Ag |
1090 |
1029 |
0.00 |
|
|
|
| 5 |
Ag |
734 |
694 |
0.00 |
|
|
|
| 6 |
Ag |
204 |
192 |
0.00 |
|
|
|
| 7 |
Au |
3242 |
3062 |
0.05 |
|
|
|
| 8 |
Au |
1133 |
1070 |
3.14 |
|
|
|
| 9 |
Au |
766 |
723 |
3.53 |
|
|
|
| 10 |
Au |
111 |
104 |
3.91 |
|
|
|
| 11 |
Bg |
3221 |
3042 |
0.00 |
|
|
|
| 12 |
Bg |
1307 |
1235 |
0.00 |
|
|
|
| 13 |
Bg |
971 |
917 |
0.00 |
|
|
|
| 14 |
Bu |
3162 |
2987 |
4.55 |
|
|
|
| 15 |
Bu |
1504 |
1420 |
6.44 |
|
|
|
| 16 |
Bu |
1233 |
1165 |
41.61 |
|
|
|
| 17 |
Bu |
669 |
632 |
43.44 |
|
|
|
| 18 |
Bu |
190 |
179 |
5.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12753.7 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12045.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.489 |
0.577 |
0.000 |
| C2 |
-0.489 |
-0.577 |
0.000 |
| Br3 |
-0.489 |
2.227 |
0.000 |
| Br4 |
0.489 |
-2.227 |
0.000 |
| H5 |
1.119 |
0.560 |
0.887 |
| H6 |
1.119 |
0.560 |
-0.887 |
| H7 |
-1.119 |
-0.560 |
0.887 |
| H8 |
-1.119 |
-0.560 |
-0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5125 | 1.9188 | 2.8037 | 1.0885 | 1.0885 | 2.1605 | 2.1605 |
C2 | 1.5125 | | 2.8037 | 1.9188 | 2.1605 | 2.1605 | 1.0885 | 1.0885 | Br3 | 1.9188 | 2.8037 | | 4.5604 | 2.4807 | 2.4807 | 2.9923 | 2.9923 | Br4 | 2.8037 | 1.9188 | 4.5604 | | 2.9923 | 2.9923 | 2.4807 | 2.4807 | H5 | 1.0885 | 2.1605 | 2.4807 | 2.9923 | | 1.7749 | 2.5034 | 3.0688 | H6 | 1.0885 | 2.1605 | 2.4807 | 2.9923 | 1.7749 | | 3.0688 | 2.5034 | H7 | 2.1605 | 1.0885 | 2.9923 | 2.4807 | 2.5034 | 3.0688 | | 1.7749 | H8 | 2.1605 | 1.0885 | 2.9923 | 2.4807 | 3.0688 | 2.5034 | 1.7749 | |
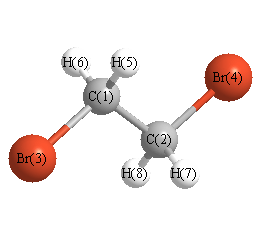 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
109.014 |
|
C1 |
C2 |
H7 |
111.288 |
| C1 |
C2 |
H8 |
111.288 |
|
C2 |
C1 |
Br3 |
109.014 |
| C2 |
C1 |
H5 |
111.288 |
|
C2 |
C1 |
H6 |
111.288 |
| Br3 |
C1 |
H5 |
107.949 |
|
Br3 |
C1 |
H6 |
107.949 |
| Br4 |
C2 |
H7 |
107.949 |
|
Br4 |
C2 |
H8 |
107.949 |
| H5 |
C1 |
H6 |
109.237 |
|
H7 |
C2 |
H8 |
109.237 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -5218.847283 |
| Energy at 298.15K | |
| HF Energy | -5218.211021 |
| Nuclear repulsion energy | 462.374073 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3209 |
3031 |
0.02 |
77.40 |
0.74 |
0.85 |
| 2 |
A |
3141 |
2967 |
9.43 |
160.81 |
0.02 |
0.04 |
| 3 |
A |
1490 |
1407 |
0.17 |
4.74 |
0.74 |
0.85 |
| 4 |
A |
1329 |
1256 |
15.78 |
2.26 |
0.40 |
0.57 |
| 5 |
A |
1215 |
1147 |
1.15 |
8.29 |
0.73 |
0.85 |
| 6 |
A |
1057 |
998 |
0.44 |
2.21 |
0.75 |
0.86 |
| 7 |
A |
941 |
889 |
5.64 |
3.56 |
0.36 |
0.53 |
| 8 |
A |
615 |
581 |
4.87 |
16.20 |
0.06 |
0.11 |
| 9 |
A |
236 |
223 |
0.87 |
0.74 |
0.28 |
0.43 |
| 10 |
A |
90 |
85 |
0.18 |
0.65 |
0.64 |
0.78 |
| 11 |
B |
3220 |
3042 |
0.23 |
26.44 |
0.75 |
0.86 |
| 12 |
B |
3136 |
2962 |
1.40 |
34.75 |
0.75 |
0.86 |
| 13 |
B |
1486 |
1404 |
10.96 |
12.72 |
0.75 |
0.86 |
| 14 |
B |
1301 |
1228 |
41.38 |
0.60 |
0.75 |
0.86 |
| 15 |
B |
1147 |
1083 |
1.65 |
2.89 |
0.75 |
0.86 |
| 16 |
B |
870 |
822 |
13.65 |
0.73 |
0.75 |
0.86 |
| 17 |
B |
649 |
613 |
6.53 |
6.07 |
0.75 |
0.86 |
| 18 |
B |
372 |
352 |
4.21 |
1.59 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12752.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12044.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.303 |
0.694 |
1.192 |
| C2 |
-0.303 |
-0.694 |
1.192 |
| Br3 |
-0.303 |
1.731 |
-0.297 |
| Br4 |
0.303 |
-1.731 |
-0.297 |
| H5 |
0.004 |
1.223 |
2.097 |
| H6 |
1.390 |
0.648 |
1.143 |
| H7 |
-0.004 |
-1.223 |
2.097 |
| H8 |
-1.390 |
-0.648 |
1.143 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5137 | 1.9130 | 2.8454 | 1.0908 | 1.0889 | 2.1420 | 2.1603 |
C2 | 1.5137 | | 2.8454 | 1.9130 | 2.1420 | 2.1603 | 1.0908 | 1.0889 | Br3 | 1.9130 | 2.8454 | | 3.5150 | 2.4667 | 2.4721 | 3.8145 | 2.9860 | Br4 | 2.8454 | 1.9130 | 3.5150 | | 3.8145 | 2.9860 | 2.4667 | 2.4721 | H5 | 1.0908 | 2.1420 | 2.4667 | 3.8145 | | 1.7782 | 2.4464 | 2.5206 | H6 | 1.0889 | 2.1603 | 2.4721 | 2.9860 | 1.7782 | | 2.5206 | 3.0667 | H7 | 2.1420 | 1.0908 | 3.8145 | 2.4667 | 2.4464 | 2.5206 | | 1.7782 | H8 | 2.1603 | 1.0889 | 2.9860 | 2.4721 | 2.5206 | 3.0667 | 1.7782 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
111.744 |
|
C1 |
C2 |
H7 |
109.588 |
| C1 |
C2 |
H8 |
111.156 |
|
C2 |
C1 |
Br3 |
111.744 |
| C2 |
C1 |
H5 |
109.588 |
|
C2 |
C1 |
H6 |
111.156 |
| Br3 |
C1 |
H5 |
107.216 |
|
Br3 |
C1 |
H6 |
107.687 |
| Br4 |
C2 |
H7 |
107.216 |
|
Br4 |
C2 |
H8 |
107.687 |
| H5 |
C1 |
H6 |
109.336 |
|
H7 |
C2 |
H8 |
109.336 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
