Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -191.449715 |
| Energy at 298.15K | |
| HF Energy | -190.780641 |
| Nuclear repulsion energy | 103.007588 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3311 |
3127 |
1.81 |
51.00 |
0.56 |
0.72 |
| 2 |
A' |
3255 |
3075 |
0.32 |
81.99 |
0.29 |
0.45 |
| 3 |
A' |
3208 |
3029 |
1.18 |
62.75 |
0.11 |
0.19 |
| 4 |
A' |
2960 |
2795 |
81.45 |
79.24 |
0.27 |
0.42 |
| 5 |
A' |
1773 |
1675 |
130.62 |
48.63 |
0.35 |
0.52 |
| 6 |
A' |
1674 |
1581 |
2.91 |
7.34 |
0.18 |
0.31 |
| 7 |
A' |
1469 |
1388 |
9.90 |
3.34 |
0.58 |
0.74 |
| 8 |
A' |
1408 |
1330 |
4.13 |
18.53 |
0.42 |
0.59 |
| 9 |
A' |
1301 |
1229 |
3.62 |
5.60 |
0.32 |
0.49 |
| 10 |
A' |
1183 |
1117 |
30.65 |
10.04 |
0.65 |
0.79 |
| 11 |
A' |
929 |
877 |
17.47 |
1.01 |
0.07 |
0.14 |
| 12 |
A' |
570 |
538 |
4.26 |
4.41 |
0.29 |
0.45 |
| 13 |
A' |
314 |
297 |
9.21 |
0.38 |
0.37 |
0.54 |
| 14 |
A" |
1056 |
998 |
10.76 |
1.00 |
0.75 |
0.86 |
| 15 |
A" |
1035 |
977 |
8.23 |
3.11 |
0.75 |
0.86 |
| 16 |
A" |
994 |
939 |
29.24 |
0.20 |
0.75 |
0.86 |
| 17 |
A" |
622 |
588 |
7.49 |
2.87 |
0.75 |
0.86 |
| 18 |
A" |
168 |
159 |
3.84 |
1.07 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13615.0 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12859.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.153 |
-0.740 |
0.000 |
| C2 |
0.000 |
0.724 |
0.000 |
| C3 |
1.221 |
1.273 |
0.000 |
| O4 |
-1.223 |
-1.318 |
0.000 |
| H5 |
0.805 |
-1.299 |
0.000 |
| H6 |
-0.908 |
1.316 |
0.000 |
| H7 |
1.373 |
2.344 |
0.000 |
| H8 |
2.106 |
0.647 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4726 | 2.4374 | 1.2155 | 1.1097 | 2.1905 | 3.4416 | 2.6512 |
C2 | 1.4726 | | 1.3382 | 2.3805 | 2.1778 | 1.0837 | 2.1237 | 2.1076 | C3 | 2.4374 | 1.3382 | | 3.5615 | 2.6055 | 2.1289 | 1.0819 | 1.0847 | O4 | 1.2155 | 2.3805 | 3.5615 | | 2.0281 | 2.6530 | 4.4892 | 3.8655 | H5 | 1.1097 | 2.1778 | 2.6055 | 2.0281 | | 3.1264 | 3.6872 | 2.3405 | H6 | 2.1905 | 1.0837 | 2.1289 | 2.6530 | 3.1264 | | 2.5023 | 3.0875 | H7 | 3.4416 | 2.1237 | 1.0819 | 4.4892 | 3.6872 | 2.5023 | | 1.8488 | H8 | 2.6512 | 2.1076 | 1.0847 | 3.8655 | 2.3405 | 3.0875 | 1.8488 | |
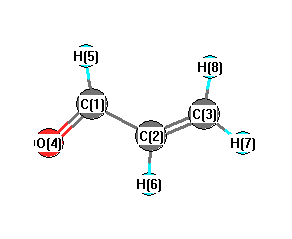 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.182 |
|
C1 |
C2 |
H6 |
117.118 |
| C2 |
C1 |
O4 |
124.366 |
|
C2 |
C1 |
H5 |
114.262 |
| C2 |
C3 |
H7 |
122.332 |
|
C2 |
C3 |
H8 |
120.522 |
| C3 |
C2 |
H6 |
122.700 |
|
O4 |
C1 |
H5 |
121.372 |
| H7 |
C3 |
H8 |
117.146 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -191.447314 |
| Energy at 298.15K | |
| HF Energy | -190.777546 |
| Nuclear repulsion energy | 104.676618 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3321 |
3137 |
0.40 |
43.46 |
0.62 |
0.77 |
| 2 |
A' |
3248 |
3068 |
5.82 |
95.24 |
0.25 |
0.40 |
| 3 |
A' |
3214 |
3036 |
0.99 |
53.80 |
0.13 |
0.22 |
| 4 |
A' |
2993 |
2827 |
117.16 |
129.45 |
0.30 |
0.47 |
| 5 |
A' |
1769 |
1671 |
61.44 |
11.74 |
0.35 |
0.51 |
| 6 |
A' |
1676 |
1583 |
37.98 |
28.93 |
0.21 |
0.34 |
| 7 |
A' |
1449 |
1369 |
25.00 |
8.55 |
0.69 |
0.82 |
| 8 |
A' |
1440 |
1360 |
12.87 |
7.49 |
0.23 |
0.37 |
| 9 |
A' |
1320 |
1246 |
1.32 |
11.84 |
0.42 |
0.59 |
| 10 |
A' |
1070 |
1011 |
4.05 |
4.00 |
0.75 |
0.86 |
| 11 |
A' |
952 |
900 |
53.34 |
3.24 |
0.14 |
0.25 |
| 12 |
A' |
679 |
642 |
10.25 |
0.90 |
0.55 |
0.71 |
| 13 |
A' |
290 |
274 |
6.87 |
2.18 |
0.44 |
0.61 |
| 14 |
A" |
1049 |
991 |
22.42 |
1.94 |
0.75 |
0.86 |
| 15 |
A" |
1042 |
985 |
3.57 |
4.27 |
0.75 |
0.86 |
| 16 |
A" |
1004 |
948 |
16.62 |
0.11 |
0.75 |
0.86 |
| 17 |
A" |
577 |
545 |
7.40 |
4.58 |
0.75 |
0.86 |
| 18 |
A" |
168 |
159 |
5.67 |
0.66 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13631.1 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12874.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.891 |
-0.282 |
0.000 |
| C2 |
0.000 |
0.904 |
0.000 |
| C3 |
1.330 |
0.756 |
0.000 |
| O4 |
-0.493 |
-1.431 |
0.000 |
| H5 |
-1.975 |
-0.062 |
0.000 |
| H6 |
-0.465 |
1.883 |
0.000 |
| H7 |
2.003 |
1.603 |
0.000 |
| H8 |
1.754 |
-0.241 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4831 | 2.4511 | 1.2166 | 1.1064 | 2.2061 | 3.4532 | 2.6459 |
C2 | 1.4831 | | 1.3378 | 2.3867 | 2.1988 | 1.0841 | 2.1211 | 2.0949 | C3 | 2.4511 | 1.3378 | | 2.8471 | 3.4046 | 2.1195 | 1.0818 | 1.0836 | O4 | 1.2166 | 2.3867 | 2.8471 | | 2.0181 | 3.3144 | 3.9287 | 2.5433 | H5 | 1.1064 | 2.1988 | 3.4046 | 2.0181 | | 2.4621 | 4.3122 | 3.7341 | H6 | 2.2061 | 1.0841 | 2.1195 | 3.3144 | 2.4621 | | 2.4838 | 3.0723 | H7 | 3.4532 | 2.1211 | 1.0818 | 3.9287 | 4.3122 | 2.4838 | | 1.8604 | H8 | 2.6459 | 2.0949 | 1.0836 | 2.5433 | 3.7341 | 3.0723 | 1.8604 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.579 |
|
C1 |
C2 |
H6 |
117.647 |
| C2 |
C1 |
O4 |
123.975 |
|
C2 |
C1 |
H5 |
115.466 |
| C2 |
C3 |
H7 |
122.119 |
|
C2 |
C3 |
H8 |
119.437 |
| C3 |
C2 |
H6 |
121.774 |
|
O4 |
C1 |
H5 |
120.558 |
| H7 |
C3 |
H8 |
118.443 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
