Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -3108.288488 |
| Energy at 298.15K | |
| HF Energy | -3107.632188 |
| Nuclear repulsion energy | 291.633753 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3164 |
2989 |
6.21 |
2.17 |
0.11 |
0.20 |
| 2 |
A' |
3156 |
2981 |
0.40 |
119.58 |
0.06 |
0.12 |
| 3 |
A' |
1510 |
1426 |
4.28 |
4.54 |
0.75 |
0.86 |
| 4 |
A' |
1506 |
1423 |
2.31 |
9.11 |
0.75 |
0.86 |
| 5 |
A' |
1337 |
1263 |
2.69 |
9.42 |
0.43 |
0.60 |
| 6 |
A' |
1248 |
1178 |
33.68 |
1.38 |
0.28 |
0.44 |
| 7 |
A' |
1093 |
1032 |
1.38 |
4.68 |
0.70 |
0.83 |
| 8 |
A' |
793 |
749 |
25.55 |
35.35 |
0.26 |
0.41 |
| 9 |
A' |
706 |
666 |
39.46 |
8.53 |
0.20 |
0.34 |
| 10 |
A' |
261 |
246 |
0.73 |
3.50 |
0.27 |
0.42 |
| 11 |
A' |
202 |
191 |
6.27 |
0.08 |
0.16 |
0.28 |
| 12 |
A" |
3245 |
3065 |
0.02 |
0.35 |
0.75 |
0.86 |
| 13 |
A" |
3225 |
3046 |
0.07 |
76.19 |
0.75 |
0.86 |
| 14 |
A" |
1309 |
1237 |
0.06 |
6.73 |
0.75 |
0.86 |
| 15 |
A" |
1150 |
1086 |
2.10 |
0.15 |
0.75 |
0.86 |
| 16 |
A" |
997 |
942 |
0.13 |
4.04 |
0.75 |
0.86 |
| 17 |
A" |
774 |
731 |
3.02 |
0.00 |
0.75 |
0.86 |
| 18 |
A" |
119 |
112 |
5.35 |
0.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12896.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12180.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.641 |
0.000 |
| C2 |
1.209 |
-0.269 |
0.000 |
| Br3 |
-1.584 |
-0.439 |
0.000 |
| Cl4 |
2.692 |
0.728 |
0.000 |
| H5 |
-0.020 |
1.269 |
0.888 |
| H6 |
-0.020 |
1.269 |
-0.888 |
| H7 |
1.232 |
-0.897 |
0.889 |
| H8 |
1.232 |
-0.897 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5133 | 1.9170 | 2.6930 | 1.0882 | 1.0882 | 2.1618 | 2.1618 |
C2 | 1.5133 | | 2.7982 | 1.7867 | 2.1600 | 2.1600 | 1.0884 | 1.0884 | Br3 | 1.9170 | 2.7982 | | 4.4319 | 2.4807 | 2.4807 | 2.9885 | 2.9885 | Cl4 | 2.6930 | 1.7867 | 4.4319 | | 2.9039 | 2.9039 | 2.3580 | 2.3580 | H5 | 1.0882 | 2.1600 | 2.4807 | 2.9039 | | 1.7762 | 2.5021 | 3.0688 | H6 | 1.0882 | 2.1600 | 2.4807 | 2.9039 | 1.7762 | | 3.0688 | 2.5021 | H7 | 2.1618 | 1.0884 | 2.9885 | 2.3580 | 2.5021 | 3.0688 | | 1.7776 | H8 | 2.1618 | 1.0884 | 2.9885 | 2.3580 | 3.0688 | 2.5021 | 1.7776 | |
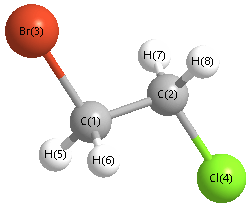 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.104 |
|
C1 |
C2 |
H7 |
111.343 |
| C1 |
C2 |
H8 |
111.343 |
|
C2 |
C1 |
Br3 |
108.749 |
| C2 |
C1 |
Cl4 |
38.824 |
|
C2 |
C1 |
H6 |
111.211 |
| Br3 |
C1 |
H5 |
108.081 |
|
Br3 |
C1 |
H6 |
108.081 |
| Cl4 |
C2 |
H7 |
107.711 |
|
Cl4 |
C2 |
H8 |
107.711 |
| H5 |
C1 |
H6 |
109.401 |
|
H7 |
C2 |
H8 |
109.492 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -3108.286393 |
| Energy at 298.15K | |
| HF Energy | -3107.628855 |
| Nuclear repulsion energy | 309.191526 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3223 |
3044 |
0.43 |
24.75 |
0.75 |
0.85 |
| 2 |
A |
3211 |
3033 |
0.19 |
79.94 |
0.74 |
0.85 |
| 3 |
A |
3144 |
2969 |
9.61 |
142.30 |
0.03 |
0.05 |
| 4 |
A |
3138 |
2963 |
3.86 |
51.27 |
0.39 |
0.56 |
| 5 |
A |
1496 |
1413 |
3.90 |
7.83 |
0.73 |
0.85 |
| 6 |
A |
1487 |
1405 |
7.02 |
10.04 |
0.75 |
0.86 |
| 7 |
A |
1350 |
1275 |
19.36 |
1.87 |
0.73 |
0.85 |
| 8 |
A |
1315 |
1242 |
34.64 |
1.17 |
0.32 |
0.49 |
| 9 |
A |
1231 |
1163 |
1.51 |
9.44 |
0.72 |
0.84 |
| 10 |
A |
1166 |
1101 |
1.12 |
3.47 |
0.73 |
0.84 |
| 11 |
A |
1065 |
1006 |
1.70 |
2.57 |
0.75 |
0.85 |
| 12 |
A |
962 |
909 |
7.48 |
3.72 |
0.35 |
0.52 |
| 13 |
A |
893 |
844 |
17.32 |
1.20 |
0.69 |
0.82 |
| 14 |
A |
718 |
678 |
13.76 |
7.70 |
0.34 |
0.51 |
| 15 |
A |
632 |
597 |
6.56 |
13.16 |
0.14 |
0.24 |
| 16 |
A |
398 |
376 |
5.59 |
1.61 |
0.71 |
0.83 |
| 17 |
A |
250 |
236 |
1.02 |
0.71 |
0.28 |
0.44 |
| 18 |
A |
108 |
102 |
0.54 |
0.87 |
0.70 |
0.82 |
Unscaled Zero Point Vibrational Energy (zpe) 12892.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 12176.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.014 |
1.106 |
-0.381 |
| C2 |
1.266 |
0.907 |
0.400 |
| Br3 |
-1.320 |
-0.227 |
0.037 |
| Cl4 |
2.115 |
-0.579 |
-0.088 |
| H5 |
-0.445 |
2.075 |
-0.127 |
| H6 |
0.173 |
1.065 |
-1.453 |
| H7 |
1.944 |
1.740 |
0.209 |
| H8 |
1.069 |
0.836 |
1.469 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5134 | 1.9120 | 2.7312 | 1.0906 | 1.0887 | 2.1418 | 2.1608 |
C2 | 1.5134 | | 2.8474 | 1.7797 | 2.1380 | 2.1577 | 1.0908 | 1.0889 | Br3 | 1.9120 | 2.8474 | | 3.4555 | 2.4679 | 2.4736 | 3.8151 | 2.9813 | Cl4 | 2.7312 | 1.7797 | 3.4555 | | 3.6878 | 2.8882 | 2.3444 | 2.3495 | H5 | 1.0906 | 2.1380 | 2.4679 | 3.6878 | | 1.7772 | 2.4360 | 2.5246 | H6 | 1.0887 | 2.1577 | 2.4736 | 2.8882 | 1.7772 | | 2.5212 | 3.0648 | H7 | 2.1418 | 1.0908 | 3.8151 | 2.3444 | 2.4360 | 2.5212 | | 1.7803 | H8 | 2.1608 | 1.0889 | 2.9813 | 2.3495 | 2.5246 | 3.0648 | 1.7803 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
111.812 |
|
C1 |
C2 |
H7 |
109.587 |
| C1 |
C2 |
H8 |
111.214 |
|
C2 |
C1 |
Br3 |
111.933 |
| C2 |
C1 |
Cl4 |
37.227 |
|
C2 |
C1 |
H6 |
110.982 |
| Br3 |
C1 |
H5 |
107.376 |
|
Br3 |
C1 |
H6 |
107.872 |
| Cl4 |
C2 |
H7 |
107.060 |
|
Cl4 |
C2 |
H8 |
107.525 |
| H5 |
C1 |
H6 |
109.276 |
|
H7 |
C2 |
H8 |
109.525 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
