Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -612.531275 |
| Energy at 298.15K | -612.534577 |
| HF Energy | -611.830687 |
| Nuclear repulsion energy | 143.312743 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3228 |
3049 |
0.19 |
|
|
|
| 2 |
A |
3139 |
2965 |
5.92 |
|
|
|
| 3 |
A |
3013 |
2846 |
50.59 |
|
|
|
| 4 |
A |
1793 |
1693 |
98.10 |
|
|
|
| 5 |
A |
1475 |
1393 |
9.20 |
|
|
|
| 6 |
A |
1425 |
1346 |
4.50 |
|
|
|
| 7 |
A |
1282 |
1211 |
15.03 |
|
|
|
| 8 |
A |
1200 |
1133 |
5.02 |
|
|
|
| 9 |
A |
1073 |
1013 |
29.33 |
|
|
|
| 10 |
A |
1042 |
984 |
1.66 |
|
|
|
| 11 |
A |
846 |
799 |
20.65 |
|
|
|
| 12 |
A |
725 |
685 |
10.65 |
|
|
|
| 13 |
A |
464 |
438 |
9.39 |
|
|
|
| 14 |
A |
290 |
274 |
9.45 |
|
|
|
| 15 |
A |
61 |
58 |
12.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10527.7 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 9943.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.011 |
0.689 |
0.139 |
| C2 |
1.153 |
-0.304 |
0.233 |
| Cl3 |
-1.527 |
-0.177 |
-0.085 |
| O4 |
2.243 |
-0.087 |
-0.243 |
| H5 |
-0.060 |
1.257 |
1.068 |
| H6 |
0.173 |
1.366 |
-0.697 |
| H7 |
0.918 |
-1.234 |
0.782 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5163 | 1.7784 | 2.3940 | 1.0915 | 1.0879 | 2.2208 |
C2 | 1.5163 | | 2.7018 | 1.2090 | 2.1462 | 2.1478 | 1.1056 | Cl3 | 1.7784 | 2.7018 | | 3.7743 | 2.3529 | 2.3753 | 2.8009 | O4 | 2.3940 | 1.2090 | 3.7743 | | 2.9717 | 2.5695 | 2.0307 | H5 | 1.0915 | 2.1462 | 2.3529 | 2.9717 | | 1.7837 | 2.6911 | H6 | 1.0879 | 2.1478 | 2.3753 | 2.5695 | 1.7837 | | 3.0823 | H7 | 2.2208 | 1.1056 | 2.8009 | 2.0307 | 2.6911 | 3.0823 | |
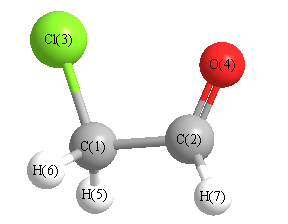 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
122.515 |
|
C1 |
C2 |
H7 |
114.883 |
| C2 |
C1 |
Cl3 |
109.929 |
|
C2 |
C1 |
H5 |
109.693 |
| C2 |
C1 |
H6 |
110.041 |
|
Cl3 |
C1 |
H5 |
107.725 |
| Cl3 |
C1 |
H6 |
109.565 |
|
O4 |
C2 |
H7 |
122.597 |
| H5 |
C1 |
H6 |
109.853 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -612.530722 |
| Energy at 298.15K | -612.534209 |
| HF Energy | -611.828657 |
| Nuclear repulsion energy | 147.380008 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3123 |
2950 |
12.38 |
|
|
|
| 2 |
A' |
2972 |
2807 |
82.79 |
|
|
|
| 3 |
A' |
1810 |
1709 |
82.17 |
|
|
|
| 4 |
A' |
1465 |
1383 |
20.47 |
|
|
|
| 5 |
A' |
1421 |
1342 |
13.81 |
|
|
|
| 6 |
A' |
1322 |
1249 |
16.88 |
|
|
|
| 7 |
A' |
971 |
917 |
5.40 |
|
|
|
| 8 |
A' |
794 |
750 |
9.35 |
|
|
|
| 9 |
A' |
653 |
616 |
37.87 |
|
|
|
| 10 |
A' |
216 |
204 |
2.80 |
|
|
|
| 11 |
A" |
3186 |
3009 |
0.00 |
|
|
|
| 12 |
A" |
1220 |
1152 |
1.67 |
|
|
|
| 13 |
A" |
1048 |
990 |
0.73 |
|
|
|
| 14 |
A" |
701 |
662 |
1.62 |
|
|
|
| 15 |
A" |
176 |
166 |
0.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10537.9 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 9953.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.929 |
0.000 |
| C2 |
1.359 |
0.262 |
0.000 |
| Cl3 |
-1.331 |
-0.227 |
0.000 |
| O4 |
1.556 |
-0.928 |
0.000 |
| H5 |
-0.080 |
1.568 |
0.882 |
| H6 |
-0.080 |
1.568 |
-0.882 |
| H7 |
2.190 |
0.996 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5136 | 1.7631 | 2.4227 | 1.0924 | 1.0924 | 2.1912 |
C2 | 1.5136 | | 2.7342 | 1.2060 | 2.1339 | 2.1339 | 1.1090 | Cl3 | 1.7631 | 2.7342 | | 2.9715 | 2.3594 | 2.3594 | 3.7278 | O4 | 2.4227 | 1.2060 | 2.9715 | | 3.1119 | 3.1119 | 2.0255 | H5 | 1.0924 | 2.1339 | 2.3594 | 3.1119 | | 1.7649 | 2.5016 | H6 | 1.0924 | 2.1339 | 2.3594 | 3.1119 | 1.7649 | | 2.5016 | H7 | 2.1912 | 1.1090 | 3.7278 | 2.0255 | 2.5016 | 2.5016 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
125.567 |
|
C1 |
C2 |
H7 |
112.419 |
| C2 |
C1 |
Cl3 |
112.895 |
|
C2 |
C1 |
H5 |
108.859 |
| C2 |
C1 |
H6 |
108.859 |
|
Cl3 |
C1 |
H5 |
109.170 |
| Cl3 |
C1 |
H6 |
109.170 |
|
O4 |
C2 |
H7 |
122.014 |
| H5 |
C1 |
H6 |
107.759 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
