Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -208.643144 |
| Energy at 298.15K | -208.649185 |
| HF Energy | -207.908606 |
| Nuclear repulsion energy | 117.422461 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3871 |
3656 |
96.55 |
|
|
|
| 2 |
A' |
3224 |
3045 |
4.67 |
|
|
|
| 3 |
A' |
3176 |
3000 |
8.26 |
|
|
|
| 4 |
A' |
3105 |
2933 |
10.67 |
|
|
|
| 5 |
A' |
1708 |
1613 |
0.67 |
|
|
|
| 6 |
A' |
1512 |
1429 |
7.00 |
|
|
|
| 7 |
A' |
1459 |
1378 |
8.33 |
|
|
|
| 8 |
A' |
1415 |
1336 |
23.19 |
|
|
|
| 9 |
A' |
1293 |
1221 |
52.14 |
|
|
|
| 10 |
A' |
1166 |
1101 |
4.74 |
|
|
|
| 11 |
A' |
1039 |
982 |
115.11 |
|
|
|
| 12 |
A' |
923 |
871 |
6.92 |
|
|
|
| 13 |
A' |
569 |
537 |
13.86 |
|
|
|
| 14 |
A' |
322 |
304 |
2.37 |
|
|
|
| 15 |
A" |
3184 |
3007 |
7.59 |
|
|
|
| 16 |
A" |
1508 |
1424 |
7.39 |
|
|
|
| 17 |
A" |
1086 |
1026 |
0.43 |
|
|
|
| 18 |
A" |
919 |
868 |
8.90 |
|
|
|
| 19 |
A" |
405 |
383 |
126.82 |
|
|
|
| 20 |
A" |
289 |
273 |
2.13 |
|
|
|
| 21 |
A" |
211 |
199 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16191.2 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 15292.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.302 |
1.293 |
0.000 |
| C2 |
0.000 |
0.564 |
0.000 |
| N3 |
0.014 |
-0.712 |
0.000 |
| O4 |
1.314 |
-1.200 |
0.000 |
| H5 |
1.167 |
-2.149 |
0.000 |
| H6 |
-2.129 |
0.584 |
0.000 |
| H7 |
-1.382 |
1.935 |
0.880 |
| H8 |
-1.382 |
1.935 |
-0.880 |
| H9 |
0.934 |
1.130 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4924 | 2.3984 | 3.6136 | 4.2359 | 1.0890 | 1.0921 | 1.0921 | 2.2422 |
C2 | 1.4924 | | 1.2762 | 2.1995 | 2.9534 | 2.1287 | 2.1358 | 2.1358 | 1.0917 | N3 | 2.3984 | 1.2762 | | 1.3881 | 1.8422 | 2.5038 | 3.1187 | 3.1187 | 2.0585 | O4 | 3.6136 | 2.1995 | 1.3881 | | 0.9602 | 3.8767 | 4.2267 | 4.2267 | 2.3605 | H5 | 4.2359 | 2.9534 | 1.8422 | 0.9602 | | 4.2808 | 4.8933 | 4.8933 | 3.2869 | H6 | 1.0890 | 2.1287 | 2.5038 | 3.8767 | 4.2808 | | 1.7771 | 1.7771 | 3.1108 | H7 | 1.0921 | 2.1358 | 3.1187 | 4.2267 | 4.8933 | 1.7771 | | 1.7604 | 2.6047 | H8 | 1.0921 | 2.1358 | 3.1187 | 4.2267 | 4.8933 | 1.7771 | 1.7604 | | 2.6047 | H9 | 2.2422 | 1.0917 | 2.0585 | 2.3605 | 3.2869 | 3.1108 | 2.6047 | 2.6047 | |
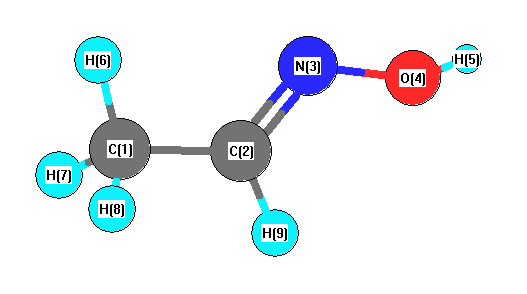 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
119.857 |
|
C1 |
C2 |
H9 |
119.577 |
| C2 |
C1 |
H6 |
110.119 |
|
C2 |
C1 |
H7 |
110.504 |
| C2 |
C1 |
H8 |
110.504 |
|
C2 |
N3 |
O4 |
111.219 |
| N3 |
C2 |
H9 |
120.566 |
|
N3 |
O4 |
H5 |
101.793 |
| H6 |
C1 |
H7 |
109.124 |
|
H6 |
C1 |
H8 |
109.124 |
| H7 |
C1 |
H8 |
107.408 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -208.642956 |
| Energy at 298.15K | |
| HF Energy | -207.907071 |
| Nuclear repulsion energy | 119.998913 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3872 |
3657 |
97.68 |
|
|
|
| 2 |
A' |
3250 |
3069 |
2.38 |
|
|
|
| 3 |
A' |
3230 |
3050 |
4.89 |
|
|
|
| 4 |
A' |
3112 |
2939 |
7.03 |
|
|
|
| 5 |
A' |
1716 |
1621 |
2.09 |
|
|
|
| 6 |
A' |
1508 |
1424 |
12.99 |
|
|
|
| 7 |
A' |
1421 |
1342 |
21.42 |
|
|
|
| 8 |
A' |
1393 |
1316 |
28.61 |
|
|
|
| 9 |
A' |
1350 |
1275 |
38.01 |
|
|
|
| 10 |
A' |
1160 |
1095 |
9.00 |
|
|
|
| 11 |
A' |
963 |
909 |
113.83 |
|
|
|
| 12 |
A' |
931 |
879 |
0.05 |
|
|
|
| 13 |
A' |
675 |
637 |
12.84 |
|
|
|
| 14 |
A' |
318 |
301 |
1.45 |
|
|
|
| 15 |
A" |
3187 |
3010 |
5.66 |
|
|
|
| 16 |
A" |
1515 |
1431 |
8.34 |
|
|
|
| 17 |
A" |
1070 |
1011 |
0.22 |
|
|
|
| 18 |
A" |
873 |
824 |
12.46 |
|
|
|
| 19 |
A" |
501 |
473 |
33.33 |
|
|
|
| 20 |
A" |
383 |
362 |
82.80 |
|
|
|
| 21 |
A" |
53i |
50i |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16186.3 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 15288.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.443 |
0.480 |
0.000 |
| C2 |
0.000 |
0.865 |
0.000 |
| N3 |
1.021 |
0.087 |
0.000 |
| O4 |
0.647 |
-1.261 |
0.000 |
| H5 |
1.506 |
-1.693 |
0.000 |
| H6 |
-1.561 |
-0.601 |
0.000 |
| H7 |
-1.941 |
0.895 |
0.879 |
| H8 |
-1.941 |
0.895 |
-0.879 |
| H9 |
0.271 |
1.916 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4939 | 2.4957 | 2.7206 | 3.6635 | 1.0865 | 1.0921 | 1.0921 | 2.2366 |
C2 | 1.4939 | | 1.2835 | 2.2222 | 2.9683 | 2.1408 | 2.1313 | 2.1313 | 1.0853 | N3 | 2.4957 | 1.2835 | | 1.3993 | 1.8451 | 2.6721 | 3.1940 | 3.1940 | 1.9762 | O4 | 2.7206 | 2.2222 | 1.3993 | | 0.9614 | 2.3048 | 3.4816 | 3.4816 | 3.1990 | H5 | 3.6635 | 2.9683 | 1.8451 | 0.9614 | | 3.2558 | 4.3995 | 4.3995 | 3.8140 | H6 | 1.0865 | 2.1408 | 2.6721 | 2.3048 | 3.2558 | | 1.7756 | 1.7756 | 3.1125 | H7 | 1.0921 | 2.1313 | 3.1940 | 3.4816 | 4.3995 | 1.7756 | | 1.7576 | 2.5905 | H8 | 1.0921 | 2.1313 | 3.1940 | 3.4816 | 4.3995 | 1.7756 | 1.7576 | | 2.5905 | H9 | 2.2366 | 1.0853 | 1.9762 | 3.1990 | 3.8140 | 3.1125 | 2.5905 | 2.5905 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.781 |
|
C1 |
C2 |
H9 |
119.412 |
| C2 |
C1 |
H6 |
111.139 |
|
C2 |
C1 |
H7 |
110.038 |
| C2 |
C1 |
H8 |
110.038 |
|
C2 |
N3 |
O4 |
111.780 |
| N3 |
C2 |
H9 |
112.808 |
|
N3 |
O4 |
H5 |
101.184 |
| H6 |
C1 |
H7 |
109.184 |
|
H6 |
C1 |
H8 |
109.184 |
| H7 |
C1 |
H8 |
107.165 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
