Jump to
S1C2
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -209.080724 |
| Energy at 298.15K | |
| HF Energy | -209.080724 |
| Nuclear repulsion energy | 120.301511 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3918 |
3918 |
92.36 |
|
|
|
| 2 |
A' |
3202 |
3202 |
6.73 |
|
|
|
| 3 |
A' |
3179 |
3179 |
7.42 |
|
|
|
| 4 |
A' |
3065 |
3065 |
12.50 |
|
|
|
| 5 |
A' |
1784 |
1784 |
6.72 |
|
|
|
| 6 |
A' |
1492 |
1492 |
17.07 |
|
|
|
| 7 |
A' |
1420 |
1420 |
24.99 |
|
|
|
| 8 |
A' |
1403 |
1403 |
39.04 |
|
|
|
| 9 |
A' |
1358 |
1358 |
29.99 |
|
|
|
| 10 |
A' |
1158 |
1158 |
10.97 |
|
|
|
| 11 |
A' |
999 |
999 |
135.87 |
|
|
|
| 12 |
A' |
929 |
929 |
2.91 |
|
|
|
| 13 |
A' |
691 |
691 |
13.27 |
|
|
|
| 14 |
A' |
321 |
321 |
1.22 |
|
|
|
| 15 |
A" |
3128 |
3128 |
11.69 |
|
|
|
| 16 |
A" |
1487 |
1487 |
8.59 |
|
|
|
| 17 |
A" |
1072 |
1072 |
0.14 |
|
|
|
| 18 |
A" |
868 |
868 |
11.67 |
|
|
|
| 19 |
A" |
514 |
514 |
30.86 |
|
|
|
| 20 |
A" |
389 |
389 |
85.64 |
|
|
|
| 21 |
A" |
60i |
60i |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16158.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16158.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.448 |
0.482 |
0.000 |
| C2 |
0.000 |
0.857 |
0.000 |
| N3 |
1.010 |
0.085 |
0.000 |
| O4 |
0.663 |
-1.257 |
0.000 |
| H5 |
1.518 |
-1.690 |
0.000 |
| H6 |
-1.580 |
-0.599 |
0.000 |
| H7 |
-1.943 |
0.904 |
0.881 |
| H8 |
-1.943 |
0.904 |
-0.881 |
| H9 |
0.264 |
1.912 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4961 | 2.4904 | 2.7350 | 3.6761 | 1.0887 | 1.0948 | 1.0948 | 2.2309 |
C2 | 1.4961 | | 1.2715 | 2.2153 | 2.9647 | 2.1485 | 2.1336 | 2.1336 | 1.0874 | N3 | 2.4904 | 1.2715 | | 1.3858 | 1.8458 | 2.6790 | 3.1887 | 3.1887 | 1.9736 | O4 | 2.7350 | 2.2153 | 1.3858 | | 0.9582 | 2.3376 | 3.4981 | 3.4981 | 3.1937 | H5 | 3.6761 | 2.9647 | 1.8458 | 0.9582 | | 3.2843 | 4.4138 | 4.4138 | 3.8137 | H6 | 1.0887 | 2.1485 | 2.6790 | 2.3376 | 3.2843 | | 1.7796 | 1.7796 | 3.1151 | H7 | 1.0948 | 2.1336 | 3.1887 | 3.4981 | 4.4138 | 1.7796 | | 1.7611 | 2.5807 | H8 | 1.0948 | 2.1336 | 3.1887 | 3.4981 | 4.4138 | 1.7796 | 1.7611 | | 2.5807 | H9 | 2.2309 | 1.0874 | 1.9736 | 3.1937 | 3.8137 | 3.1151 | 2.5807 | 2.5807 | |
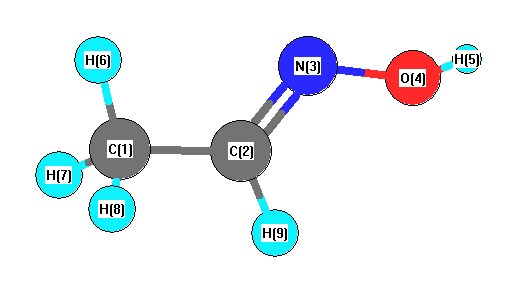 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.086 |
|
C1 |
C2 |
H9 |
118.565 |
| C2 |
C1 |
H6 |
111.472 |
|
C2 |
C1 |
H7 |
109.905 |
| C2 |
C1 |
H8 |
109.905 |
|
C2 |
N3 |
O4 |
112.883 |
| N3 |
C2 |
H9 |
113.349 |
|
N3 |
O4 |
H5 |
102.362 |
| H6 |
C1 |
H7 |
109.182 |
|
H6 |
C1 |
H8 |
109.182 |
| H7 |
C1 |
H8 |
107.088 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.495 |
|
|
|
| 2 |
C |
0.039 |
|
|
|
| 3 |
N |
-0.081 |
|
|
|
| 4 |
O |
-0.422 |
|
|
|
| 5 |
H |
0.328 |
|
|
|
| 6 |
H |
0.177 |
|
|
|
| 7 |
H |
0.156 |
|
|
|
| 8 |
H |
0.156 |
|
|
|
| 9 |
H |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.644 |
0.119 |
0.000 |
0.655 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.455 |
-3.791 |
0.000 |
| y |
-3.791 |
-20.642 |
0.000 |
| z |
0.000 |
0.000 |
-24.578 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.155 |
-3.791 |
0.000 |
| y |
-3.791 |
2.874 |
0.000 |
| z |
0.000 |
0.000 |
-3.029 |
|
| Polar |
|---|
| 3z2-r2 | -6.058 |
|---|
| x2-y2 | -1.813 |
|---|
| xy | -3.791 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.730 |
-1.179 |
0.000 |
| y |
-1.179 |
5.901 |
0.000 |
| z |
0.000 |
0.000 |
3.362 |
<r2> (average value of r
2) Å
2
| <r2> |
78.296 |
| (<r2>)1/2 |
8.849 |
