All results from a given calculation for C6H13N (cyclohexanamine)
using model chemistry: MP2/6-31G(2df,p)
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C1 |
1A |
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -290.445895 |
| Energy at 298.15K | |
| HF Energy | -289.267175 |
| Nuclear repulsion energy | 331.182944 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.870 |
0.011 |
0.294 |
| C2 |
1.174 |
-1.248 |
-0.223 |
| C3 |
-0.304 |
-1.258 |
0.166 |
| C4 |
-1.024 |
-0.010 |
-0.332 |
| C5 |
-0.318 |
1.246 |
0.185 |
| C6 |
1.159 |
1.266 |
-0.211 |
| N7 |
-2.441 |
-0.104 |
0.025 |
| H8 |
2.920 |
0.019 |
-0.010 |
| H9 |
1.857 |
0.007 |
1.390 |
| H10 |
1.259 |
-1.283 |
-1.314 |
| H11 |
1.671 |
-2.144 |
0.160 |
| H12 |
-0.812 |
-2.143 |
-0.225 |
| H13 |
-0.392 |
-1.296 |
1.260 |
| H14 |
-0.972 |
-0.004 |
-1.429 |
| H15 |
-0.405 |
1.265 |
1.279 |
| H16 |
-0.831 |
2.139 |
-0.190 |
| H17 |
1.645 |
2.166 |
0.177 |
| H18 |
1.239 |
1.311 |
-1.303 |
| H19 |
-2.917 |
0.740 |
-0.281 |
| H20 |
-2.515 |
-0.098 |
1.039 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
N7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
| C1 | | 1.5286 | 2.5208 | 2.9611 | 2.5148 | 1.5284 | 4.3209 | 1.0934 | 1.0960 | 2.1533 | 2.1682 | 3.4787 | 2.7851 | 3.3240 | 2.7780 | 3.4717 | 2.1696 | 2.1536 | 4.8762 | 4.4491 |
C2 | 1.5286 | | 1.5286 | 2.5255 | 2.9350 | 2.5145 | 3.8000 | 2.1672 | 2.1544 | 1.0959 | 1.0933 | 2.1783 | 2.1568 | 2.7589 | 3.3264 | 3.9356 | 3.4693 | 2.7782 | 4.5492 | 4.0652 | C3 | 2.5208 | 1.5286 | | 1.5250 | 2.5047 | 2.9423 | 2.4328 | 3.4722 | 2.7877 | 2.1533 | 2.1644 | 1.0925 | 1.0980 | 2.1363 | 2.7599 | 3.4557 | 3.9402 | 3.3374 | 3.3198 | 2.6453 | C4 | 2.9611 | 2.5255 | 1.5250 | | 1.5310 | 2.5318 | 1.4643 | 3.9572 | 3.3572 | 2.7930 | 3.4727 | 2.1469 | 2.1426 | 1.0979 | 2.1460 | 2.1615 | 3.4810 | 2.7941 | 2.0368 | 2.0276 | C5 | 2.5148 | 2.9350 | 2.5047 | 1.5310 | | 1.5293 | 2.5212 | 3.4683 | 2.7788 | 3.3369 | 3.9304 | 3.4499 | 2.7616 | 2.1440 | 1.0976 | 1.0952 | 2.1678 | 2.1546 | 2.6886 | 2.7135 | C6 | 1.5284 | 2.5145 | 2.9423 | 2.5318 | 1.5293 | | 3.8594 | 2.1673 | 2.1536 | 2.7801 | 3.4681 | 3.9383 | 3.3370 | 2.7644 | 2.1601 | 2.1727 | 1.0934 | 1.0960 | 4.1107 | 4.1137 | N7 | 4.3209 | 3.8000 | 2.4328 | 1.4643 | 2.5212 | 3.8594 | | 5.3625 | 4.5115 | 4.1082 | 4.5920 | 2.6223 | 2.6734 | 2.0689 | 2.7556 | 2.7691 | 4.6768 | 4.1602 | 1.0161 | 1.0167 | H8 | 1.0934 | 2.1672 | 3.4722 | 3.9572 | 3.4683 | 2.1673 | 5.3625 | | 1.7581 | 2.4805 | 2.5030 | 4.3179 | 3.7828 | 4.1428 | 3.7778 | 4.3119 | 2.5041 | 2.4829 | 5.8876 | 5.5367 | H9 | 1.0960 | 2.1544 | 2.7877 | 3.3572 | 2.7788 | 2.1536 | 4.5115 | 1.7581 | | 3.0556 | 2.4842 | 3.7887 | 2.6025 | 3.9946 | 2.5914 | 3.7773 | 2.4860 | 3.0556 | 5.1114 | 4.3879 | H10 | 2.1533 | 1.0959 | 2.1533 | 2.7930 | 3.3369 | 2.7801 | 4.1082 | 2.4805 | 3.0556 | | 1.7563 | 2.4930 | 3.0583 | 2.5750 | 3.9988 | 4.1642 | 3.7775 | 2.5939 | 4.7546 | 4.6033 | H11 | 2.1682 | 1.0933 | 2.1644 | 3.4727 | 3.9304 | 3.4681 | 4.5920 | 2.5030 | 2.4842 | 1.7563 | | 2.5121 | 2.4862 | 3.7538 | 4.1450 | 4.9716 | 4.3097 | 3.7762 | 5.4370 | 4.7414 | H12 | 3.4787 | 2.1783 | 1.0925 | 2.1469 | 3.4499 | 3.9383 | 2.6223 | 4.3179 | 3.7887 | 2.4930 | 2.5121 | | 1.7602 | 2.4602 | 3.7478 | 4.2820 | 4.9766 | 4.1589 | 3.5709 | 2.9469 | H13 | 2.7851 | 2.1568 | 1.0980 | 2.1426 | 2.7616 | 3.3370 | 2.6734 | 3.7828 | 2.6025 | 3.0583 | 2.4862 | 1.7602 | | 3.0393 | 2.5618 | 3.7540 | 4.1604 | 4.0030 | 3.5914 | 2.4485 | H14 | 3.3240 | 2.7589 | 2.1363 | 1.0979 | 2.1440 | 2.7644 | 2.0689 | 4.1428 | 3.9946 | 2.5750 | 3.7538 | 2.4602 | 3.0393 | | 3.0440 | 2.4790 | 3.7602 | 2.5758 | 2.3779 | 2.9118 | H15 | 2.7780 | 3.3264 | 2.7599 | 2.1460 | 1.0976 | 2.1601 | 2.7556 | 3.7778 | 2.5914 | 3.9988 | 4.1450 | 3.7478 | 2.5618 | 3.0440 | | 1.7611 | 2.4958 | 3.0613 | 3.0031 | 2.5235 | H16 | 3.4717 | 3.9356 | 3.4557 | 2.1615 | 1.0952 | 2.1727 | 2.7691 | 4.3119 | 3.7773 | 4.1642 | 4.9716 | 4.2820 | 3.7540 | 2.4790 | 1.7611 | | 2.5030 | 2.4916 | 2.5133 | 3.0573 | H17 | 2.1696 | 3.4693 | 3.9402 | 3.4810 | 2.1678 | 1.0934 | 4.6768 | 2.5041 | 2.4860 | 3.7775 | 4.3097 | 4.9766 | 4.1604 | 3.7602 | 2.4958 | 2.5030 | | 1.7568 | 4.8017 | 4.8140 | H18 | 2.1536 | 2.7782 | 3.3374 | 2.7941 | 2.1546 | 1.0960 | 4.1602 | 2.4829 | 3.0556 | 2.5939 | 3.7762 | 4.1589 | 4.0030 | 2.5758 | 3.0613 | 2.4916 | 1.7568 | | 4.3179 | 4.6434 | H19 | 4.8762 | 4.5492 | 3.3198 | 2.0368 | 2.6886 | 4.1107 | 1.0161 | 5.8876 | 5.1114 | 4.7546 | 5.4370 | 3.5709 | 3.5914 | 2.3779 | 3.0031 | 2.5133 | 4.8017 | 4.3179 | | 1.6137 | H20 | 4.4491 | 4.0652 | 2.6453 | 2.0276 | 2.7135 | 4.1137 | 1.0167 | 5.5367 | 4.3879 | 4.6033 | 4.7414 | 2.9469 | 2.4485 | 2.9118 | 2.5235 | 3.0573 | 4.8140 | 4.6434 | 1.6137 | |
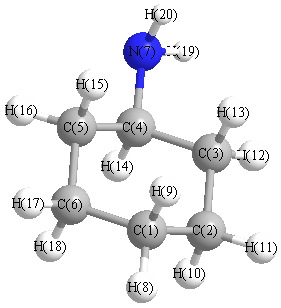 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.085 |
|
C1 |
C2 |
H10 |
109.150 |
| C1 |
C2 |
H11 |
110.470 |
|
C1 |
C6 |
C5 |
110.661 |
| C1 |
C6 |
H17 |
110.596 |
|
C1 |
C6 |
H18 |
109.184 |
| C2 |
C1 |
C6 |
110.680 |
|
C2 |
C1 |
H8 |
110.394 |
| C2 |
C1 |
H9 |
109.236 |
|
C2 |
C3 |
C4 |
111.595 |
| C2 |
C3 |
H12 |
111.324 |
|
C2 |
C3 |
H13 |
109.299 |
| C3 |
C2 |
H10 |
109.156 |
|
C3 |
C2 |
H11 |
110.172 |
| C3 |
C4 |
C5 |
110.092 |
|
C3 |
C4 |
N7 |
108.929 |
| C3 |
C4 |
H14 |
107.966 |
|
C4 |
C3 |
H12 |
109.093 |
| C4 |
C3 |
H13 |
108.441 |
|
C4 |
C5 |
C6 |
111.642 |
| C4 |
C5 |
H15 |
108.322 |
|
C4 |
C5 |
H16 |
109.674 |
| C4 |
N7 |
H19 |
109.061 |
|
C4 |
N7 |
H20 |
108.264 |
| C5 |
C4 |
N7 |
114.631 |
|
C5 |
C4 |
H14 |
108.154 |
| C5 |
C6 |
H17 |
110.387 |
|
C5 |
C6 |
H18 |
109.196 |
| C6 |
C1 |
H8 |
110.412 |
|
C6 |
C1 |
H9 |
109.186 |
| C6 |
C5 |
H15 |
109.530 |
|
C6 |
C5 |
H16 |
110.668 |
| N7 |
C4 |
H14 |
106.826 |
|
H8 |
C1 |
H9 |
106.841 |
| H10 |
C2 |
H11 |
106.693 |
|
H12 |
C3 |
H13 |
106.938 |
| H15 |
C5 |
H16 |
106.860 |
|
H17 |
C6 |
H18 |
106.720 |
| H19 |
N7 |
H20 |
105.087 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
