return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12 (Pentane)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-197.192778
Energy at 298.15K-197.205691
HF Energy-196.360057
Nuclear repulsion energy185.283701
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3196 3018 27.32      
2 A1 3102 2930 41.49      
3 A1 3099 2927 28.53      
4 A1 3080 2909 4.55      
5 A1 1541 1456 5.77      
6 A1 1526 1442 0.00      
7 A1 1512 1428 0.01      
8 A1 1433 1354 3.22      
9 A1 1379 1302 0.10      
10 A1 1183 1118 1.03      
11 A1 1081 1021 0.42      
12 A1 892 842 1.29      
13 A1 403 380 0.00      
14 A1 176 166 0.03      
15 A2 3194 3017 0.00      
16 A2 3144 2970 0.00      
17 A2 1532 1447 0.00      
18 A2 1347 1272 0.00      
19 A2 1283 1212 0.00      
20 A2 1000 945 0.00      
21 A2 771 728 0.00      
22 A2 262 248 0.00      
23 A2 110 104 0.00      
24 B1 3194 3017 62.61      
25 B1 3157 2982 36.13      
26 B1 3125 2952 0.56      
27 B1 1531 1446 12.42      
28 B1 1343 1268 0.51      
29 B1 1224 1156 0.13      
30 B1 873 824 1.40      
31 B1 740 699 4.46      
32 B1 253 239 0.00      
33 B1 117 111 0.00      
34 B2 3196 3018 18.41      
35 B2 3101 2929 29.76      
36 B2 3094 2923 0.18      
37 B2 1536 1451 1.41      
38 B2 1516 1432 0.52      
39 B2 1430 1351 0.95      
40 B2 1417 1338 1.50      
41 B2 1297 1225 3.02      
42 B2 1111 1049 0.44      
43 B2 1061 1002 1.94      
44 B2 952 899 3.19      
45 B2 395 373 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 35953.9 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 33958.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.56710 0.06528 0.06188

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.313
C2 0.000 1.273 -0.528
C3 0.000 -1.273 -0.528
C4 0.000 2.537 0.328
C5 0.000 -2.537 0.328
H6 0.877 0.000 0.971
H7 -0.877 0.000 0.971
H8 0.876 1.272 -1.184
H9 -0.876 1.272 -1.184
H10 -0.876 -1.272 -1.184
H11 0.876 -1.272 -1.184
H12 0.000 3.439 -0.287
H13 0.000 -3.439 -0.287
H14 -0.881 2.571 0.972
H15 0.881 2.571 0.972
H16 0.881 -2.571 0.972
H17 -0.881 -2.571 0.972

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17
C11.52591.52592.53752.53751.09601.09602.15072.15072.15072.15073.49123.49122.79682.79682.79682.7968
C21.52592.54681.52633.90572.15312.15311.09461.09462.77062.77062.17934.71892.17002.17004.21974.2197
C31.52592.54683.90571.52632.15312.15312.77062.77061.09461.09464.71892.17934.21974.21972.17002.1700
C42.53751.52633.90575.07492.76062.76062.15732.15734.19084.19081.09126.00821.09201.09205.22415.2241
C52.53753.90571.52635.07492.76062.76064.19084.19082.15732.15736.00821.09125.22415.22411.09201.0920
H61.09602.15312.15312.76062.76061.75302.50223.05493.05492.50223.76553.76553.11482.57132.57133.1148
H71.09602.15312.15312.76062.76061.75303.05492.50222.50223.05493.76553.76552.57133.11483.11482.5713
H82.15071.09462.77062.15734.19082.50223.05491.75213.08862.54362.50414.87513.06972.51684.40634.7438
H92.15071.09462.77062.15734.19083.05492.50221.75212.54363.08862.50414.87512.51683.06974.74384.4063
H102.15072.77061.09464.19082.15733.05492.50223.08862.54361.75214.87512.50414.40634.74383.06972.5168
H112.15072.77061.09464.19082.15732.50223.05492.54363.08861.75214.87512.50414.74384.40632.51683.0697
H123.49122.17934.71891.09126.00823.76553.76552.50412.50414.87514.87516.87861.76441.76446.20386.2038
H133.49124.71892.17936.00821.09123.76553.76554.87514.87512.50412.50416.87866.20386.20381.76441.7644
H142.79682.17004.21971.09205.22413.11482.57133.06972.51684.40634.74381.76446.20381.76265.43645.1427
H152.79682.17004.21971.09205.22412.57133.11482.51683.06974.74384.40631.76446.20381.76265.14275.4364
H162.79684.21972.17005.22411.09202.57133.11484.40634.74383.06972.51686.20381.76445.43645.14271.7626
H172.79684.21972.17005.22411.09203.11482.57134.74384.40632.51683.06976.20381.76445.14275.43641.7626

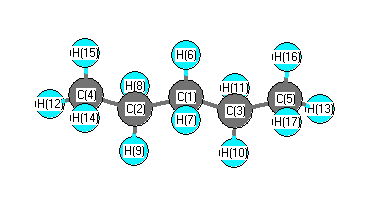
picture of Pentane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 112.482 C1 C2 H8 109.213
C1 C2 H9 109.213 C1 C3 C5 112.482
C1 C3 H10 109.213 C1 C3 H11 109.213
C2 C1 C3 113.133 C2 C1 H6 109.316
C2 C1 H7 109.316 C2 C4 H12 111.654
C2 C4 H14 110.863 C2 C4 H15 110.863
C3 C1 H6 109.316 C3 C1 H7 109.316
C3 C5 H13 111.654 C3 C5 H16 110.863
C3 C5 H17 110.863 C4 C2 H8 109.705
C4 C2 H9 109.705 C5 C3 H10 109.705
C5 C3 H11 109.705 H6 C1 H7 106.209
H8 C2 H9 106.332 H10 C3 H11 106.332
H12 C4 H14 107.835 H12 C4 H15 107.835
H13 C5 H16 107.835 H13 C5 H17 107.835
H14 C4 H15 107.626 H16 C5 H17 107.626
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability