return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(NH2)H2CH2CH2CH3 (1-Butanamine)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-213.213673
Energy at 298.15K-213.226383
Nuclear repulsion energy185.876551
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3545 3348 0.12      
2 A' 3196 3019 22.35      
3 A' 3103 2930 53.55      
4 A' 3101 2929 28.55      
5 A' 3096 2924 1.14      
6 A' 3077 2906 8.04      
7 A' 1682 1588 22.14      
8 A' 1539 1454 3.90      
9 A' 1530 1445 0.36      
10 A' 1517 1433 0.47      
11 A' 1509 1426 0.22      
12 A' 1432 1353 1.91      
13 A' 1415 1336 8.02      
14 A' 1389 1312 2.47      
15 A' 1295 1223 5.28      
16 A' 1168 1103 6.50      
17 A' 1122 1060 8.25      
18 A' 1098 1037 5.01      
19 A' 1048 990 26.06      
20 A' 934 882 55.03      
21 A' 896 846 117.31      
22 A' 432 408 4.20      
23 A' 393 371 0.55      
24 A' 179 169 1.91      
25 A" 3640 3438 1.26      
26 A" 3195 3018 31.04      
27 A" 3162 2986 46.99      
28 A" 3147 2973 1.40      
29 A" 3124 2951 1.62      
30 A" 1532 1447 6.27      
31 A" 1410 1332 0.75      
32 A" 1345 1270 0.30      
33 A" 1323 1250 0.00      
34 A" 1245 1176 0.02      
35 A" 1057 998 0.62      
36 A" 929 877 0.00      
37 A" 793 749 0.38      
38 A" 744 703 3.73      
39 A" 297 280 46.33      
40 A" 260 246 0.30      
41 A" 126 119 0.08      
42 A" 111 105 2.97      

Unscaled Zero Point Vibrational Energy (zpe) 33565.3 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 31702.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.59780 0.06522 0.06206

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 2.535 0.400 0.000
C2 1.328 -0.427 0.000
C3 0.000 0.329 0.000
C4 -1.211 -0.600 0.000
C5 -2.533 0.163 0.000
H6 2.511 1.016 0.806
H7 2.511 1.016 -0.806
H8 1.371 -1.083 0.875
H9 1.371 -1.083 -0.875
H10 -0.040 0.984 -0.878
H11 -0.040 0.984 0.878
H12 -1.163 -1.255 0.876
H13 -1.163 -1.255 -0.876
H14 -3.388 -0.515 0.000
H15 -2.612 0.803 0.881
H16 -2.612 0.803 -0.881

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.46312.53613.87695.07311.01491.01492.07822.07822.78252.78254.14474.14475.99305.23795.2379
C21.46311.52852.54473.90562.03232.03231.09431.09432.15292.15292.76712.76714.71674.22114.2211
C32.53611.52851.52602.53802.72472.72472.15372.15371.09631.09632.15112.15113.49122.79752.7975
C43.87692.54471.52601.52624.13624.13622.76842.76842.15672.15671.09461.09462.17872.16992.1699
C55.07313.90562.53801.52625.17785.17784.18984.18982.76752.76752.15702.15701.09121.09191.0919
H61.01492.03232.72474.13625.17781.61232.38892.92053.05642.55154.31894.63446.14675.12795.3979
H71.01492.03232.72474.13625.17781.61232.92052.38892.55153.05644.63444.31896.14675.39795.1279
H82.07821.09432.15372.76844.18982.38892.92051.75063.05552.50242.53963.08494.87174.40694.7441
H92.07821.09432.15372.76844.18982.92052.38891.75062.50243.05553.08492.53964.87174.74414.4069
H102.78252.15291.09632.15672.76753.05642.55153.05552.50241.75613.05782.50473.77183.12212.5791
H112.78252.15291.09632.15672.76752.55153.05642.50243.05551.75612.50473.05783.77182.57913.1221
H124.14472.76712.15111.09462.15704.31894.63442.53963.08493.05782.50471.75192.50292.51653.0694
H134.14472.76712.15111.09462.15704.63444.31893.08492.53962.50473.05781.75192.50293.06942.5165
H145.99304.71673.49122.17871.09126.14676.14674.87174.87173.77183.77182.50292.50291.76441.7644
H155.23794.22112.79752.16991.09195.12795.39794.40694.74413.12212.57912.51653.06941.76441.7628
H165.23794.22112.79752.16991.09195.39795.12794.74414.40692.57913.12213.06942.51651.76441.7628

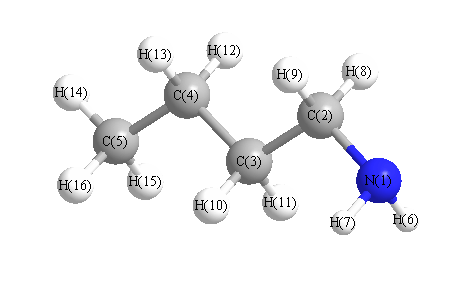
picture of 1-Butanamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 115.914 N1 C2 H8 107.838
N1 C2 H9 107.838 C2 N1 H6 108.854
C2 N1 H7 108.854 C2 C3 C4 112.840
C2 C3 H10 109.104 C2 C3 H11 109.104
C3 C2 H8 109.285 C3 C2 H9 109.285
C3 C4 C5 112.509 C3 C4 H12 109.233
C3 C4 H13 109.233 C4 C3 H10 109.574
C4 C3 H11 109.574 C4 C5 H14 111.613
C4 C5 H15 110.864 C4 C5 H16 110.864
C5 C4 H12 109.682 C5 C4 H13 109.682
H6 N1 H7 105.189 H8 C2 H9 106.245
H10 C3 H11 106.440 H12 C4 H13 106.309
H14 C5 H15 107.844 H14 C5 H16 107.844
H15 C5 H16 107.648
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability