return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2CN (Butanenitrile)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-210.836136
Energy at 298.15K-210.843426
HF Energy-210.016878
Nuclear repulsion energy154.062732
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3206 3028 13.48      
2 A' 3126 2952 14.63      
3 A' 3112 2939 0.22      
4 A' 3106 2934 12.14      
5 A' 2217 2094 0.01      
6 A' 1536 1451 5.64      
7 A' 1521 1437 1.38      
8 A' 1495 1412 2.89      
9 A' 1435 1356 2.31      
10 A' 1399 1321 2.08      
11 A' 1311 1239 1.72      
12 A' 1133 1070 3.12      
13 A' 1082 1022 0.09      
14 A' 976 922 1.53      
15 A' 895 845 1.84      
16 A' 528 498 0.67      
17 A' 347 327 0.10      
18 A' 163 154 5.90      
19 A" 3202 3024 24.15      
20 A" 3181 3004 0.85      
21 A" 3162 2986 0.00      
22 A" 1530 1446 7.67      
23 A" 1344 1269 0.01      
24 A" 1275 1204 0.01      
25 A" 1140 1077 0.21      
26 A" 881 832 0.17      
27 A" 752 710 3.96      
28 A" 384 363 0.65      
29 A" 252 238 0.01      
30 A" 101 96 4.04      

Unscaled Zero Point Vibrational Energy (zpe) 22894.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 21624.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.79184 0.07568 0.07184

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.615 0.331 0.000
C2 -1.451 0.482 0.000
C3 0.000 0.638 0.000
C4 0.722 -0.715 0.000
C5 2.236 -0.529 0.000
H6 0.289 1.221 0.879
H7 0.289 1.221 -0.879
H8 0.408 -1.286 0.877
H9 0.408 -1.286 -0.877
H10 2.747 -1.492 0.000
H11 2.563 0.024 0.883
H12 2.563 0.024 -0.883

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.17352.63313.49784.92663.16163.16163.53923.53925.66415.26145.2614
C21.17351.45972.48183.82332.08462.08462.71172.71174.63984.13534.1353
C32.63311.45971.53422.52211.09331.09332.15342.15343.47652.77902.7790
C43.49782.48181.53421.52482.17012.17011.09221.09222.16902.17072.1707
C54.92663.82332.52211.52482.76152.76152.16342.16341.09031.09171.0917
H63.16162.08461.09332.17012.76151.75792.50933.06273.76532.56963.1154
H73.16162.08461.09332.17012.76151.75793.06272.50933.76533.11542.5696
H83.53922.71172.15341.09222.16342.50933.06271.75402.50662.52103.0743
H93.53922.71172.15341.09222.16343.06272.50931.75402.50663.07432.5210
H105.66414.63983.47652.16901.09033.76533.76532.50662.50661.76391.7639
H115.26144.13532.77902.17071.09172.56963.11542.52103.07431.76391.7652
H125.26144.13532.77902.17071.09173.11542.56963.07432.52101.76391.7652

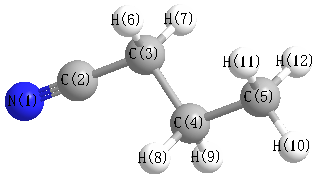
picture of Butanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 178.740 C2 C3 C4 111.956
C2 C3 H6 108.628 C2 C3 H7 108.628
C3 C4 C5 111.076 C3 C4 H8 108.987
C3 C4 H9 108.987 C4 C3 H6 110.233
C4 C3 H7 110.233 C4 C5 H10 110.987
C4 C5 H11 111.040 C4 C5 H12 111.040
C5 C4 H8 110.425 C5 C4 H9 110.425
H6 C3 H7 107.016 H8 C4 H9 106.828
H10 C5 H11 107.873 H10 C5 H12 107.873
H11 C5 H12 107.884
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability