return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9SH (1-Butanethiol)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-555.663730
Energy at 298.15K 
HF Energy-554.836293
Nuclear repulsion energy223.965446
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3199 3022 19.77      
2 A' 3124 2950 22.44      
3 A' 3105 2932 29.11      
4 A' 3102 2930 10.84      
5 A' 3094 2922 0.88      
6 A' 2785 2630 1.79      
7 A' 1539 1454 4.26      
8 A' 1526 1441 1.53      
9 A' 1514 1430 0.45      
10 A' 1510 1426 2.48      
11 A' 1433 1353 2.20      
12 A' 1407 1329 3.62      
13 A' 1346 1272 7.84      
14 A' 1263 1193 20.44      
15 A' 1154 1090 2.09      
16 A' 1103 1041 0.23      
17 A' 1064 1005 0.23      
18 A' 944 891 2.37      
19 A' 866 818 1.45      
20 A' 781 738 2.32      
21 A' 392 370 0.79      
22 A' 325 307 0.78      
23 A' 151 143 1.30      
24 A" 3198 3020 32.04      
25 A" 3192 3015 4.91      
26 A" 3160 2985 11.74      
27 A" 3141 2967 0.72      
28 A" 1531 1446 6.65      
29 A" 1347 1272 0.19      
30 A" 1323 1250 0.70      
31 A" 1246 1177 0.57      
32 A" 1092 1031 0.92      
33 A" 936 884 1.61      
34 A" 801 756 0.00      
35 A" 748 707 3.91      
36 A" 258 244 0.07      
37 A" 203 192 17.36      
38 A" 115 108 1.63      
39 A" 99 93 3.40      

Unscaled Zero Point Vibrational Energy (zpe) 29555.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 27915.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.53148 0.04494 0.04275

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.387 -1.829 0.000
C2 -0.230 -0.989 0.000
C3 0.000 0.518 0.000
C4 -1.316 1.293 0.000
C5 -1.090 2.803 0.000
H6 0.923 -3.083 0.000
H7 -0.792 -1.285 0.887
H8 -0.792 -1.285 -0.887
H9 0.591 0.795 -0.879
H10 0.591 0.795 0.879
H11 -1.904 1.007 0.877
H12 -1.904 1.007 -0.877
H13 -2.036 3.347 0.000
H14 -0.525 3.112 0.882
H15 -0.525 3.112 -0.882

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
S11.82252.72664.12975.25301.33682.41502.41502.87992.87994.43254.43256.20585.37115.3711
C21.82251.52402.52683.88772.39071.09111.09112.15112.15112.74872.74874.69684.20434.2043
C32.72661.52401.52702.53153.71732.15972.15971.09461.09462.15292.15293.48552.78932.7893
C44.12972.52681.52701.52634.91532.77602.77602.15762.15761.09431.09432.17662.16992.1699
C55.25303.88772.53151.52636.22044.19324.19322.76212.76212.15782.15781.09101.09171.0917
H61.33682.39073.71734.91536.22042.63842.63843.99023.99025.04875.04877.07816.42256.4225
H72.41501.09112.15972.77604.19322.63841.77383.05882.49782.54743.09844.87734.40454.7463
H82.41501.09112.15972.77604.19322.63841.77382.49783.05883.09842.54744.87734.74634.4045
H92.87992.15111.09462.15762.76213.99023.05882.49781.75743.05842.50433.76633.11632.5715
H102.87992.15111.09462.15762.76213.99022.49783.05881.75742.50433.05843.76632.57153.1163
H114.43252.74872.15291.09432.15785.04872.54743.09843.05842.50431.75402.50252.51613.0698
H124.43252.74872.15291.09432.15785.04873.09842.54742.50433.05841.75402.50253.06982.5161
H136.20584.69683.48552.17661.09107.07814.87734.87733.76633.76632.50252.50251.76461.7646
H145.37114.20432.78932.16991.09176.42254.40454.74633.11632.57152.51613.06981.76461.7633
H155.37114.20432.78932.16991.09176.42254.74634.40452.57153.11633.06982.51611.76461.7633

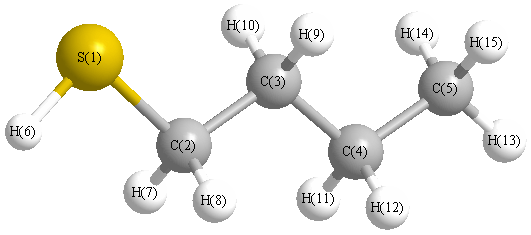
picture of 1-Butanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C3 108.800 S1 C2 H7 109.389
S1 C2 H8 109.389 C2 S1 H6 97.156
C2 C3 C4 111.828 C2 C3 H9 109.372
C2 C3 H10 109.372 C3 C2 H7 110.249
C3 C2 H8 110.249 C3 C4 C5 112.019
C3 C4 H11 109.319 C3 C4 H12 109.319
C4 C3 H9 109.673 C4 C3 H10 109.673
C4 C5 H13 111.456 C4 C5 H14 110.872
C4 C5 H15 110.872 C5 C4 H11 109.753
C5 C4 H12 109.753 H7 C2 H8 108.749
H9 C3 H10 106.791 H11 C4 H12 106.524
H13 C5 H14 107.888 H13 C5 H15 107.888
H14 C5 H15 107.713
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability