Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -230.619530 |
| Energy at 298.15K | -230.625289 |
| HF Energy | -229.787676 |
| Nuclear repulsion energy | 159.147513 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3354 |
3168 |
0.68 |
|
|
|
| 2 |
A1 |
3247 |
3067 |
2.34 |
|
|
|
| 3 |
A1 |
3220 |
3041 |
13.88 |
|
|
|
| 4 |
A1 |
1745 |
1648 |
2.68 |
|
|
|
| 5 |
A1 |
1448 |
1368 |
0.29 |
|
|
|
| 6 |
A1 |
1375 |
1299 |
5.93 |
|
|
|
| 7 |
A1 |
1224 |
1156 |
17.31 |
|
|
|
| 8 |
A1 |
874 |
825 |
2.90 |
|
|
|
| 9 |
A1 |
527 |
498 |
0.54 |
|
|
|
| 10 |
A1 |
235 |
222 |
0.00 |
|
|
|
| 11 |
A2 |
983 |
929 |
0.00 |
|
|
|
| 12 |
A2 |
852 |
805 |
0.00 |
|
|
|
| 13 |
A2 |
733 |
692 |
0.00 |
|
|
|
| 14 |
A2 |
105 |
99 |
0.00 |
|
|
|
| 15 |
B1 |
993 |
938 |
39.47 |
|
|
|
| 16 |
B1 |
855 |
807 |
87.81 |
|
|
|
| 17 |
B1 |
701 |
662 |
4.74 |
|
|
|
| 18 |
B1 |
72 |
68 |
1.32 |
|
|
|
| 19 |
B2 |
3354 |
3168 |
0.14 |
|
|
|
| 20 |
B2 |
3246 |
3066 |
2.02 |
|
|
|
| 21 |
B2 |
3214 |
3035 |
9.98 |
|
|
|
| 22 |
B2 |
1699 |
1605 |
392.09 |
|
|
|
| 23 |
B2 |
1438 |
1358 |
63.17 |
|
|
|
| 24 |
B2 |
1343 |
1269 |
11.42 |
|
|
|
| 25 |
B2 |
1240 |
1171 |
378.90 |
|
|
|
| 26 |
B2 |
1037 |
980 |
4.46 |
|
|
|
| 27 |
B2 |
482 |
455 |
5.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19796.5 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 18697.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.343 |
| C2 |
0.000 |
1.161 |
-0.371 |
| C3 |
0.000 |
-1.161 |
-0.371 |
| C4 |
0.000 |
2.349 |
0.228 |
| C5 |
0.000 |
-2.349 |
0.228 |
| H6 |
0.000 |
1.044 |
-1.450 |
| H7 |
0.000 |
-1.044 |
-1.450 |
| H8 |
0.000 |
3.244 |
-0.374 |
| H9 |
0.000 |
2.435 |
1.305 |
| H10 |
0.000 |
-3.244 |
-0.374 |
| H11 |
0.000 |
-2.435 |
1.305 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3633 | 1.3633 | 2.3523 | 2.3523 | 2.0757 | 2.0757 | 3.3225 | 2.6179 | 3.3225 | 2.6179 |
C2 | 1.3633 | | 2.3226 | 1.3306 | 3.5615 | 1.0861 | 2.4558 | 2.0827 | 2.1048 | 4.4053 | 3.9675 | C3 | 1.3633 | 2.3226 | | 3.5615 | 1.3306 | 2.4558 | 1.0861 | 4.4053 | 3.9675 | 2.0827 | 2.1048 | C4 | 2.3523 | 1.3306 | 3.5615 | | 4.6990 | 2.1263 | 3.7864 | 1.0786 | 1.0801 | 5.6259 | 4.9041 | C5 | 2.3523 | 3.5615 | 1.3306 | 4.6990 | | 3.7864 | 2.1263 | 5.6259 | 4.9041 | 1.0786 | 1.0801 | H6 | 2.0757 | 1.0861 | 2.4558 | 2.1263 | 3.7864 | | 2.0888 | 2.4486 | 3.0864 | 4.4213 | 4.4383 | H7 | 2.0757 | 2.4558 | 1.0861 | 3.7864 | 2.1263 | 2.0888 | | 4.4213 | 4.4383 | 2.4486 | 3.0864 | H8 | 3.3225 | 2.0827 | 4.4053 | 1.0786 | 5.6259 | 2.4486 | 4.4213 | | 1.8642 | 6.4880 | 5.9221 | H9 | 2.6179 | 2.1048 | 3.9675 | 1.0801 | 4.9041 | 3.0864 | 4.4383 | 1.8642 | | 5.9221 | 4.8699 | H10 | 3.3225 | 4.4053 | 2.0827 | 5.6259 | 1.0786 | 4.4213 | 2.4486 | 6.4880 | 5.9221 | | 1.8642 | H11 | 2.6179 | 3.9675 | 2.1048 | 4.9041 | 1.0801 | 4.4383 | 3.0864 | 5.9221 | 4.8699 | 1.8642 | |
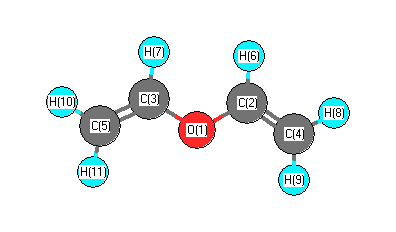 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
121.662 |
|
O1 |
C2 |
H6 |
115.406 |
| O1 |
C3 |
C5 |
121.662 |
|
O1 |
C3 |
H7 |
115.406 |
| C2 |
O1 |
C3 |
116.828 |
|
C2 |
C4 |
H8 |
119.273 |
| C2 |
C4 |
H9 |
121.290 |
|
C3 |
C5 |
H10 |
119.273 |
| C3 |
C5 |
H11 |
121.290 |
|
C4 |
C2 |
H6 |
122.932 |
| C5 |
C3 |
H7 |
122.932 |
|
H8 |
C4 |
H9 |
119.436 |
| H10 |
C5 |
H11 |
119.436 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -230.620807 |
| Energy at 298.15K | -230.626917 |
| HF Energy | -229.787499 |
| Nuclear repulsion energy | 162.127388 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3348 |
3163 |
0.86 |
|
|
|
| 2 |
A |
3347 |
3161 |
1.77 |
|
|
|
| 3 |
A |
3265 |
3084 |
6.81 |
|
|
|
| 4 |
A |
3243 |
3063 |
1.46 |
|
|
|
| 5 |
A |
3242 |
3062 |
4.36 |
|
|
|
| 6 |
A |
3218 |
3040 |
7.39 |
|
|
|
| 7 |
A |
1716 |
1621 |
85.38 |
|
|
|
| 8 |
A |
1698 |
1604 |
139.26 |
|
|
|
| 9 |
A |
1452 |
1371 |
2.13 |
|
|
|
| 10 |
A |
1433 |
1354 |
21.89 |
|
|
|
| 11 |
A |
1379 |
1302 |
49.62 |
|
|
|
| 12 |
A |
1338 |
1264 |
3.70 |
|
|
|
| 13 |
A |
1261 |
1191 |
232.03 |
|
|
|
| 14 |
A |
1148 |
1085 |
13.41 |
|
|
|
| 15 |
A |
1027 |
970 |
32.17 |
|
|
|
| 16 |
A |
1016 |
960 |
7.20 |
|
|
|
| 17 |
A |
1007 |
951 |
32.45 |
|
|
|
| 18 |
A |
897 |
847 |
20.46 |
|
|
|
| 19 |
A |
883 |
834 |
31.27 |
|
|
|
| 20 |
A |
858 |
810 |
45.22 |
|
|
|
| 21 |
A |
756 |
714 |
4.72 |
|
|
|
| 22 |
A |
737 |
696 |
1.39 |
|
|
|
| 23 |
A |
587 |
555 |
2.50 |
|
|
|
| 24 |
A |
455 |
430 |
2.95 |
|
|
|
| 25 |
A |
300 |
284 |
1.64 |
|
|
|
| 26 |
A |
211 |
199 |
7.98 |
|
|
|
| 27 |
A |
94 |
89 |
0.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19957.8 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 18850.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.039 |
-0.829 |
0.048 |
| C2 |
1.279 |
-0.485 |
-0.011 |
| C3 |
-0.912 |
0.188 |
0.334 |
| C4 |
1.786 |
0.742 |
-0.147 |
| C5 |
-2.122 |
0.240 |
-0.214 |
| H6 |
1.898 |
-1.371 |
0.055 |
| H7 |
-0.552 |
0.895 |
1.076 |
| H8 |
2.859 |
0.858 |
-0.169 |
| H9 |
1.170 |
1.621 |
-0.262 |
| H10 |
-2.816 |
1.006 |
0.098 |
| H11 |
-2.431 |
-0.483 |
-0.956 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3635 | 1.3699 | 2.4157 | 2.3561 | 2.0112 | 2.0710 | 3.3601 | 2.7504 | 3.3286 | 2.6173 |
C2 | 1.3635 | | 2.3172 | 1.3342 | 3.4834 | 1.0828 | 2.5367 | 2.0793 | 2.1240 | 4.3590 | 3.8286 | C3 | 1.3699 | 2.3172 | | 2.7954 | 1.3296 | 3.2248 | 1.0861 | 3.8624 | 2.5972 | 2.0857 | 2.1024 | C4 | 2.4157 | 1.3342 | 2.7954 | | 3.9406 | 2.1253 | 2.6426 | 1.0795 | 1.0796 | 4.6156 | 4.4650 | C5 | 2.3561 | 3.4834 | 1.3296 | 3.9406 | | 4.3391 | 2.1348 | 5.0193 | 3.5710 | 1.0794 | 1.0804 | H6 | 2.0112 | 1.0828 | 3.2248 | 2.1253 | 4.3391 | | 3.4892 | 2.4374 | 3.0958 | 5.2790 | 4.5334 | H7 | 2.0710 | 2.5367 | 1.0861 | 2.6426 | 2.1348 | 3.4892 | | 3.6309 | 2.2986 | 2.4683 | 3.0909 | H8 | 3.3601 | 2.0793 | 3.8624 | 1.0795 | 5.0193 | 2.4374 | 3.6309 | | 1.8553 | 5.6828 | 5.5135 | H9 | 2.7504 | 2.1240 | 2.5972 | 1.0796 | 3.5710 | 3.0958 | 2.2986 | 1.8553 | | 4.0495 | 4.2286 | H10 | 3.3286 | 4.3590 | 2.0857 | 4.6156 | 1.0794 | 5.2790 | 2.4683 | 5.6828 | 4.0495 | | 1.8638 | H11 | 2.6173 | 3.8286 | 2.1024 | 4.4650 | 1.0804 | 4.5334 | 3.0909 | 5.5135 | 4.2286 | 1.8638 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
127.139 |
|
O1 |
C2 |
H6 |
110.074 |
| O1 |
C3 |
C5 |
121.558 |
|
O1 |
C3 |
H7 |
114.476 |
| C2 |
O1 |
C3 |
115.937 |
|
C2 |
C4 |
H8 |
118.586 |
| C2 |
C4 |
H9 |
122.921 |
|
C3 |
C5 |
H10 |
119.584 |
| C3 |
C5 |
H11 |
121.117 |
|
C4 |
C2 |
H6 |
122.781 |
| C5 |
C3 |
H7 |
123.871 |
|
H8 |
C4 |
H9 |
118.475 |
| H10 |
C5 |
H11 |
119.290 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
