return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HCOOC2H5 (Ethyl formate)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS trans 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-267.769597
Energy at 298.15K-267.776849
HF Energy-266.855536
Nuclear repulsion energy178.192960
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3215 3036 9.76      
2 A' 3127 2953 9.45      
3 A' 3118 2945 7.82      
4 A' 3111 2938 53.36      
5 A' 1813 1712 244.21      
6 A' 1547 1461 8.42      
7 A' 1527 1442 2.49      
8 A' 1443 1363 14.64      
9 A' 1420 1341 1.38      
10 A' 1407 1329 2.19      
11 A' 1244 1175 382.90      
12 A' 1152 1088 8.19      
13 A' 1066 1007 16.07      
14 A' 880 831 8.13      
15 A' 799 755 3.23      
16 A' 383 362 5.92      
17 A' 233 220 6.54      
18 A" 3224 3045 14.70      
19 A" 3183 3006 7.97      
20 A" 1516 1432 5.48      
21 A" 1315 1242 1.15      
22 A" 1201 1134 4.32      
23 A" 1062 1004 0.08      
24 A" 819 773 0.88      
25 A" 358 338 18.62      
26 A" 247 233 2.03      
27 A" 71 67 0.45      

Unscaled Zero Point Vibrational Energy (zpe) 20239.3 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 19116.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.59055 0.09809 0.08685

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.166 -0.250 0.000
C2 -0.686 -0.556 0.000
O3 0.000 0.712 0.000
C4 1.334 0.608 0.000
O5 1.965 -0.419 0.000
H6 -2.737 -1.180 0.000
H7 -2.440 0.324 0.885
H8 -2.440 0.324 -0.885
H9 -0.381 -1.122 -0.881
H10 -0.381 -1.122 0.881
H11 1.768 1.616 0.000

Atom - Atom Distances (Å)
  C1 C2 O3 C4 O5 H6 H7 H8 H9 H10 H11
C11.51132.37063.60444.13521.09081.08981.08982.17392.17394.3546
C21.51131.44162.33162.65522.14402.15212.15211.09151.09153.2771
O32.37061.44161.33832.26763.32762.62392.62392.07072.07071.9856
C43.60442.33161.33831.20554.44683.88653.88652.59092.59091.0973
O54.13522.65522.26761.20554.76394.55404.55402.60312.60312.0445
H61.09082.14403.32764.44684.76391.77021.77022.51672.51675.3023
H71.08982.15212.62393.88654.55401.77021.76943.07432.51644.4894
H81.08982.15212.62393.88654.55401.77021.76942.51643.07434.4894
H92.17391.09152.07072.59092.60312.51673.07432.51641.76273.5907
H102.17391.09152.07072.59092.60312.51672.51643.07431.76273.5907
H114.35463.27711.98561.09732.04455.30234.48944.48943.59073.5907

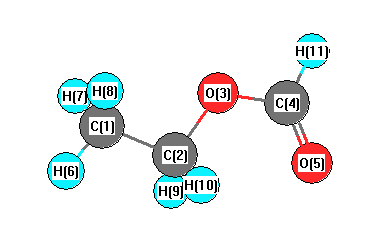
picture of Ethyl formate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 106.774 C1 C2 H9 112.273
C1 C2 H10 112.273 C2 C1 H6 109.909
C2 C1 H7 110.613 C2 C1 H8 110.613
C2 O3 C4 113.966 O3 C2 H9 108.874
O3 C2 H10 108.874 O3 C4 O5 126.034
O3 C4 H11 108.824 O5 C4 H11 125.142
H6 C1 H7 108.548 H6 C1 H8 108.548
H7 C1 H8 108.552 H9 C2 H10 107.691
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability