return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H9N (Pentanenitrile)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-250.039235
Energy at 298.15K-250.048739
HF Energy-249.056410
Nuclear repulsion energy211.136563
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3202 3024 17.38      
2 A' 3118 2945 18.52      
3 A' 3109 2936 9.49      
4 A' 3106 2934 12.04      
5 A' 3096 2925 8.99      
6 A' 2216 2093 0.06      
7 A' 1539 1453 5.72      
8 A' 1525 1440 1.51      
9 A' 1514 1430 0.84      
10 A' 1494 1411 2.73      
11 A' 1435 1355 2.69      
12 A' 1413 1334 0.36      
13 A' 1366 1290 2.32      
14 A' 1283 1212 1.38      
15 A' 1143 1080 3.76      
16 A' 1098 1037 0.28      
17 A' 1053 995 0.64      
18 A' 951 898 0.11      
19 A' 922 871 2.63      
20 A' 535 505 0.52      
21 A' 387 366 0.75      
22 A' 278 263 1.33      
23 A' 122 116 5.42      
24 A" 3199 3022 26.07      
25 A" 3180 3004 10.78      
26 A" 3162 2986 3.34      
27 A" 3144 2969 3.29      
28 A" 1531 1446 7.11      
29 A" 1348 1273 0.13      
30 A" 1330 1257 0.44      
31 A" 1255 1186 0.06      
32 A" 1148 1084 0.02      
33 A" 941 889 0.79      
34 A" 799 755 0.04      
35 A" 743 702 4.50      
36 A" 383 362 0.65      
37 A" 255 241 0.11      
38 A" 119 113 0.17      
39 A" 91 86 3.79      

Unscaled Zero Point Vibrational Energy (zpe) 29266.5 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 27642.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.49857 0.04465 0.04226

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.402 0.000
C2 1.415 0.971 0.000
C3 1.417 2.496 0.000
C4 0.007 -1.132 0.000
N5 -2.442 -2.098 0.000
C6 -1.345 -1.682 0.000
H7 2.433 2.893 0.000
H8 -0.547 0.758 0.878
H9 -0.547 0.758 -0.878
H10 1.955 0.599 0.877
H11 1.955 0.599 -0.877
H12 0.905 2.886 -0.882
H13 0.905 2.886 0.882
H14 0.537 -1.508 -0.879
H15 0.537 -1.508 0.879

Atom - Atom Distances (Å)
  C1 C2 C3 C4 N5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.52522.52871.53393.49492.48033.48181.09351.09352.15142.15142.78732.78732.17022.1702
C21.52521.52592.53064.92923.82832.17492.15972.15971.09441.09442.16982.16982.77292.7729
C32.52871.52593.89266.00015.00871.09072.76542.76542.15812.15811.09161.09164.19344.1934
C41.53392.53063.89262.63311.45974.69922.15602.15602.74952.74954.21074.21071.09321.0932
N53.49494.92926.00012.63311.17366.97673.53833.53835.23255.23256.06836.06833.16183.1618
C62.48033.82835.00871.45971.17365.93302.71312.71314.10634.10635.16815.16812.08442.0844
H73.48182.17491.09074.69926.97675.93303.76893.76892.50142.50141.76431.76434.87204.8720
H81.09352.15972.76542.15603.53832.71313.76891.75512.50633.05943.11962.57623.06522.5119
H91.09352.15972.76542.15603.53832.71313.76891.75513.05942.50632.57623.11962.51193.0652
H102.15141.09442.15812.74955.23254.10632.50142.50633.05941.75393.07012.51643.08802.5401
H112.15141.09442.15812.74955.23254.10632.50143.05942.50631.75392.51643.07012.54013.0880
H122.78732.16981.09164.21076.06835.16811.76433.11962.57623.07012.51641.76364.41004.7486
H132.78732.16981.09164.21076.06835.16811.76432.57623.11962.51643.07011.76364.74864.4100
H142.17022.77294.19341.09323.16182.08444.87203.06522.51193.08802.54014.41004.74861.7581
H152.17022.77294.19341.09323.16182.08444.87202.51193.06522.54013.08804.74864.41001.7581

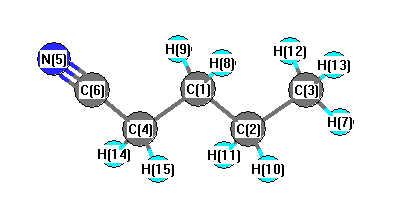
picture of Pentanenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.951 C1 C2 H10 109.321
C1 C2 H11 109.321 C1 C4 C6 111.877
C1 C4 H14 110.272 C1 C4 H15 110.272
C2 C1 C4 111.631 C2 C1 H8 110.025
C2 C1 H9 110.025 C2 C3 H7 111.358
C2 C3 H12 110.898 C2 C3 H13 110.898
C3 C2 H10 109.798 C3 C2 H11 109.798
C4 C1 H8 109.140 C4 C1 H9 109.140
C4 C6 N5 178.612 C6 C4 H14 108.616
C6 C4 H15 108.616 H7 C3 H12 107.889
H7 C3 H13 107.889 H8 C1 H9 106.750
H10 C2 H11 106.499 H12 C3 H13 107.760
H14 C4 H15 107.043
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability