return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H5N (Pyridine)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-247.656728
Energy at 298.15K-247.662845
HF Energy-246.717576
Nuclear repulsion energy206.082277
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3262 3081 2.48      
2 A1 3238 3059 3.54      
3 A1 3218 3039 3.56      
4 A1 1631 1540 15.20      
5 A1 1511 1428 4.48      
6 A1 1252 1182 1.16      
7 A1 1097 1036 3.73      
8 A1 1046 988 4.92      
9 A1 1008 952 7.50      
10 A1 598 565 4.61      
11 A2 998 943 0.00      
12 A2 914 863 0.00      
13 A2 383 362 0.00      
14 B1 1008 952 0.04      
15 B1 960 907 0.00      
16 B1 762 720 15.27      
17 B1 729 689 38.05      
18 B1 413 390 3.71      
19 B2 3255 3075 12.01      
20 B2 3217 3038 18.14      
21 B2 1621 1531 2.45      
22 B2 1475 1394 22.69      
23 B2 1409 1331 1.68      
24 B2 1385 1308 0.01      
25 B2 1179 1114 0.95      
26 B2 1083 1023 0.29      
27 B2 656 619 0.41      

Unscaled Zero Point Vibrational Energy (zpe) 19655.0 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 18564.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.20220 0.19365 0.09892

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 1.424
C2 0.000 0.000 -1.386
C3 0.000 1.141 0.720
C4 0.000 -1.141 0.720
C5 0.000 1.195 -0.672
C6 0.000 -1.195 -0.672
H7 0.000 0.000 -2.470
H8 0.000 2.056 1.303
H9 0.000 -2.056 1.303
H10 0.000 2.152 -1.179
H11 0.000 -2.152 -1.179

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11
N12.81031.34091.34092.41342.41343.89402.06012.06013.37803.3780
C22.81032.39482.39481.39191.39191.08383.38533.38532.16222.1622
C31.34092.39482.28161.39312.71923.38731.08583.25012.15153.8013
C41.34092.39482.28162.71921.39313.38733.25011.08583.80132.1515
C52.41341.39191.39312.71922.39022.15842.15513.80461.08313.3855
C62.41341.39192.71921.39312.39022.15843.80462.15513.38551.0831
H73.89401.08383.38733.38732.15842.15844.29694.29692.50962.5096
H82.06013.38531.08583.25012.15513.80464.29694.11302.48414.8863
H92.06013.38533.25011.08583.80462.15514.29694.11304.88632.4841
H103.37802.16222.15153.80131.08313.38552.50962.48414.88634.3046
H113.37802.16223.80132.15153.38551.08312.50964.88632.48414.3046

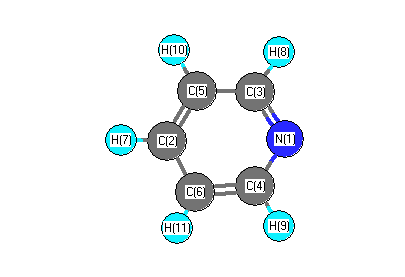
picture of Pyridine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 C5 123.937 N1 C3 H8 115.795
N1 C4 C6 123.937 N1 C4 H9 115.795
C2 C5 C3 118.607 C2 C5 H10 121.264
C2 C6 C4 118.607 C2 C6 H11 121.264
C3 N1 C4 116.593 C3 C5 H10 120.129
C4 C6 H11 120.129 C5 C2 C6 118.321
C5 C2 H7 120.840 C5 C3 H8 120.269
C6 C2 H7 120.840 C6 C4 H9 120.269
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability