Jump to
S1C2
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -251.230295 |
| Energy at 298.15K | -251.244737 |
| Nuclear repulsion energy | 260.059707 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3575 |
3377 |
0.75 |
|
|
|
| 2 |
A' |
3169 |
2993 |
42.10 |
|
|
|
| 3 |
A' |
3149 |
2974 |
22.79 |
|
|
|
| 4 |
A' |
3141 |
2967 |
19.74 |
|
|
|
| 5 |
A' |
3104 |
2931 |
11.39 |
|
|
|
| 6 |
A' |
3089 |
2918 |
21.08 |
|
|
|
| 7 |
A' |
2996 |
2830 |
97.16 |
|
|
|
| 8 |
A' |
1536 |
1451 |
3.42 |
|
|
|
| 9 |
A' |
1516 |
1432 |
4.82 |
|
|
|
| 10 |
A' |
1506 |
1423 |
6.89 |
|
|
|
| 11 |
A' |
1442 |
1362 |
3.22 |
|
|
|
| 12 |
A' |
1395 |
1317 |
0.13 |
|
|
|
| 13 |
A' |
1334 |
1260 |
0.46 |
|
|
|
| 14 |
A' |
1304 |
1232 |
3.16 |
|
|
|
| 15 |
A' |
1185 |
1119 |
4.88 |
|
|
|
| 16 |
A' |
1086 |
1026 |
4.63 |
|
|
|
| 17 |
A' |
1071 |
1011 |
8.63 |
|
|
|
| 18 |
A' |
938 |
886 |
1.73 |
|
|
|
| 19 |
A' |
889 |
840 |
16.28 |
|
|
|
| 20 |
A' |
851 |
804 |
0.97 |
|
|
|
| 21 |
A' |
800 |
756 |
70.74 |
|
|
|
| 22 |
A' |
546 |
516 |
17.49 |
|
|
|
| 23 |
A' |
430 |
406 |
5.84 |
|
|
|
| 24 |
A' |
414 |
391 |
1.34 |
|
|
|
| 25 |
A' |
253 |
239 |
1.22 |
|
|
|
| 26 |
A" |
3164 |
2989 |
10.32 |
|
|
|
| 27 |
A" |
3141 |
2966 |
50.56 |
|
|
|
| 28 |
A" |
3104 |
2932 |
21.91 |
|
|
|
| 29 |
A" |
2996 |
2829 |
22.43 |
|
|
|
| 30 |
A" |
1523 |
1438 |
0.13 |
|
|
|
| 31 |
A" |
1499 |
1416 |
0.14 |
|
|
|
| 32 |
A" |
1488 |
1406 |
4.23 |
|
|
|
| 33 |
A" |
1397 |
1319 |
0.00 |
|
|
|
| 34 |
A" |
1375 |
1299 |
17.97 |
|
|
|
| 35 |
A" |
1361 |
1285 |
12.84 |
|
|
|
| 36 |
A" |
1311 |
1239 |
2.10 |
|
|
|
| 37 |
A" |
1207 |
1140 |
9.95 |
|
|
|
| 38 |
A" |
1180 |
1114 |
2.67 |
|
|
|
| 39 |
A" |
1168 |
1103 |
6.79 |
|
|
|
| 40 |
A" |
1089 |
1029 |
2.56 |
|
|
|
| 41 |
A" |
991 |
936 |
0.78 |
|
|
|
| 42 |
A" |
909 |
859 |
0.37 |
|
|
|
| 43 |
A" |
831 |
785 |
0.19 |
|
|
|
| 44 |
A" |
444 |
419 |
0.77 |
|
|
|
| 45 |
A" |
255 |
241 |
0.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35575.4 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 33601.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.652 |
1.314 |
0.000 |
| H2 |
0.610 |
2.406 |
0.000 |
| H3 |
1.712 |
1.036 |
0.000 |
| C4 |
-0.014 |
0.748 |
1.254 |
| C5 |
-0.014 |
0.748 |
-1.254 |
| C6 |
-0.014 |
-0.776 |
-1.202 |
| C7 |
-0.014 |
-0.776 |
1.202 |
| N8 |
-0.719 |
-1.211 |
0.000 |
| H9 |
-0.784 |
-2.223 |
0.000 |
| H10 |
0.502 |
1.091 |
2.155 |
| H11 |
0.502 |
1.091 |
-2.155 |
| H12 |
-1.050 |
1.092 |
1.310 |
| H13 |
-1.050 |
1.092 |
-1.310 |
| H14 |
1.031 |
-1.133 |
-1.230 |
| H15 |
1.031 |
-1.133 |
1.230 |
| H16 |
-0.525 |
-1.189 |
-2.075 |
| H17 |
-0.525 |
-1.189 |
2.075 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0931 | 1.0962 | 1.5286 | 1.5286 | 2.5016 | 2.5016 | 2.8734 | 3.8177 | 2.1717 | 2.1717 | 2.1590 | 2.1590 | 2.7649 | 2.7649 | 3.4580 | 3.4580 |
H2 | 1.0931 | | 1.7584 | 2.1713 | 2.1713 | 3.4590 | 3.4590 | 3.8542 | 4.8351 | 2.5273 | 2.5273 | 2.4900 | 2.4900 | 3.7705 | 3.7705 | 4.3040 | 4.3040 | H3 | 1.0962 | 1.7584 | | 2.1527 | 2.1527 | 2.7761 | 2.7761 | 3.3105 | 4.1051 | 2.4720 | 2.4720 | 3.0575 | 3.0575 | 2.5844 | 2.5844 | 3.7765 | 3.7765 | C4 | 1.5286 | 2.1713 | 2.1527 | | 2.5083 | 2.8904 | 1.5248 | 2.4304 | 3.3154 | 1.0933 | 3.4650 | 1.0937 | 2.7870 | 3.2860 | 2.1515 | 3.8853 | 2.1649 | C5 | 1.5286 | 2.1713 | 2.1527 | 2.5083 | | 1.5248 | 2.8904 | 2.4304 | 3.3154 | 3.4650 | 1.0933 | 2.7870 | 1.0937 | 2.1515 | 3.2860 | 2.1649 | 3.8853 | C6 | 2.5016 | 3.4590 | 2.7761 | 2.8904 | 1.5248 | | 2.4037 | 1.4597 | 2.0325 | 3.8756 | 2.1589 | 3.2977 | 2.1396 | 1.1046 | 2.6706 | 1.0928 | 3.3420 | C7 | 2.5016 | 3.4590 | 2.7761 | 1.5248 | 2.8904 | 2.4037 | | 1.4597 | 2.0325 | 2.1589 | 3.8756 | 2.1396 | 3.2977 | 2.6706 | 1.1046 | 3.3420 | 1.0928 | N8 | 2.8734 | 3.8542 | 3.3105 | 2.4304 | 2.4304 | 1.4597 | 1.4597 | | 1.0142 | 3.3814 | 3.3814 | 2.6705 | 2.6705 | 2.1407 | 2.1407 | 2.0840 | 2.0840 | H9 | 3.8177 | 4.8351 | 4.1051 | 3.3154 | 3.3154 | 2.0325 | 2.0325 | 1.0142 | | 4.1570 | 4.1570 | 3.5749 | 3.5749 | 2.4488 | 2.4488 | 2.3326 | 2.3326 | H10 | 2.1717 | 2.5273 | 2.4720 | 1.0933 | 3.4650 | 3.8756 | 2.1589 | 3.3814 | 4.1570 | | 4.3100 | 1.7674 | 3.7967 | 4.0843 | 2.4658 | 4.9138 | 2.5021 | H11 | 2.1717 | 2.5273 | 2.4720 | 3.4650 | 1.0933 | 2.1589 | 3.8756 | 3.3814 | 4.1570 | 4.3100 | | 3.7967 | 1.7674 | 2.4658 | 4.0843 | 2.5021 | 4.9138 | H12 | 2.1590 | 2.4900 | 3.0575 | 1.0937 | 2.7870 | 3.2977 | 2.1396 | 2.6705 | 3.5749 | 1.7674 | 3.7967 | | 2.6197 | 3.9666 | 3.0480 | 4.1156 | 2.4631 | H13 | 2.1590 | 2.4900 | 3.0575 | 2.7870 | 1.0937 | 2.1396 | 3.2977 | 2.6705 | 3.5749 | 3.7967 | 1.7674 | 2.6197 | | 3.0480 | 3.9666 | 2.4631 | 4.1156 | H14 | 2.7649 | 3.7705 | 2.5844 | 3.2860 | 2.1515 | 1.1046 | 2.6706 | 2.1407 | 2.4488 | 4.0843 | 2.4658 | 3.9666 | 3.0480 | | 2.4596 | 1.7721 | 3.6533 | H15 | 2.7649 | 3.7705 | 2.5844 | 2.1515 | 3.2860 | 2.6706 | 1.1046 | 2.1407 | 2.4488 | 2.4658 | 4.0843 | 3.0480 | 3.9666 | 2.4596 | | 3.6533 | 1.7721 | H16 | 3.4580 | 4.3040 | 3.7765 | 3.8853 | 2.1649 | 1.0928 | 3.3420 | 2.0840 | 2.3326 | 4.9138 | 2.5021 | 4.1156 | 2.4631 | 1.7721 | 3.6533 | | 4.1497 | H17 | 3.4580 | 4.3040 | 3.7765 | 2.1649 | 3.8853 | 3.3420 | 1.0928 | 2.0840 | 2.3326 | 2.5021 | 4.9138 | 2.4631 | 4.1156 | 3.6533 | 1.7721 | 4.1497 | |
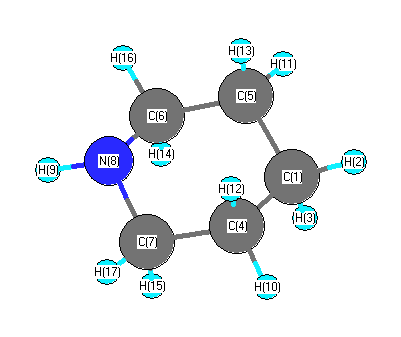 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.019 |
|
C1 |
C4 |
H10 |
110.750 |
| C1 |
C4 |
H12 |
109.724 |
|
C1 |
C5 |
C6 |
110.019 |
| C1 |
C5 |
H11 |
110.750 |
|
C1 |
C5 |
H13 |
109.724 |
| H2 |
C1 |
H3 |
106.874 |
|
H2 |
C1 |
C4 |
110.729 |
| H2 |
C1 |
C5 |
110.729 |
|
H3 |
C1 |
C4 |
109.083 |
| H3 |
C1 |
C5 |
109.083 |
|
C4 |
C1 |
C5 |
110.256 |
| C4 |
C7 |
N8 |
109.023 |
|
C4 |
C7 |
H15 |
108.758 |
| C4 |
C7 |
H17 |
110.507 |
|
C5 |
C6 |
N8 |
109.023 |
| C5 |
C6 |
H14 |
108.758 |
|
C5 |
C6 |
H16 |
110.507 |
| C6 |
C5 |
H11 |
109.996 |
|
C6 |
C5 |
H13 |
108.464 |
| C6 |
N8 |
C7 |
110.837 |
|
C6 |
N8 |
H9 |
109.156 |
| C7 |
C4 |
H10 |
109.996 |
|
C7 |
C4 |
H12 |
108.464 |
| C7 |
N8 |
H9 |
109.156 |
|
N8 |
C6 |
H14 |
112.450 |
| N8 |
C6 |
H16 |
108.607 |
|
N8 |
C7 |
H15 |
112.450 |
| N8 |
C7 |
H17 |
108.607 |
|
H10 |
C4 |
H12 |
107.830 |
| H11 |
C5 |
H13 |
107.830 |
|
H14 |
C6 |
H16 |
107.492 |
| H15 |
C7 |
H17 |
107.492 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -251.229308 |
| Energy at 298.15K | -251.243699 |
| Nuclear repulsion energy | 259.495745 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3552 |
3354 |
0.08 |
|
|
|
| 2 |
A' |
3157 |
2982 |
61.44 |
|
|
|
| 3 |
A' |
3151 |
2976 |
18.66 |
|
|
|
| 4 |
A' |
3143 |
2968 |
15.93 |
|
|
|
| 5 |
A' |
3090 |
2918 |
22.61 |
|
|
|
| 6 |
A' |
3088 |
2917 |
30.13 |
|
|
|
| 7 |
A' |
3086 |
2915 |
15.13 |
|
|
|
| 8 |
A' |
1527 |
1442 |
2.33 |
|
|
|
| 9 |
A' |
1511 |
1427 |
9.06 |
|
|
|
| 10 |
A' |
1508 |
1424 |
3.58 |
|
|
|
| 11 |
A' |
1411 |
1333 |
3.30 |
|
|
|
| 12 |
A' |
1396 |
1319 |
1.76 |
|
|
|
| 13 |
A' |
1356 |
1281 |
0.04 |
|
|
|
| 14 |
A' |
1299 |
1227 |
8.93 |
|
|
|
| 15 |
A' |
1208 |
1141 |
3.44 |
|
|
|
| 16 |
A' |
1072 |
1013 |
0.62 |
|
|
|
| 17 |
A' |
1037 |
979 |
8.55 |
|
|
|
| 18 |
A' |
946 |
894 |
10.82 |
|
|
|
| 19 |
A' |
881 |
832 |
40.39 |
|
|
|
| 20 |
A' |
842 |
795 |
0.16 |
|
|
|
| 21 |
A' |
796 |
752 |
84.52 |
|
|
|
| 22 |
A' |
554 |
523 |
1.37 |
|
|
|
| 23 |
A' |
432 |
408 |
0.95 |
|
|
|
| 24 |
A' |
402 |
379 |
4.95 |
|
|
|
| 25 |
A' |
245 |
232 |
4.44 |
|
|
|
| 26 |
A" |
3154 |
2979 |
8.22 |
|
|
|
| 27 |
A" |
3144 |
2969 |
49.62 |
|
|
|
| 28 |
A" |
3087 |
2916 |
4.00 |
|
|
|
| 29 |
A" |
3084 |
2913 |
41.80 |
|
|
|
| 30 |
A" |
1510 |
1426 |
5.24 |
|
|
|
| 31 |
A" |
1506 |
1422 |
0.36 |
|
|
|
| 32 |
A" |
1486 |
1403 |
8.04 |
|
|
|
| 33 |
A" |
1395 |
1317 |
0.57 |
|
|
|
| 34 |
A" |
1385 |
1308 |
1.99 |
|
|
|
| 35 |
A" |
1360 |
1284 |
0.93 |
|
|
|
| 36 |
A" |
1315 |
1242 |
0.65 |
|
|
|
| 37 |
A" |
1236 |
1167 |
14.34 |
|
|
|
| 38 |
A" |
1162 |
1098 |
13.21 |
|
|
|
| 39 |
A" |
1130 |
1067 |
1.93 |
|
|
|
| 40 |
A" |
1087 |
1027 |
3.32 |
|
|
|
| 41 |
A" |
959 |
906 |
4.87 |
|
|
|
| 42 |
A" |
901 |
851 |
0.15 |
|
|
|
| 43 |
A" |
819 |
773 |
0.05 |
|
|
|
| 44 |
A" |
452 |
427 |
0.87 |
|
|
|
| 45 |
A" |
244 |
230 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35551.0 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 33577.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.658 |
1.310 |
0.000 |
| H2 |
0.621 |
2.402 |
0.000 |
| H3 |
1.717 |
1.028 |
0.000 |
| C4 |
-0.010 |
0.744 |
1.254 |
| C5 |
-0.010 |
0.744 |
-1.254 |
| C6 |
-0.010 |
-0.786 |
-1.205 |
| C7 |
-0.010 |
-0.786 |
1.205 |
| N8 |
-0.623 |
-1.342 |
0.000 |
| H9 |
-1.609 |
-1.094 |
0.000 |
| H10 |
0.499 |
1.094 |
2.157 |
| H11 |
0.499 |
1.094 |
-2.157 |
| H12 |
-1.045 |
1.100 |
1.309 |
| H13 |
-1.045 |
1.100 |
-1.309 |
| H14 |
1.025 |
-1.146 |
-1.240 |
| H15 |
1.025 |
-1.146 |
1.240 |
| H16 |
-0.521 |
-1.202 |
-2.077 |
| H17 |
-0.521 |
-1.202 |
2.077 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.0930 | 1.0961 | 1.5290 | 1.5290 | 2.5081 | 2.5081 | 2.9451 | 3.3043 | 2.1741 | 2.1741 | 2.1583 | 2.1583 | 2.7761 | 2.7761 | 3.4661 | 3.4661 |
H2 | 1.0930 | | 1.7577 | 2.1725 | 2.1725 | 3.4662 | 3.4662 | 3.9457 | 4.1474 | 2.5261 | 2.5261 | 2.4874 | 2.4874 | 3.7808 | 3.7808 | 4.3140 | 4.3140 | H3 | 1.0961 | 1.7577 | | 2.1528 | 2.1528 | 2.7792 | 2.7792 | 3.3305 | 3.9453 | 2.4786 | 2.4786 | 3.0575 | 3.0575 | 2.5971 | 2.5971 | 3.7811 | 3.7811 | C4 | 1.5290 | 2.1725 | 2.1528 | | 2.5077 | 2.8963 | 1.5307 | 2.5101 | 2.7404 | 1.0942 | 3.4667 | 1.0961 | 2.7872 | 3.2963 | 2.1552 | 3.8914 | 2.1742 | C5 | 1.5290 | 2.1725 | 2.1528 | 2.5077 | | 1.5307 | 2.8963 | 2.5101 | 2.7404 | 3.4667 | 1.0942 | 2.7872 | 1.0961 | 2.1552 | 3.2963 | 2.1742 | 3.8914 | C6 | 2.5081 | 3.4662 | 2.7792 | 2.8963 | 1.5307 | | 2.4108 | 1.4623 | 2.0261 | 3.8859 | 2.1675 | 3.3093 | 2.1537 | 1.0963 | 2.6800 | 1.0930 | 3.3479 | C7 | 2.5081 | 3.4662 | 2.7792 | 1.5307 | 2.8963 | 2.4108 | | 1.4623 | 2.0261 | 2.1675 | 3.8859 | 2.1537 | 3.3093 | 2.6800 | 1.0963 | 3.3479 | 1.0930 | N8 | 2.9451 | 3.9457 | 3.3305 | 2.5101 | 2.5101 | 1.4623 | 1.4623 | | 1.0164 | 3.4419 | 3.4419 | 2.8029 | 2.8029 | 2.0717 | 2.0717 | 2.0840 | 2.0840 | H9 | 3.3043 | 4.1474 | 3.9453 | 2.7404 | 2.7404 | 2.0261 | 2.0261 | 1.0164 | | 3.7263 | 3.7263 | 2.6169 | 2.6169 | 2.9115 | 2.9115 | 2.3466 | 2.3466 | H10 | 2.1741 | 2.5261 | 2.4786 | 1.0942 | 3.4667 | 3.8859 | 2.1675 | 3.4419 | 3.7263 | | 4.3149 | 1.7613 | 3.7949 | 4.1038 | 2.4773 | 4.9236 | 2.5137 | H11 | 2.1741 | 2.5261 | 2.4786 | 3.4667 | 1.0942 | 2.1675 | 3.8859 | 3.4419 | 3.7263 | 4.3149 | | 3.7949 | 1.7613 | 2.4773 | 4.1038 | 2.5137 | 4.9236 | H12 | 2.1583 | 2.4874 | 3.0575 | 1.0961 | 2.7872 | 3.3093 | 2.1537 | 2.8029 | 2.6169 | 1.7613 | 3.7949 | | 2.6187 | 3.9789 | 3.0554 | 4.1279 | 2.4824 | H13 | 2.1583 | 2.4874 | 3.0575 | 2.7872 | 1.0961 | 2.1537 | 3.3093 | 2.8029 | 2.6169 | 3.7949 | 1.7613 | 2.6187 | | 3.0554 | 3.9789 | 2.4824 | 4.1279 | H14 | 2.7761 | 3.7808 | 2.5971 | 3.2963 | 2.1552 | 1.0963 | 2.6800 | 2.0717 | 2.9115 | 4.1038 | 2.4773 | 3.9789 | 3.0554 | | 2.4807 | 1.7591 | 3.6604 | H15 | 2.7761 | 3.7808 | 2.5971 | 2.1552 | 3.2963 | 2.6800 | 1.0963 | 2.0717 | 2.9115 | 2.4773 | 4.1038 | 3.0554 | 3.9789 | 2.4807 | | 3.6604 | 1.7591 | H16 | 3.4661 | 4.3140 | 3.7811 | 3.8914 | 2.1742 | 1.0930 | 3.3479 | 2.0840 | 2.3466 | 4.9236 | 2.5137 | 4.1279 | 2.4824 | 1.7591 | 3.6604 | | 4.1537 | H17 | 3.4661 | 4.3140 | 3.7811 | 2.1742 | 3.8914 | 3.3479 | 1.0930 | 2.0840 | 2.3466 | 2.5137 | 4.9236 | 2.4824 | 4.1279 | 3.6604 | 1.7591 | 4.1537 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.112 |
|
C1 |
C4 |
H10 |
110.858 |
| C1 |
C4 |
H12 |
109.498 |
|
C1 |
C5 |
C6 |
110.112 |
| C1 |
C5 |
H11 |
110.858 |
|
C1 |
C5 |
H13 |
109.498 |
| H2 |
C1 |
H3 |
106.826 |
|
H2 |
C1 |
C4 |
110.806 |
| H2 |
C1 |
C5 |
110.806 |
|
H3 |
C1 |
C4 |
109.068 |
| H3 |
C1 |
C5 |
109.068 |
|
C4 |
C1 |
C5 |
110.178 |
| C4 |
C7 |
N8 |
113.978 |
|
C4 |
C7 |
H15 |
109.134 |
| C4 |
C7 |
H17 |
110.820 |
|
C5 |
C6 |
N8 |
113.978 |
| C5 |
C6 |
H14 |
109.134 |
|
C5 |
C6 |
H16 |
110.820 |
| C6 |
C5 |
H11 |
110.223 |
|
C6 |
C5 |
H13 |
109.028 |
| C6 |
N8 |
C7 |
111.034 |
|
C6 |
N8 |
H9 |
108.299 |
| C7 |
C4 |
H10 |
110.223 |
|
C7 |
C4 |
H12 |
109.028 |
| C7 |
N8 |
H9 |
108.299 |
|
N8 |
C6 |
H14 |
107.266 |
| N8 |
C6 |
H16 |
108.419 |
|
N8 |
C7 |
H15 |
107.266 |
| N8 |
C7 |
H17 |
108.419 |
|
H10 |
C4 |
H12 |
107.053 |
| H11 |
C5 |
H13 |
107.053 |
|
H14 |
C6 |
H16 |
106.934 |
| H15 |
C7 |
H17 |
106.934 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
