return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6OS (Vinyl sulfoxide)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-628.272127
Energy at 298.15K-628.278411
Nuclear repulsion energy291.958855
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3318 3134 0.90      
2 A' 3247 3067 1.80      
3 A' 3207 3029 2.60      
4 A' 1652 1560 2.62      
5 A' 1414 1336 13.33      
6 A' 1278 1207 0.36      
7 A' 1147 1083 79.28      
8 A' 1014 958 32.89      
9 A' 1002 947 2.45      
10 A' 962 909 19.45      
11 A' 719 679 55.97      
12 A' 656 619 4.13      
13 A' 506 478 0.78      
14 A' 310 293 1.53      
15 A' 217 205 2.13      
16 A' 93 88 0.33      
17 A" 3318 3134 0.00      
18 A" 3243 3063 2.23      
19 A" 3207 3029 1.16      
20 A" 1644 1552 2.80      
21 A" 1409 1331 5.41      
22 A" 1261 1191 6.89      
23 A" 1010 954 16.94      
24 A" 981 926 18.72      
25 A" 960 907 11.07      
26 A" 670 633 11.85      
27 A" 573 541 3.96      
28 A" 483 456 8.40      
29 A" 265 250 5.76      
30 A" 170 160 0.78      

Unscaled Zero Point Vibrational Energy (zpe) 19968.2 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 18859.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.16493 0.07697 0.06824

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.598 -0.513 0.000
O2 1.279 0.812 0.000
C3 -0.618 -0.492 1.308
C4 -0.618 -0.492 -1.308
C5 -0.618 0.539 2.151
C6 -0.618 0.539 -2.151
H7 -1.263 -1.361 1.382
H8 -1.263 -1.361 -1.382
H9 -1.309 0.590 2.982
H10 -1.309 0.590 -2.982
H11 0.085 1.349 1.997
H12 0.085 1.349 -1.997

Atom - Atom Distances (Å)
  S1 O2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
S11.48941.78591.78592.68552.68552.46792.46793.70773.70772.77832.7783
O21.48942.64762.64762.88122.88123.61823.61823.95513.95512.38812.3881
C31.78592.64762.61571.33203.60941.08452.89882.11014.47802.08813.8478
C41.78592.64762.61573.60941.33202.89881.08454.47802.11013.84782.0881
C52.68552.88121.33203.60944.30232.14894.06261.08215.17981.08394.2844
C62.68552.88123.60941.33204.30234.06262.14895.17981.08214.28441.0839
H72.46793.61821.08452.89882.14894.06262.76302.52414.78023.08894.5360
H82.46793.61822.89881.08454.06262.14892.76304.78022.52414.53603.0889
H93.70773.95512.11014.47801.08215.17982.52414.78025.96411.86845.2257
H103.70773.95514.47802.11015.17981.08214.78022.52415.96415.22571.8684
H112.77832.38812.08813.84781.08394.28443.08894.53601.86845.22573.9934
H122.77832.38813.84782.08814.28441.08394.53603.08895.22571.86843.9934

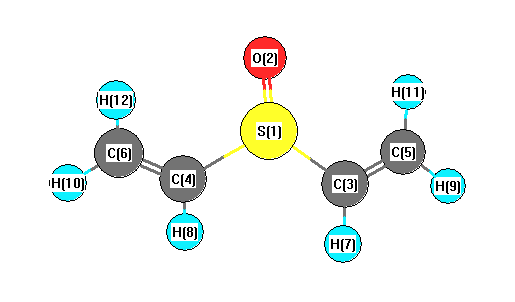
picture of Vinyl sulfoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 118.202 S1 C3 H7 116.445
S1 C4 C6 118.202 S1 C4 H8 116.445
O2 S1 C3 107.528 O2 S1 C4 107.528
C3 S1 C4 94.161 C3 C5 H9 121.524
C3 C5 H11 119.258 C4 C6 H10 121.524
C4 C6 H12 119.258 C5 C3 H7 125.248
C6 C4 H8 125.248 H9 C5 H11 119.215
H10 C6 H12 119.215
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability