return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH5N3 (Guanidine)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-204.921497
Energy at 298.15K-204.929218
HF Energy-204.152269
Nuclear repulsion energy124.047213
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3717 3521 16.57      
2 A 3711 3516 22.53      
3 A 3612 3422 12.55      
4 A 3606 3416 21.47      
5 A 3558 3370 2.18      
6 A 1795 1700 245.45      
7 A 1676 1588 119.97      
8 A 1658 1571 31.63      
9 A 1491 1413 106.70      
10 A 1223 1158 22.63      
11 A 1161 1100 68.64      
12 A 1132 1072 12.62      
13 A 955 904 3.92      
14 A 860 815 54.51      
15 A 815 772 235.91      
16 A 707 670 169.44      
17 A 609 577 183.19      
18 A 543 515 14.54      
19 A 479 454 0.08      
20 A 405 384 16.25      
21 A 362 343 43.34      

Unscaled Zero Point Vibrational Energy (zpe) 17036.5 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 16140.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
0.35058 0.34150 0.17565

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.021 0.125 0.000
N2 -0.230 1.383 0.008
N3 -0.965 -0.904 0.087
N4 1.279 -0.375 -0.092
H5 -1.227 1.580 -0.030
H6 -1.908 -0.593 -0.086
H7 -0.733 -1.697 -0.493
H8 1.949 0.368 0.040
H9 1.450 -1.140 0.545

Atom - Atom Distances (Å)
  C1 N2 N3 N4 H5 H6 H7 H8 H9
C11.27581.39891.39571.88992.02112.01731.98472.0147
N21.27582.40342.31971.01652.59433.16102.40383.0785
N31.39892.40342.31202.49981.00811.00973.17932.4690
N41.39572.31972.31203.17893.19442.44031.00951.0099
H51.88991.01652.49983.17892.27763.34573.39943.8587
H62.02112.59431.00813.19442.27761.66273.97673.4602
H72.01733.16101.00972.44033.34571.66273.42642.4806
H81.98472.40383.17931.00953.39943.97673.42641.6668
H92.01473.07852.46901.00993.85873.46022.48061.6668

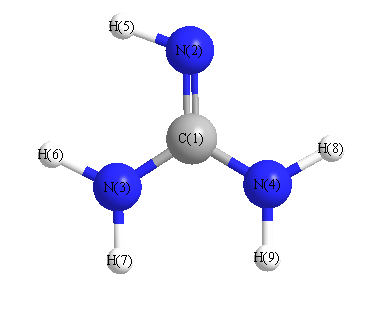
picture of Guanidine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H5 110.560 C1 N3 H6 113.216
C1 N3 H7 112.768 C1 N4 H8 110.175
C1 N4 H9 112.767 N2 C1 N3 127.888
N2 C1 N4 120.458 N3 C1 N4 111.652
H6 N3 H7 110.970 H8 N4 H9 111.256
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability