return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH5N3 (Guanidine)

using model chemistry: CCD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at CCD/6-31G(2df,p)
 hartrees
Energy at 0K-204.915304
Energy at 298.15K-204.923041
HF Energy-204.152612
Nuclear repulsion energy124.205577
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3731 3534 18.26      
2 A 3725 3529 24.20      
3 A 3625 3434 14.51      
4 A 3619 3428 23.08      
5 A 3575 3386 2.55      
6 A 1822 1725 260.47      
7 A 1681 1592 120.06      
8 A 1665 1577 33.37      
9 A 1497 1418 107.07      
10 A 1229 1165 20.56      
11 A 1163 1102 67.77      
12 A 1136 1076 12.34      
13 A 961 910 5.16      
14 A 870 824 59.01      
15 A 819 776 225.58      
16 A 708 670 175.02      
17 A 612 579 189.41      
18 A 548 519 14.64      
19 A 482 457 0.10      
20 A 405 384 17.73      
21 A 363 344 44.38      

Unscaled Zero Point Vibrational Energy (zpe) 17117.5 cm-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 16213.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G(2df,p)
ABC
0.35181 0.34212 0.17611

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.021 0.126 0.000
N2 -0.234 1.379 0.008
N3 -0.961 -0.904 0.087
N4 1.279 -0.371 -0.092
H5 -1.231 1.574 -0.027
H6 -1.905 -0.598 -0.086
H7 -0.727 -1.697 -0.491
H8 1.948 0.373 0.040
H9 1.452 -1.136 0.544

Atom - Atom Distances (Å)
  C1 N2 N3 N4 H5 H6 H7 H8 H9
C11.27181.39781.39481.88712.02082.01621.98462.0140
N21.27182.39812.31631.01572.59063.15572.40363.0753
N31.39782.39812.31032.49531.00751.00903.17792.4675
N41.39482.31632.31033.17593.19292.43841.00891.0092
H51.88711.01572.49533.17592.27483.34193.39863.8555
H62.02082.59061.00753.19292.27481.66173.97593.4584
H72.01623.15571.00902.43843.34191.66173.42432.4775
H81.98462.40363.17791.00893.39863.97593.42431.6659
H92.01403.07532.46751.00923.85553.45842.47751.6659

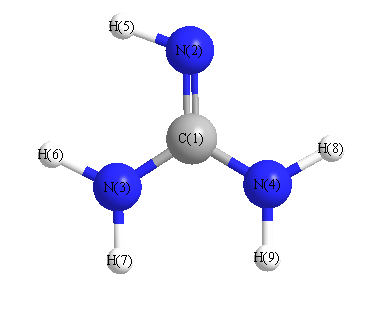
picture of Guanidine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H5 110.683 C1 N3 H6 113.316
C1 N3 H7 112.801 C1 N4 H8 110.280
C1 N4 H9 112.831 N2 C1 N3 127.812
N2 C1 N4 120.535 N3 C1 N4 111.651
H6 N3 H7 110.989 H8 N4 H9 111.280
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability