return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2C(CH3)CH3 (1-Propene, 2-methyl-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-156.774818
Energy at 298.15K-156.782723
HF Energy-156.131530
Nuclear repulsion energy119.818264
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3207 3029 5.02      
2 A1 3202 3024 12.53      
3 A1 3091 2920 18.41      
4 A1 1730 1634 7.58      
5 A1 1528 1443 10.72      
6 A1 1466 1385 0.10      
7 A1 1427 1348 0.37      
8 A1 1095 1034 4.78      
9 A1 834 788 0.13      
10 A1 372 352 0.26      
11 A2 3164 2988 0.00      
12 A2 1499 1416 0.00      
13 A2 1034 977 0.00      
14 A2 719 679 0.00      
15 A2 177 167 0.00      
16 B1 3166 2990 25.56      
17 B1 1518 1434 14.74      
18 B1 1112 1050 0.18      
19 B1 930 879 37.38      
20 B1 433 409 5.05      
21 B1 216 204 0.52      
22 B2 3302 3119 9.46      
23 B2 3201 3023 12.65      
24 B2 3088 2917 14.52      
25 B2 1514 1430 3.83      
26 B2 1429 1350 9.04      
27 B2 1319 1246 2.71      
28 B2 1003 947 0.03      
29 B2 974 920 0.18      
30 B2 430 406 1.43      

Unscaled Zero Point Vibrational Energy (zpe) 24089.4 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 22752.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.30558 0.28100 0.15472

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.111
C2 0.000 0.000 1.450
H3 0.000 0.922 2.017
H4 0.000 -0.922 2.017
C5 0.000 1.287 -0.674
C6 0.000 -1.287 -0.674
H7 0.000 2.149 -0.009
H8 0.878 1.345 -1.322
H9 -0.878 1.345 -1.322
H10 0.000 -2.149 -0.009
H11 -0.878 -1.345 -1.322
H12 0.878 -1.345 -1.322

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 H7 H8 H9 H10 H11 H12
C11.33922.11822.11821.50711.50712.15232.15272.15272.15232.15272.1527
C21.33921.08311.08312.48342.48342.59763.20403.20402.59763.20403.2040
H32.11821.08311.84502.71613.48212.36893.47903.47903.67994.13134.1313
H42.11821.08311.84503.48212.71613.67994.13134.13132.36893.47903.4790
C51.50712.48342.71613.48212.57321.08901.09271.09273.49932.84922.8492
C61.50712.48343.48212.71612.57323.49932.84922.84921.08901.09271.0927
H72.15232.59762.36893.67991.08903.49931.77211.77214.29803.83473.8347
H82.15273.20403.47904.13131.09272.84921.77211.75543.83473.21282.6908
H92.15273.20403.47904.13131.09272.84921.77211.75543.83472.69083.2128
H102.15232.59763.67992.36893.49931.08904.29803.83473.83471.77211.7721
H112.15273.20404.13133.47902.84921.09273.83473.21282.69081.77211.7554
H122.15273.20404.13133.47902.84921.09273.83472.69083.21281.77211.7554

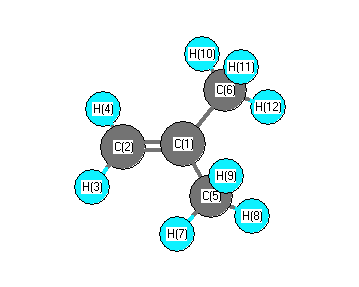
picture of 1-Propene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H3 121.600 C1 C2 H4 121.600
C1 C5 H7 110.981 C1 C5 H8 110.784
C1 C5 H9 110.784 C1 C6 H10 110.981
C1 C6 H11 110.784 C1 C6 H12 110.784
C2 C1 C5 121.385 C2 C1 C6 121.385
H3 C2 H4 116.799 C5 C1 C6 117.229
H7 C5 H8 108.638 H7 C5 H9 108.638
H8 C5 H9 106.888 H10 C6 H11 108.638
H10 C6 H12 108.638 H11 C6 H12 106.888
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability