return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10S2 (1,4-Butanedithiol)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-953.337149
Energy at 298.15K-953.347886
HF Energy-952.351327
Nuclear repulsion energy336.525253
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3124 2951 0.00      
2 Ag 3096 2924 0.00      
3 Ag 2786 2631 0.00      
4 Ag 1514 1430 0.00      
5 Ag 1512 1428 0.00      
6 Ag 1404 1326 0.00      
7 Ag 1289 1217 0.00      
8 Ag 1128 1065 0.00      
9 Ag 1086 1026 0.00      
10 Ag 868 820 0.00      
11 Ag 800 756 0.00      
12 Ag 339 320 0.00      
13 Ag 208 196 0.00      
14 Au 3196 3018 17.93      
15 Au 3164 2988 7.03      
16 Au 1334 1260 1.32      
17 Au 1119 1057 3.26      
18 Au 905 855 1.67      
19 Au 752 710 3.86      
20 Au 202 191 33.27      
21 Au 103 97 11.19      
22 Au 56 53 6.56      
23 Bg 3193 3015 0.00      
24 Bg 3147 2972 0.00      
25 Bg 1345 1271 0.00      
26 Bg 1271 1200 0.00      
27 Bg 1060 1001 0.00      
28 Bg 789 745 0.00      
29 Bg 197 186 0.00      
30 Bg 139 131 0.00      
31 Bu 3126 2952 35.90      
32 Bu 3104 2931 8.39      
33 Bu 2786 2631 2.80      
34 Bu 1531 1446 4.40      
35 Bu 1508 1424 4.13      
36 Bu 1356 1281 35.18      
37 Bu 1238 1169 23.25      
38 Bu 1077 1018 0.17      
39 Bu 903 852 2.32      
40 Bu 762 720 2.81      
41 Bu 393 371 4.22      
42 Bu 100 94 3.49      

Unscaled Zero Point Vibrational Energy (zpe) 29502.4 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 27865.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.47732 0.01898 0.01851

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.334 3.139 0.000
S2 -1.334 -3.139 0.000
C3 1.420 1.319 0.000
C4 -1.420 -1.319 0.000
C5 0.000 0.764 0.000
C6 0.000 -0.764 0.000
H7 2.651 3.366 0.000
H8 -2.651 -3.366 0.000
H9 -0.535 1.135 0.880
H10 -0.535 1.135 -0.880
H11 0.535 -1.135 0.880
H12 0.535 -1.135 -0.880
H13 -1.957 -0.981 -0.887
H14 -1.957 -0.981 0.887
H15 1.957 0.981 -0.887
H16 1.957 0.981 0.887

Atom - Atom Distances (Å)
  S1 S2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
S16.82201.82205.24062.72444.12491.33677.62902.87812.87814.43644.43645.34735.34732.41562.4156
S26.82205.24061.82204.12492.72447.62901.33674.43644.43642.87812.87812.41562.41565.34735.3473
C31.82205.24063.87641.52462.52112.38856.20702.15142.15142.75332.75334.18114.18111.09101.0910
C45.24061.82203.87642.52111.52466.20702.38852.75332.75332.15142.15141.09101.09104.18114.1811
C52.72444.12491.52462.52111.52783.71494.90771.09431.09432.15992.15992.76792.76792.15972.1597
C64.12492.72442.52111.52461.52784.90773.71492.15992.15991.09431.09432.15972.15972.76792.7679
H71.33677.62902.38856.20703.71494.90778.56953.98783.98785.05095.05096.39686.39682.63782.6378
H87.62901.33676.20702.38854.90773.71498.56955.05095.05093.98783.98782.63782.63786.39686.3968
H92.87814.43642.15142.75331.09432.15993.98785.05091.75942.50913.06453.10172.54933.05862.4967
H102.87814.43642.15142.75331.09432.15993.98785.05091.75943.06452.50912.54933.10172.49673.0586
H114.43642.87812.75332.15142.15991.09435.05093.98782.50913.06451.75943.05862.49673.10172.5493
H124.43642.87812.75332.15142.15991.09435.05093.98783.06452.50911.75942.49673.05862.54933.1017
H135.34732.41564.18111.09102.76792.15976.39682.63783.10172.54933.05862.49671.77414.37834.7241
H145.34732.41564.18111.09102.76792.15976.39682.63782.54933.10172.49673.05861.77414.72414.3783
H152.41565.34731.09104.18112.15972.76792.63786.39683.05862.49673.10172.54934.37834.72411.7741
H162.41565.34731.09104.18112.15972.76792.63786.39682.49673.05862.54933.10174.72414.37831.7741

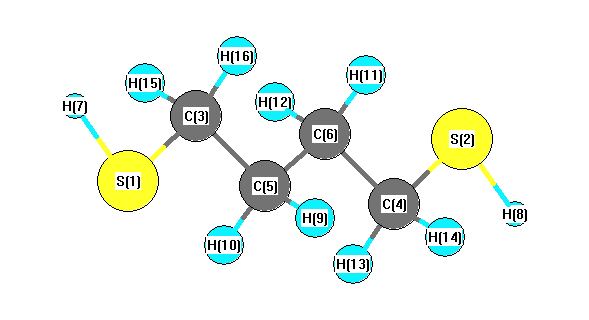
picture of 1,4-Butanedithiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C3 C5 108.663 S1 C3 H15 109.470
S1 C3 H16 109.470 S2 C4 C6 108.663
S2 C4 H13 109.470 S2 C4 H14 109.470
C3 S1 H7 97.058 C3 C5 C6 111.366
C3 C5 H9 109.363 C3 C5 H10 109.363
C4 S2 H8 97.058 C4 C6 C5 111.366
C4 C6 H11 109.363 C4 C6 H12 109.363
C5 C3 H15 110.217 C5 C3 H16 110.217
C5 C6 H11 109.818 C5 C6 H12 109.818
C6 C4 H13 110.217 C6 C4 H14 110.217
C6 C5 H9 109.818 C6 C5 H10 109.818
H9 C5 H10 107.011 H11 C6 H12 107.011
H13 C4 H14 108.791 H15 C3 H16 108.791
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability