return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H15N (triethylamine)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-291.692798
Energy at 298.15K-291.710051
HF Energy-290.409033
Nuclear repulsion energy336.902062
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3174 3007 36.72      
2 A 3160 2994 5.46      
3 A 3132 2967 0.58      
4 A 3086 2924 5.43      
5 A 2997 2840 142.26      
6 A 1550 1469 8.35      
7 A 1525 1445 0.39      
8 A 1519 1439 1.21      
9 A 1452 1376 14.59      
10 A 1432 1357 0.05      
11 A 1342 1271 3.63      
12 A 1198 1135 9.68      
13 A 1111 1052 2.54      
14 A 1056 1000 5.89      
15 A 825 782 0.07      
16 A 769 729 8.46      
17 A 448 424 3.27      
18 A 311 295 0.79      
19 A 191 181 0.54      
20 A 95 90 0.00      
21 E 3176 3009 29.24      
22 E 3163 2997 38.50      
23 E 3130 2965 20.14      
24 E 3086 2924 20.74      
25 E 2989 2832 10.42      
26 E 1546 1464 3.37      
26 E 1546 1464 3.37      
27 E 1532 1452 0.74      
27 E 1532 1452 0.74      
28 E 1519 1439 2.00      
28 E 1519 1439 2.00      
29 E 1450 1374 20.40      
30 E 1420 1345 5.59      
30 E 1420 1345 5.59      
31 E 1354 1283 20.81      
32 E 1267 1201 17.58      
33 E 1128 1068 16.73      
34 E 1116 1058 6.95      
34 E 1116 1058 6.95      
35 E 948 898 0.98      
35 E 948 898 0.98      
36 E 812 769 2.55      
36 E 812 769 2.55      
37 E 479 453 2.12      
37 E 479 453 2.12      
38 E 319 302 0.47      
38 E 319 302 0.47      
39 E 217 206 0.02      
39 E 217 206 0.02      
40 E 92 87 0.19      
40 E 92 87 0.19      

Unscaled Zero Point Vibrational Energy (zpe) 45928.4 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 43512.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
0.07732 0.07732 0.04446

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.002
C2 0.332 1.353 -0.448
C3 1.005 -0.964 -0.448
C4 -1.337 -0.389 -0.448
C5 -0.297 2.428 0.435
C6 2.251 -0.957 0.435
C7 -1.954 -1.471 0.435
H8 1.418 1.468 -0.408
H9 0.044 1.504 -1.503
H10 0.562 -1.962 -0.408
H11 1.280 -0.790 -1.503
H12 -1.980 0.494 -0.408
H13 -1.324 -0.714 -1.503
H14 -1.389 2.387 0.420
H15 0.030 2.299 1.469
H16 0.000 3.424 0.094
H17 2.761 0.009 0.420
H18 1.975 -1.176 1.469
H19 2.965 -1.712 0.094
H20 -1.373 -2.396 0.420
H21 -2.006 -1.123 1.469
H22 -2.965 -1.712 0.094

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.46371.46371.46372.48402.48402.48402.08142.12812.08142.12812.08142.12812.79262.72663.42482.79262.72663.42482.79262.72663.4248
C21.46372.41242.41241.52703.12973.73901.09241.10423.32252.56942.46682.85082.18722.15852.16642.90843.57294.07644.20843.90744.5337
C31.46372.41242.41243.73901.52703.12982.46682.85081.09241.10423.32252.56944.20843.90744.53372.18722.15852.16642.90843.57294.0764
C41.46372.41242.41243.12983.73901.52703.32252.56942.46682.85081.09241.10422.90853.57294.07644.20843.90744.53372.18722.15852.1664
C52.48401.52703.73903.12984.23674.23672.13832.17404.55164.07402.69883.83181.09231.09201.09353.89914.38395.28154.94224.07404.9366
C62.48403.12971.52703.73904.23674.23672.69883.83182.13832.17404.55164.07404.94224.07404.93661.09231.09201.09353.89914.38395.2815
C72.48403.73903.12981.52704.23674.23674.55164.07402.69883.83182.13832.17403.89914.38395.28154.94224.07404.93661.09231.09201.0935
H82.08141.09242.46683.32252.13832.69884.55161.75753.53452.51293.53453.67113.06672.47702.46712.14913.28943.57164.83734.68545.4377
H92.12811.10422.85082.56942.17403.83184.07401.75753.67112.60552.51292.60552.55553.07622.49793.64944.44314.62874.57294.46444.6843
H102.08143.32251.09242.46684.55162.13832.69883.53453.67111.75753.53452.51294.83734.68545.43773.06672.47702.46712.14913.28943.5716
H112.12812.56941.10422.85084.07402.17403.83182.51292.60551.75753.67112.60554.57294.46444.68432.55553.07622.49793.64944.44314.6287
H122.08142.46683.32251.09242.69884.55162.13833.53452.51293.53453.67111.75752.14913.28943.57164.83734.68545.43773.06672.47702.4671
H132.12812.85082.56941.10423.83184.07402.17403.67112.60552.51292.60551.75753.64944.44314.62874.57294.46444.68432.55553.07622.4979
H142.79262.18724.20842.90851.09234.94223.89913.06672.55554.83734.57292.14913.64941.76631.76354.78245.01045.98804.78243.71444.4031
H152.72662.15853.90743.57291.09204.07404.38392.47703.07624.68544.46443.28944.44311.76631.77633.71443.98165.15595.01043.98165.1907
H163.42482.16644.53374.07641.09354.93665.28152.46712.49795.43774.68433.57164.62871.76351.77634.40315.19075.92995.98805.15595.9299
H172.79262.90842.18724.20843.89911.09234.94222.14913.64943.06672.55554.83734.57294.78243.71444.40311.76631.76354.78245.01045.9880
H182.72663.57292.15853.90744.38391.09204.07403.28944.44312.47703.07624.68544.46445.01043.98165.19071.76631.77633.71443.98165.1559
H193.42484.07642.16644.53375.28151.09354.93663.57164.62872.46712.49795.43774.68435.98805.15595.92991.76351.77634.40315.19075.9299
H202.79264.20842.90842.18724.94223.89911.09234.83734.57292.14913.64943.06672.55554.78245.01045.98804.78243.71444.40311.76631.7635
H212.72663.90743.57292.15854.07404.38391.09204.68544.46443.28944.44312.47703.07623.71443.98165.15595.01043.98165.19071.76631.7763
H223.42484.53374.07642.16644.93665.28151.09355.43774.68433.57164.62872.46712.49794.40315.19075.92995.98805.15595.92991.76351.7763

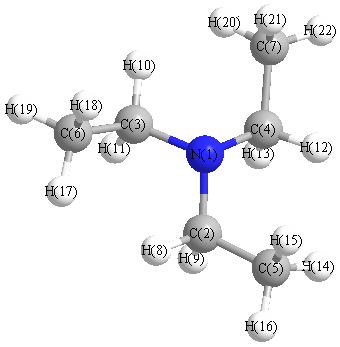
picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 112.298 N1 C2 H8 108.156
N1 C2 H9 111.165 N1 C3 C6 112.298
N1 C3 H10 108.156 N1 C3 H11 111.165
N1 C4 C7 112.298 N1 C4 H12 108.156
N1 C4 H13 111.165 C2 N1 C3 110.984
C2 N1 C4 110.984 C2 C5 H14 112.172
C2 C5 H15 109.897 C2 C5 H16 110.432
C3 N1 C4 110.984 C3 C6 H17 112.172
C3 C6 H18 109.897 C3 C6 H19 110.432
C4 C7 H20 112.172 C4 C7 H21 109.897
C4 C7 H22 110.432 C5 C2 H8 108.298
C5 C2 H9 110.398 C6 C3 H10 108.298
C6 C3 H11 110.398 C7 C4 H12 108.298
C7 C4 H13 110.399 H8 C2 H9 106.274
H10 C3 H11 106.274 H12 C4 H13 106.274
H14 C5 H15 107.926 H14 C5 H16 107.570
H15 C5 H16 108.736 H17 C6 H18 107.926
H17 C6 H19 107.570 H18 C6 H19 108.736
H20 C7 H21 107.926 H20 C7 H22 107.570
H21 C7 H22 108.737
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability