return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H15N (triethylamine)

using model chemistry: CCD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C3 1A
Energy calculated at CCD/6-31G(2df,p)
 hartrees
Energy at 0K-291.684847
Energy at 298.15K-291.702120
HF Energy-290.409188
Nuclear repulsion energy337.041858
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3190 3022 33.02      
2 A 3177 3009 4.56      
3 A 3146 2980 0.70      
4 A 3099 2935 5.29      
5 A 3013 2854 133.60      
6 A 1554 1472 8.97      
7 A 1529 1448 0.39      
8 A 1522 1442 1.24      
9 A 1457 1380 14.23      
10 A 1436 1360 0.16      
11 A 1346 1275 3.69      
12 A 1201 1138 10.07      
13 A 1114 1055 2.65      
14 A 1060 1004 5.92      
15 A 828 784 0.07      
16 A 772 732 8.29      
17 A 449 425 3.33      
18 A 312 295 0.79      
19 A 191 181 0.59      
20 A 95 90 0.00      
21 E 3192 3023 25.33      
22 E 3180 3012 36.27      
23 E 3145 2979 19.54      
24 E 3099 2935 18.70      
25 E 3006 2847 9.70      
25 E 3006 2847 9.70      
26 E 1550 1468 3.66      
26 E 1550 1468 3.66      
27 E 1536 1455 0.71      
27 E 1536 1455 0.71      
28 E 1523 1442 2.14      
28 E 1523 1442 2.14      
29 E 1454 1377 21.28      
30 E 1425 1349 5.01      
30 E 1425 1349 5.01      
31 E 1359 1288 20.03      
32 E 1272 1205 18.03      
33 E 1132 1072 17.26      
34 E 1120 1061 5.72      
34 E 1120 1061 5.72      
35 E 951 901 0.89      
35 E 951 901 0.89      
36 E 814 771 2.64      
36 E 814 771 2.64      
37 E 480 455 2.16      
37 E 480 455 2.16      
38 E 320 303 0.48      
38 E 320 303 0.48      
39 E 218 206 0.02      
39 E 218 206 0.02      
40 E 92 87 0.19      
40 E 92 87 0.19      

Unscaled Zero Point Vibrational Energy (zpe) 46110.0 cm-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 43675.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G(2df,p)
ABC
0.07737 0.07737 0.04449

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G(2df,p)

Point Group is C3

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 0.002
C2 0.332 1.352 -0.448
C3 1.005 -0.963 -0.448
C4 -1.337 -0.389 -0.448
C5 -0.297 2.427 0.435
C6 2.251 -0.956 0.435
C7 -1.954 -1.471 0.435
H8 1.416 1.468 -0.408
H9 0.044 1.503 -1.502
H10 0.563 -1.961 -0.408
H11 1.280 -0.789 -1.502
H12 -1.979 0.493 -0.408
H13 -1.324 -0.714 -1.502
H14 -1.388 2.386 0.421
H15 0.030 2.298 1.468
H16 0.000 3.422 0.094
H17 2.760 0.009 0.421
H18 1.975 -1.175 1.468
H19 2.964 -1.711 0.094
H20 -1.372 -2.395 0.421
H21 -2.006 -1.123 1.468
H22 -2.964 -1.711 0.094

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21 H22
N11.46291.46291.46292.48362.48362.48362.08042.12652.08042.12652.08042.12652.79192.72633.42342.79192.72633.42342.79192.72633.4234
C21.46292.41132.41131.52673.12893.73781.09161.10333.32092.56842.46592.84952.18632.15762.16542.90793.57184.07474.20693.90604.5316
C31.46292.41132.41133.73781.52673.12892.46592.84951.09161.10333.32092.56844.20693.90604.53162.18632.15762.16542.90793.57184.0747
C41.46292.41132.41133.12893.73781.52673.32092.56842.46592.84951.09161.10332.90793.57184.07474.20693.90604.53162.18632.15762.1654
C52.48361.52673.73783.12894.23574.23572.13712.17294.55014.07272.69843.83041.09151.09121.09273.89844.38295.27974.94074.07304.9348
C62.48363.12891.52673.73784.23574.23572.69843.83042.13712.17294.55014.07274.94074.07304.93481.09151.09121.09273.89844.38295.2797
C72.48363.73783.12891.52674.23574.23574.55014.07272.69843.83042.13712.17293.89844.38295.27974.94074.07304.93481.09151.09121.0927
H82.08041.09162.46593.32092.13712.69844.55011.75603.53302.51223.53303.66953.06482.47552.46532.14943.28863.57074.83574.68385.4352
H92.12651.10332.84952.56842.17293.83044.07271.75603.66952.60472.51222.60472.55443.07432.49643.64824.44134.62664.57134.46264.6822
H102.08043.32091.09162.46594.55012.13712.69843.53303.66951.75603.53302.51224.83574.68385.43523.06482.47552.46532.14943.28863.5707
H112.12652.56841.10332.84954.07272.17293.83042.51222.60471.75603.66952.60474.57134.46264.68222.55443.07432.49643.64824.44134.6266
H122.08042.46593.32091.09162.69844.55012.13713.53302.51223.53303.66951.75602.14943.28863.57074.83574.68385.43523.06482.47552.4653
H132.12652.84952.56841.10333.83044.07272.17293.66952.60472.51222.60471.75603.64824.44134.62664.57134.46264.68222.55443.07432.4964
H142.79192.18634.20692.90791.09154.94073.89843.06482.55444.83574.57132.14943.64821.76511.76244.78105.00905.98564.78103.71364.4018
H152.72632.15763.90603.57181.09124.07304.38292.47553.07434.68384.46263.28864.44131.76511.77503.71363.98105.15425.00903.98105.1889
H163.42342.16544.53164.07471.09274.93485.27972.46532.49645.43524.68223.57074.62661.76241.77504.40185.18895.92735.98565.15425.9273
H172.79192.90792.18634.20693.89841.09154.94072.14943.64823.06482.55444.83574.57134.78103.71364.40181.76511.76244.78105.00905.9856
H182.72633.57182.15763.90604.38291.09124.07303.28864.44132.47553.07434.68384.46265.00903.98105.18891.76511.77503.71363.98105.1542
H193.42344.07472.16544.53165.27971.09274.93483.57074.62662.46532.49645.43524.68225.98565.15425.92731.76241.77504.40185.18895.9273
H202.79194.20692.90792.18634.94073.89841.09154.83574.57132.14943.64823.06482.55444.78105.00905.98564.78103.71364.40181.76511.7624
H212.72633.90603.57182.15764.07304.38291.09124.68384.46263.28864.44132.47553.07433.71363.98105.15425.00903.98105.18891.76511.7750
H223.42344.53164.07472.16544.93485.27971.09275.43524.68223.57074.62662.46532.49644.40185.18895.92735.98565.15425.92731.76241.7750

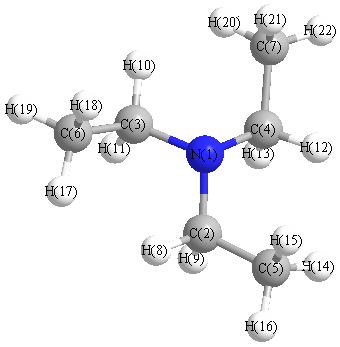
picture of triethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C5 112.334 N1 C2 H8 108.177
N1 C2 H9 111.155 N1 C3 C6 112.334
N1 C3 H10 108.177 N1 C3 H11 111.155
N1 C4 C7 112.334 N1 C4 H12 108.177
N1 C4 H13 111.155 C2 N1 C3 111.009
C2 N1 C4 111.009 C2 C5 H14 112.171
C2 C5 H15 109.893 C2 C5 H16 110.422
C3 N1 C4 111.009 C3 C6 H17 112.171
C3 C6 H18 109.893 C3 C6 H19 110.422
C4 C7 H20 112.171 C4 C7 H21 109.893
C4 C7 H22 110.422 C5 C2 H8 108.266
C5 C2 H9 110.389 C6 C3 H10 108.266
C6 C3 H11 110.389 C7 C4 H12 108.266
C7 C4 H13 110.389 H8 C2 H9 106.266
H10 C3 H11 106.266 H12 C4 H13 106.266
H14 C5 H15 107.929 H14 C5 H16 107.583
H15 C5 H16 108.737 H17 C6 H18 107.929
H17 C6 H19 107.583 H18 C6 H19 108.737
H20 C7 H21 107.929 H20 C7 H22 107.583
H21 C7 H22 108.737
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability