Jump to
S2C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -151.669156 |
| Energy at 298.15K | |
| HF Energy | -151.199638 |
| Nuclear repulsion energy | 63.390842 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
2150 |
2031 |
0.00 |
66.63 |
0.39 |
0.56 |
| 2 |
Σg |
946 |
893 |
0.00 |
0.39 |
0.56 |
0.72 |
| 3 |
Σu |
1579 |
1491 |
534.64 |
0.00 |
0.33 |
0.50 |
| 4 |
Πg |
394 |
372 |
0.00 |
2.40 |
0.75 |
0.86 |
| 4 |
Πg |
394 |
372 |
0.00 |
2.40 |
0.75 |
0.86 |
| 5 |
Πu |
179 |
169 |
18.46 |
0.00 |
0.75 |
0.86 |
| 5 |
Πu |
179 |
169 |
18.46 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 2909.6 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 2748.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.646 |
| C2 |
0.000 |
0.000 |
-0.646 |
| C3 |
0.000 |
0.000 |
1.954 |
| C4 |
0.000 |
0.000 |
-1.954 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
| C1 | | 1.2911 | 1.3089 | 2.6000 |
C2 | 1.2911 | | 2.6000 | 1.3089 | C3 | 1.3089 | 2.6000 | | 3.9090 | C4 | 2.6000 | 1.3089 | 3.9090 | |
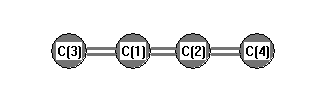 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
180.000 |
|
C2 |
C1 |
C3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -151.656938 |
| Energy at 298.15K | |
| HF Energy | -151.148280 |
| Nuclear repulsion energy | 63.038550 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
2108 |
1991 |
0.00 |
17.59 |
0.38 |
0.55 |
| 2 |
Σg |
919 |
868 |
0.00 |
40.64 |
0.30 |
0.46 |
| 3 |
Σu |
1574 |
1487 |
420.28 |
0.00 |
0.33 |
0.50 |
| 4 |
Πg |
443 |
419 |
0.00 |
2.65 |
0.75 |
0.86 |
| 4 |
Πg |
329 |
311 |
0.00 |
4.00 |
0.75 |
0.86 |
| 5 |
Πu |
207 |
196 |
8.94 |
0.00 |
0.75 |
0.86 |
| 5 |
Πu |
148 |
140 |
35.73 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 2864.6 cm
-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 2705.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/6-31G(2df,p)
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.650 |
| C2 |
0.000 |
0.000 |
-0.650 |
| C3 |
0.000 |
0.000 |
1.965 |
| C4 |
0.000 |
0.000 |
-1.965 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
| C1 | | 1.3003 | 1.3151 | 2.6153 |
C2 | 1.3003 | | 2.6153 | 1.3151 | C3 | 1.3151 | 2.6153 | | 3.9304 | C4 | 2.6153 | 1.3151 | 3.9304 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
