Jump to
S1C2
S1C3
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -208.742510 |
| Energy at 298.15K | -208.748520 |
| HF Energy | -207.983408 |
| Nuclear repulsion energy | 119.427338 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3688 |
3494 |
36.22 |
|
|
|
| 2 |
A' |
3175 |
3008 |
13.64 |
|
|
|
| 3 |
A' |
3079 |
2917 |
43.71 |
|
|
|
| 4 |
A' |
3007 |
2848 |
67.88 |
|
|
|
| 5 |
A' |
1843 |
1746 |
436.86 |
|
|
|
| 6 |
A' |
1565 |
1483 |
15.06 |
|
|
|
| 7 |
A' |
1514 |
1434 |
12.31 |
|
|
|
| 8 |
A' |
1503 |
1424 |
10.78 |
|
|
|
| 9 |
A' |
1431 |
1356 |
13.64 |
|
|
|
| 10 |
A' |
1327 |
1257 |
113.57 |
|
|
|
| 11 |
A' |
1194 |
1131 |
27.77 |
|
|
|
| 12 |
A' |
1030 |
976 |
44.14 |
|
|
|
| 13 |
A' |
620 |
587 |
13.75 |
|
|
|
| 14 |
A' |
348 |
330 |
7.96 |
|
|
|
| 15 |
A" |
3142 |
2976 |
24.45 |
|
|
|
| 16 |
A" |
1517 |
1437 |
4.61 |
|
|
|
| 17 |
A" |
1176 |
1114 |
0.59 |
|
|
|
| 18 |
A" |
1078 |
1021 |
0.06 |
|
|
|
| 19 |
A" |
603 |
572 |
117.09 |
|
|
|
| 20 |
A" |
195 |
185 |
0.97 |
|
|
|
| 21 |
A" |
74 |
70 |
1.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16553.4 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 15682.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.287 |
-0.755 |
0.000 |
| O2 |
1.403 |
-1.226 |
0.000 |
| N3 |
0.000 |
0.572 |
0.000 |
| C4 |
-1.336 |
1.125 |
0.000 |
| H5 |
-0.628 |
-1.375 |
0.000 |
| H6 |
0.794 |
1.191 |
0.000 |
| H7 |
-2.055 |
0.304 |
0.000 |
| H8 |
-1.519 |
1.736 |
0.888 |
| H9 |
-1.519 |
1.736 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2108 | 1.3578 | 2.4834 | 1.1052 | 2.0110 | 2.5704 | 3.2024 | 3.2024 |
O2 | 1.2108 | | 2.2802 | 3.6089 | 2.0359 | 2.4920 | 3.7810 | 4.2539 | 4.2539 | N3 | 1.3578 | 2.2802 | | 1.4456 | 2.0459 | 1.0069 | 2.0726 | 2.1097 | 2.1097 | C4 | 2.4834 | 3.6089 | 1.4456 | | 2.5983 | 2.1311 | 1.0916 | 1.0934 | 1.0934 | H5 | 1.1052 | 2.0359 | 2.0459 | 2.5983 | | 2.9338 | 2.2037 | 3.3557 | 3.3557 | H6 | 2.0110 | 2.4920 | 1.0069 | 2.1311 | 2.9338 | | 2.9843 | 2.5373 | 2.5373 | H7 | 2.5704 | 3.7810 | 2.0726 | 1.0916 | 2.2037 | 2.9843 | | 1.7681 | 1.7681 | H8 | 3.2024 | 4.2539 | 2.1097 | 1.0934 | 3.3557 | 2.5373 | 1.7681 | | 1.7763 | H9 | 3.2024 | 4.2539 | 2.1097 | 1.0934 | 3.3557 | 2.5373 | 1.7681 | 1.7763 | |
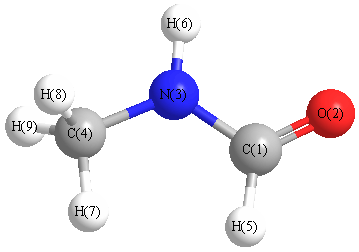 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.684 |
|
C1 |
N3 |
H6 |
115.720 |
| O2 |
C1 |
N3 |
125.079 |
|
O2 |
C1 |
H5 |
122.995 |
| N3 |
C1 |
H5 |
111.926 |
|
N3 |
C4 |
H7 |
108.747 |
| N3 |
C4 |
H8 |
111.631 |
|
N3 |
C4 |
H9 |
111.631 |
| C4 |
N3 |
H6 |
119.597 |
|
H7 |
C4 |
H8 |
108.037 |
| H7 |
C4 |
H9 |
108.037 |
|
H8 |
C4 |
H9 |
108.630 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -208.742510 |
| Energy at 298.15K | -208.748520 |
| HF Energy | -207.983408 |
| Nuclear repulsion energy | 119.427338 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -208.744642 |
| Energy at 298.15K | -208.750630 |
| HF Energy | -207.984915 |
| Nuclear repulsion energy | 121.637892 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3723 |
3527 |
36.76 |
|
|
|
| 2 |
A |
3209 |
3040 |
0.59 |
|
|
|
| 3 |
A |
3150 |
2984 |
24.84 |
|
|
|
| 4 |
A |
3086 |
2924 |
37.90 |
|
|
|
| 5 |
A |
3021 |
2862 |
90.70 |
|
|
|
| 6 |
A |
1828 |
1732 |
301.98 |
|
|
|
| 7 |
A |
1587 |
1504 |
96.16 |
|
|
|
| 8 |
A |
1533 |
1452 |
5.51 |
|
|
|
| 9 |
A |
1527 |
1447 |
44.87 |
|
|
|
| 10 |
A |
1471 |
1394 |
19.00 |
|
|
|
| 11 |
A |
1453 |
1377 |
3.25 |
|
|
|
| 12 |
A |
1250 |
1185 |
84.31 |
|
|
|
| 13 |
A |
1181 |
1119 |
15.84 |
|
|
|
| 14 |
A |
1179 |
1117 |
0.89 |
|
|
|
| 15 |
A |
1060 |
1004 |
0.63 |
|
|
|
| 16 |
A |
985 |
933 |
20.21 |
|
|
|
| 17 |
A |
778 |
737 |
0.82 |
|
|
|
| 18 |
A |
528 |
500 |
36.16 |
|
|
|
| 19 |
A |
306 |
290 |
13.46 |
|
|
|
| 20 |
A |
237 |
225 |
80.13 |
|
|
|
| 21 |
A |
52 |
49 |
0.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16572.6 cm
-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 15700.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.862 |
0.435 |
0.004 |
| O2 |
1.365 |
-0.672 |
-0.000 |
| N3 |
-0.471 |
0.662 |
-0.022 |
| C4 |
-1.414 |
-0.442 |
0.005 |
| H5 |
1.455 |
1.363 |
0.021 |
| H6 |
-0.801 |
1.605 |
0.061 |
| H7 |
-2.412 |
-0.060 |
-0.208 |
| H8 |
-1.139 |
-1.176 |
-0.753 |
| H9 |
-1.419 |
-0.942 |
0.977 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2158 | 1.3522 | 2.4394 | 1.1015 | 2.0344 | 3.3185 | 2.6780 | 2.8370 |
O2 | 1.2158 | | 2.2692 | 2.7888 | 2.0371 | 3.1434 | 3.8324 | 2.6630 | 2.9632 | N3 | 1.3522 | 2.2692 | | 1.4523 | 2.0498 | 1.0030 | 2.0799 | 2.0875 | 2.1144 | C4 | 2.4394 | 2.7888 | 1.4523 | | 3.3898 | 2.1378 | 1.0899 | 1.0905 | 1.0931 | H5 | 1.1015 | 2.0371 | 2.0498 | 3.3898 | | 2.2693 | 4.1273 | 3.7113 | 3.8065 | H6 | 2.0344 | 3.1434 | 1.0030 | 2.1378 | 2.2693 | | 2.3328 | 2.9172 | 2.7769 | H7 | 3.3185 | 3.8324 | 2.0799 | 1.0899 | 4.1273 | 2.3328 | | 1.7787 | 1.7801 | H8 | 2.6780 | 2.6630 | 2.0875 | 1.0905 | 3.7113 | 2.9172 | 1.7787 | | 1.7684 | H9 | 2.8370 | 2.9632 | 2.1144 | 1.0931 | 3.8065 | 2.7769 | 1.7801 | 1.7684 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
120.827 |
|
C1 |
N3 |
H6 |
118.737 |
| O2 |
C1 |
N3 |
124.086 |
|
O2 |
C1 |
H5 |
122.993 |
| N3 |
C1 |
H5 |
112.920 |
|
N3 |
C4 |
H7 |
108.964 |
| N3 |
C4 |
H8 |
109.534 |
|
N3 |
C4 |
H9 |
111.559 |
| C4 |
N3 |
H6 |
119.961 |
|
H7 |
C4 |
H8 |
109.325 |
| H7 |
C4 |
H9 |
109.264 |
|
H8 |
C4 |
H9 |
108.166 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
