Jump to
S1C2
S1C3
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -208.734430 |
| Energy at 298.15K | -208.740454 |
| HF Energy | -207.983885 |
| Nuclear repulsion energy | 119.642315 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3703 |
3507 |
39.08 |
|
|
|
| 2 |
A' |
3190 |
3021 |
12.35 |
|
|
|
| 3 |
A' |
3090 |
2927 |
42.01 |
|
|
|
| 4 |
A' |
3019 |
2860 |
63.81 |
|
|
|
| 5 |
A' |
1882 |
1782 |
477.23 |
|
|
|
| 6 |
A' |
1572 |
1489 |
19.18 |
|
|
|
| 7 |
A' |
1527 |
1446 |
15.96 |
|
|
|
| 8 |
A' |
1509 |
1429 |
11.34 |
|
|
|
| 9 |
A' |
1440 |
1364 |
13.65 |
|
|
|
| 10 |
A' |
1335 |
1264 |
123.80 |
|
|
|
| 11 |
A' |
1201 |
1138 |
26.86 |
|
|
|
| 12 |
A' |
1036 |
981 |
46.35 |
|
|
|
| 13 |
A' |
626 |
593 |
14.09 |
|
|
|
| 14 |
A' |
351 |
332 |
7.95 |
|
|
|
| 15 |
A" |
3156 |
2989 |
22.72 |
|
|
|
| 16 |
A" |
1521 |
1440 |
4.85 |
|
|
|
| 17 |
A" |
1181 |
1118 |
0.60 |
|
|
|
| 18 |
A" |
1089 |
1031 |
0.03 |
|
|
|
| 19 |
A" |
605 |
573 |
119.57 |
|
|
|
| 20 |
A" |
196 |
185 |
0.98 |
|
|
|
| 21 |
A" |
74 |
70 |
1.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16649.5 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 15770.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.289 |
-0.754 |
0.000 |
| O2 |
1.401 |
-1.222 |
0.000 |
| N3 |
0.000 |
0.571 |
0.000 |
| C4 |
-1.336 |
1.121 |
0.000 |
| H5 |
-0.623 |
-1.377 |
0.000 |
| H6 |
0.792 |
1.192 |
0.000 |
| H7 |
-2.054 |
0.300 |
0.000 |
| H8 |
-1.520 |
1.731 |
0.888 |
| H9 |
-1.520 |
1.731 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2059 | 1.3558 | 2.4805 | 1.1046 | 2.0093 | 2.5688 | 3.1992 | 3.1992 |
O2 | 1.2059 | | 2.2747 | 3.6016 | 2.0298 | 2.4887 | 3.7745 | 4.2468 | 4.2468 | N3 | 1.3558 | 2.2747 | | 1.4442 | 2.0447 | 1.0061 | 2.0716 | 2.1082 | 2.1082 | C4 | 2.4805 | 3.6016 | 1.4442 | | 2.5966 | 2.1286 | 1.0908 | 1.0928 | 1.0928 | H5 | 1.1046 | 2.0298 | 2.0447 | 2.5966 | | 2.9322 | 2.2035 | 3.3537 | 3.3537 | H6 | 2.0093 | 2.4887 | 1.0061 | 2.1286 | 2.9322 | | 2.9822 | 2.5347 | 2.5347 | H7 | 2.5688 | 3.7745 | 2.0716 | 1.0908 | 2.2035 | 2.9822 | | 1.7665 | 1.7665 | H8 | 3.1992 | 4.2468 | 2.1082 | 1.0928 | 3.3537 | 2.5347 | 1.7665 | | 1.7750 | H9 | 3.1992 | 4.2468 | 2.1082 | 1.0928 | 3.3537 | 2.5347 | 1.7665 | 1.7750 | |
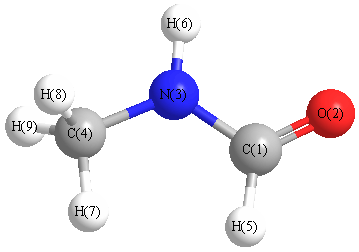 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.682 |
|
C1 |
N3 |
H6 |
115.781 |
| O2 |
C1 |
N3 |
125.133 |
|
O2 |
C1 |
H5 |
122.866 |
| N3 |
C1 |
H5 |
112.001 |
|
N3 |
C4 |
H7 |
108.812 |
| N3 |
C4 |
H8 |
111.649 |
|
N3 |
C4 |
H9 |
111.649 |
| C4 |
N3 |
H6 |
119.537 |
|
H7 |
C4 |
H8 |
107.993 |
| H7 |
C4 |
H9 |
107.993 |
|
H8 |
C4 |
H9 |
108.612 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -208.734430 |
| Energy at 298.15K | -208.740454 |
| HF Energy | -207.983885 |
| Nuclear repulsion energy | 119.642315 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -208.736504 |
| Energy at 298.15K | -208.742476 |
| HF Energy | -207.985410 |
| Nuclear repulsion energy | 121.833764 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3740 |
3543 |
42.05 |
|
|
|
| 2 |
A |
3223 |
3053 |
0.44 |
|
|
|
| 3 |
A |
3164 |
2997 |
23.02 |
|
|
|
| 4 |
A |
3098 |
2934 |
36.01 |
|
|
|
| 5 |
A |
3033 |
2873 |
86.36 |
|
|
|
| 6 |
A |
1868 |
1770 |
337.68 |
|
|
|
| 7 |
A |
1596 |
1512 |
112.07 |
|
|
|
| 8 |
A |
1536 |
1455 |
5.83 |
|
|
|
| 9 |
A |
1533 |
1452 |
42.53 |
|
|
|
| 10 |
A |
1479 |
1401 |
19.60 |
|
|
|
| 11 |
A |
1465 |
1388 |
1.20 |
|
|
|
| 12 |
A |
1260 |
1194 |
85.82 |
|
|
|
| 13 |
A |
1187 |
1124 |
17.18 |
|
|
|
| 14 |
A |
1184 |
1121 |
0.84 |
|
|
|
| 15 |
A |
1071 |
1015 |
0.70 |
|
|
|
| 16 |
A |
992 |
939 |
22.49 |
|
|
|
| 17 |
A |
785 |
743 |
1.13 |
|
|
|
| 18 |
A |
529 |
501 |
37.16 |
|
|
|
| 19 |
A |
308 |
292 |
13.43 |
|
|
|
| 20 |
A |
240 |
227 |
81.99 |
|
|
|
| 21 |
A |
36 |
34 |
0.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16662.3 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 15782.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.862 |
0.432 |
0.004 |
| O2 |
1.361 |
-0.671 |
0.000 |
| N3 |
-0.468 |
0.661 |
-0.022 |
| C4 |
-1.413 |
-0.440 |
0.005 |
| H5 |
1.459 |
1.356 |
0.021 |
| H6 |
-0.795 |
1.604 |
0.062 |
| H7 |
-2.410 |
-0.056 |
-0.206 |
| H8 |
-1.141 |
-1.172 |
-0.754 |
| H9 |
-1.418 |
-0.941 |
0.976 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2105 | 1.3498 | 2.4365 | 1.1004 | 2.0311 | 3.3152 | 2.6760 | 2.8336 |
O2 | 1.2105 | | 2.2622 | 2.7832 | 2.0299 | 3.1352 | 3.8262 | 2.6604 | 2.9572 | N3 | 1.3498 | 2.2622 | | 1.4510 | 2.0489 | 1.0020 | 2.0786 | 2.0859 | 2.1127 | C4 | 2.4365 | 2.7832 | 1.4510 | | 3.3874 | 2.1358 | 1.0892 | 1.0899 | 1.0924 | H5 | 1.1004 | 2.0299 | 2.0489 | 3.3874 | | 2.2686 | 4.1251 | 3.7088 | 3.8035 | H6 | 2.0311 | 3.1352 | 1.0020 | 2.1358 | 2.2686 | | 2.3310 | 2.9145 | 2.7749 | H7 | 3.3152 | 3.8262 | 2.0786 | 1.0892 | 4.1251 | 2.3310 | | 1.7773 | 1.7790 | H8 | 2.6760 | 2.6604 | 2.0859 | 1.0899 | 3.7088 | 2.9145 | 1.7773 | | 1.7675 | H9 | 2.8336 | 2.9572 | 2.1127 | 1.0924 | 3.8035 | 2.7749 | 1.7790 | 1.7675 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
120.856 |
|
C1 |
N3 |
H6 |
118.713 |
| O2 |
C1 |
N3 |
124.068 |
|
O2 |
C1 |
H5 |
122.833 |
| N3 |
C1 |
H5 |
113.098 |
|
N3 |
C4 |
H7 |
108.989 |
| N3 |
C4 |
H8 |
109.533 |
|
N3 |
C4 |
H9 |
111.555 |
| C4 |
N3 |
H6 |
119.955 |
|
H7 |
C4 |
H8 |
109.295 |
| H7 |
C4 |
H9 |
109.261 |
|
H8 |
C4 |
H9 |
108.177 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
