return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (1-Butanol, 3-methyl-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-272.266052
Energy at 298.15K-272.279535
HF Energy-271.213316
Nuclear repulsion energy257.371565
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3887 3671 32.20      
2 A' 3192 3015 39.88      
3 A' 3189 3012 33.56      
4 A' 3103 2931 25.80      
5 A' 3097 2926 14.22      
6 A' 3090 2918 9.18      
7 A' 3060 2891 32.80      
8 A' 1560 1474 1.42      
9 A' 1541 1456 5.53      
10 A' 1537 1452 10.30      
11 A' 1513 1429 2.18      
12 A' 1480 1398 3.40      
13 A' 1439 1360 4.15      
14 A' 1386 1309 1.62      
15 A' 1354 1279 26.98      
16 A' 1266 1196 23.81      
17 A' 1227 1159 4.18      
18 A' 1144 1080 0.52      
19 A' 1097 1036 87.84      
20 A' 1024 967 5.89      
21 A' 981 927 9.00      
22 A' 823 777 4.46      
23 A' 560 529 1.02      
24 A' 405 382 5.90      
25 A' 326 308 4.94      
26 A' 269 254 0.00      
27 A' 217 205 2.27      
28 A" 3189 3012 8.93      
29 A" 3187 3010 3.37      
30 A" 3159 2984 28.12      
31 A" 3107 2935 46.70      
32 A" 3094 2922 6.77      
33 A" 1528 1443 0.95      
34 A" 1514 1430 0.59      
35 A" 1421 1342 8.13      
36 A" 1385 1308 0.69      
37 A" 1337 1263 0.11      
38 A" 1265 1195 0.07      
39 A" 1201 1134 3.96      
40 A" 1028 970 0.52      
41 A" 979 925 0.00      
42 A" 936 884 0.97      
43 A" 802 758 0.04      
44 A" 369 349 0.03      
45 A" 290 274 102.66      
46 A" 255 240 3.40      
47 A" 110 104 0.00      
48 A" 61 58 9.02      

Unscaled Zero Point Vibrational Energy (zpe) 36992.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 34939.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.20150 0.06296 0.05649

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 1.217 -0.337 2.163
H2 1.217 -0.337 -2.163
H3 0.484 -1.679 -1.279
H4 0.484 -1.679 1.279
H5 2.233 -1.493 -1.295
H6 2.233 -1.493 1.295
H7 0.034 1.504 0.878
H8 0.034 1.504 -0.878
H9 -1.450 -0.487 -0.885
H10 -1.450 -0.487 0.885
C11 1.290 -0.943 -1.257
C12 1.290 -0.943 1.257
C13 0.000 0.851 0.000
C14 -1.349 0.153 0.000
H15 -3.192 0.750 0.000
O16 -2.332 1.177 0.000
H17 2.101 0.576 0.000
C18 1.221 -0.076 0.000

Atom - Atom Distances (Å)
  H1 H2 H3 H4 H5 H6 H7 H8 H9 H10 C11 C12 C13 C14 H15 O16 H17 C18
H14.32623.76641.76663.78551.76662.53793.74664.05352.96143.47441.09232.75213.39185.03004.42402.50902.1789
H24.32621.76663.76641.76663.78553.74662.53792.96144.05351.09233.47442.75213.39185.03004.42402.50902.1789
H33.76641.76662.55771.75933.11803.87113.23952.30583.13791.09212.76122.87652.88974.58764.21003.05602.1794
H41.76663.76642.55773.11801.75933.23953.87113.13792.30582.76121.09212.87652.88974.58764.21003.05602.1794
H53.78551.76661.75933.11802.59104.30603.74063.84024.39721.09212.77633.48754.14956.01165.44542.44512.1709
H61.76663.78553.11801.75932.59103.74064.30604.39723.84022.77631.09213.48754.14956.01165.44542.44512.1709
H72.53793.74663.87113.23954.30603.74061.75603.04532.48283.48212.77671.09432.12333.42722.54512.42992.1618
H83.74662.53793.23953.87113.74064.30601.75602.48283.04532.77673.48211.09432.12333.42722.54512.42992.1618
H94.05352.96142.30583.13793.84024.39723.04532.48281.77082.80303.50852.16281.09692.31222.08093.81132.8435
H102.96144.05353.13792.30584.39723.84022.48283.04531.77083.50852.80302.16281.09692.31222.08093.81132.8435
C113.47441.09231.09212.76121.09212.77633.48212.77672.80303.50852.51452.54303.12214.95354.38182.13241.5291
C121.09233.47442.76121.09212.77631.09212.77673.48213.50852.80302.51452.54303.12214.95354.38182.13241.5291
C132.75212.75212.87652.87653.48753.48751.09431.09432.16282.16282.54302.54301.51913.19352.35502.11921.5328
C143.39183.39182.88972.88974.14954.14952.12332.12331.09691.09693.12213.12211.51911.93731.42003.47602.5796
H155.03005.03004.58764.58766.01166.01163.42723.42722.31222.31224.95354.95353.19351.93730.95995.29604.4890
O164.42404.42404.21004.21005.44545.44542.54512.54512.08092.08094.38184.38182.35501.42000.95994.47423.7673
H172.50902.50903.05603.05602.44512.44512.42992.42993.81133.81132.13242.13242.11923.47605.29604.47421.0958
C182.17892.17892.17942.17942.17092.17092.16182.16182.84352.84351.52911.52911.53282.57964.48903.76731.0958

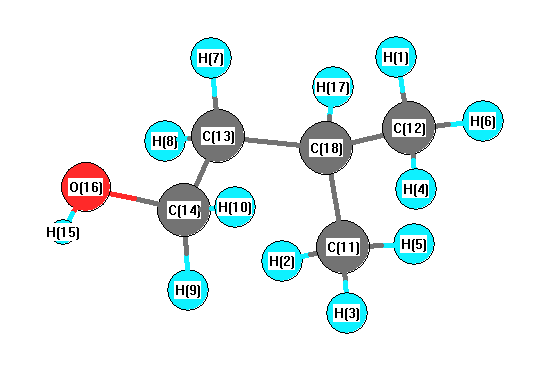
picture of 1-Butanol, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C12 H4 107.941 H1 C12 H6 107.945
H1 C12 C18 111.349 H2 C11 H3 107.941
H2 C11 H5 107.945 H2 C11 C18 111.349
H3 C11 H5 107.307 H3 C11 C18 111.400
H4 C12 H6 107.307 H4 C12 C18 111.400
H5 C11 C18 110.727 H6 C12 C18 110.727
H7 C13 H8 106.712 H7 C13 C14 107.562
H7 C13 C18 109.621 H8 C13 C14 107.562
H8 C13 C18 109.621 H9 C14 H10 107.651
H9 C14 C13 110.493 H9 C14 O16 110.891
H10 C14 C13 110.493 H10 C14 O16 110.891
C11 C18 C12 110.613 C11 C18 C13 112.306
C11 C18 H17 107.510 C12 C18 C13 112.306
C12 C18 H17 107.510 C13 C14 O16 106.453
C13 C18 H17 106.266 C14 C13 C18 115.394
C14 O16 H15 107.403
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability