return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CHOCH2CH2CH3 (Butanal)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-231.871769
Energy at 298.15K-231.880411
HF Energy-231.012094
Nuclear repulsion energy172.373913
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3199 3022 18.70      
2 A' 3126 2952 11.95      
3 A' 3102 2930 16.49      
4 A' 3080 2909 12.25      
5 A' 2961 2797 122.78      
6 A' 1790 1691 91.89      
7 A' 1537 1451 4.83      
8 A' 1516 1431 2.90      
9 A' 1477 1395 12.19      
10 A' 1438 1358 4.69      
11 A' 1434 1354 3.35      
12 A' 1409 1331 16.23      
13 A' 1319 1246 4.95      
14 A' 1155 1091 10.06      
15 A' 1083 1023 0.05      
16 A' 979 925 5.42      
17 A' 877 829 7.67      
18 A' 701 662 13.00      
19 A' 347 327 0.85      
20 A' 196 185 7.59      
21 A" 3196 3019 30.23      
22 A" 3173 2997 1.03      
23 A" 3124 2951 4.39      
24 A" 1532 1447 6.48      
25 A" 1338 1263 0.15      
26 A" 1267 1197 0.08      
27 A" 1166 1101 0.02      
28 A" 955 902 0.00      
29 A" 796 752 1.47      
30 A" 678 640 3.40      
31 A" 258 244 0.02      
32 A" 181 171 1.65      
33 A" 84 79 1.61      

Unscaled Zero Point Vibrational Energy (zpe) 25235.3 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 23834.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.49967 0.08626 0.07665

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.269 -0.348 0.000
H2 2.800 -1.301 0.000
H3 2.588 0.212 0.882
H4 2.588 0.212 -0.882
C5 0.758 -0.558 0.000
H6 0.454 -1.141 -0.872
H7 0.454 -1.141 0.872
C8 0.000 0.761 0.000
H9 0.262 1.375 0.871
H10 0.262 1.375 -0.871
C11 -1.496 0.594 0.000
H12 -2.076 1.540 0.000
O13 -2.065 -0.478 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 O13
C11.09081.09211.09211.52542.16382.16382.52562.78502.78503.88134.73734.3353
H21.09081.76361.76362.17312.50762.50763.47753.78973.78974.69575.64314.9337
H31.09211.76361.76312.17233.07552.52682.78882.60083.13624.19594.92894.7854
H41.09211.76361.76312.17232.52683.07552.78883.13622.60084.19594.92894.7854
C51.52542.17312.17232.17231.09231.09231.52142.17752.17752.53153.52582.8236
H62.16382.50763.07552.52681.09231.74382.14173.06732.52402.75263.78842.7468
H72.16382.50762.52683.07551.09231.74382.14172.52403.06732.75263.78842.7468
C82.52563.47752.78882.78881.52142.14172.14171.09741.09741.50572.21712.4080
H92.78503.78972.60083.13622.17753.06732.52401.09741.74222.11212.50043.0994
H102.78503.78973.13622.60082.17752.52403.06731.09741.74222.11212.50043.0994
C113.88134.69574.19594.19592.53152.75262.75261.50572.11212.11211.10941.2132
H124.73735.64314.92894.92893.52583.78843.78842.21712.50042.50041.10942.0179
O134.33534.93374.78544.78542.82362.74682.74682.40803.09943.09941.21322.0179

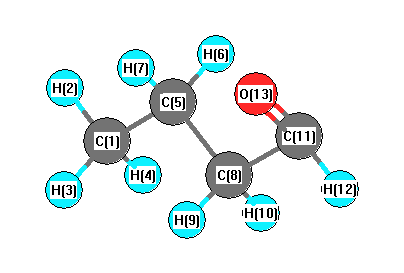
picture of Butanal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.417 C1 C5 H7 110.417
C1 C5 C8 111.980 H2 C1 H3 107.788
H2 C1 H4 107.788 H2 C1 C5 111.243
H3 C1 H4 107.652 H3 C1 C5 111.102
H4 C1 C5 111.102 C5 C8 H9 111.474
C5 C8 H10 111.474 C5 C8 C11 113.497
H6 C5 H7 105.926 H6 C5 C8 108.952
H7 C5 C8 108.952 C8 C11 H12 115.112
C8 C11 O13 124.308 H9 C8 H10 105.081
H9 C8 C11 107.424 H10 C8 C11 107.424
H12 C11 O13 120.580
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability