return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NHCH3 (Dimethylamine)

using model chemistry: QCISD/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G(2df,p)
 hartrees
Energy at 0K-134.836489
Energy at 298.15K-134.844712
HF Energy-134.258652
Nuclear repulsion energy84.196909
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3581 3393 0.29      
2 A' 3163 2996 30.89      
3 A' 3116 2952 46.82      
4 A' 3016 2857 94.57      
5 A' 1551 1470 1.24      
6 A' 1533 1453 9.62      
7 A' 1502 1423 0.44      
8 A' 1297 1229 0.55      
9 A' 1221 1156 8.90      
10 A' 970 919 1.06      
11 A' 822 779 104.77      
12 A' 393 372 5.04      
13 A' 279 264 2.28      
14 A" 3161 2995 19.25      
15 A" 3115 2952 18.30      
16 A" 3012 2854 35.03      
17 A" 1548 1467 9.96      
18 A" 1519 1439 3.70      
19 A" 1506 1427 5.41      
20 A" 1472 1395 2.06      
21 A" 1202 1139 26.67      
22 A" 1119 1060 0.15      
23 A" 1050 994 8.10      
24 A" 235 223 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 20690.6 cm-1
Scaled (by 0.9474) Zero Point Vibrational Energy (zpe) 19602.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G(2df,p)
ABC
1.14163 0.31368 0.27612

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.027 0.588 0.000
H2 -0.788 1.187 0.000
C3 0.027 -0.223 1.207
C4 0.027 -0.223 -1.207
H5 -0.798 -0.953 1.262
H6 -0.798 -0.953 -1.262
H7 0.965 -0.783 1.266
H8 0.965 -0.783 -1.266
H9 -0.029 0.424 2.085
H10 -0.029 0.424 -2.085

Atom - Atom Distances (Å)
  N1 H2 C3 C4 H5 H6 H7 H8 H9 H10
N11.01121.45401.45402.15572.15572.08812.08812.09222.0922
H21.01122.02682.02682.48432.48432.92462.92462.34612.3461
C31.45402.02682.41391.10252.70371.09412.70321.09233.3555
C41.45402.02682.41392.70371.10252.70321.09413.35551.0923
H52.15572.48431.10252.70372.52461.77113.08671.77883.7003
H62.15572.48432.70371.10252.52463.08671.77113.70031.7788
H72.08812.92461.09412.70321.77113.08672.53131.76543.6976
H82.08812.92462.70321.09413.08671.77112.53133.69761.7654
H92.09222.34611.09233.35551.77883.70031.76543.69764.1700
H102.09222.34613.35551.09233.70031.77883.69761.76544.1700

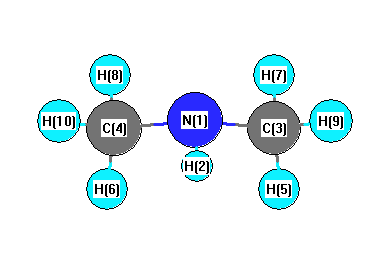
picture of Dimethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C3 H5 114.257 N1 C3 H7 109.250
N1 C3 H9 109.683 N1 C4 H6 114.257
N1 C4 H8 109.250 N1 C4 H10 109.683
H2 N1 C3 109.286 H2 N1 C4 109.286
C3 N1 C4 112.217 H5 C3 H7 107.468
H5 C3 H9 108.277 H6 C4 H8 107.468
H6 C4 H10 108.277 H7 C3 H9 107.691
H8 C4 H10 107.691
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability