Jump to
S1C2
Energy calculated at PBEPBE/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -399.475490 |
| Energy at 298.15K | |
| HF Energy | -399.475490 |
| Nuclear repulsion energy | 62.599580 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBEPBE/6-31G(2df,p)
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.000 |
| Na2 |
0.000 |
0.000 |
1.999 |
| Na3 |
0.000 |
0.000 |
-1.999 |
Atom - Atom Distances (Å)
| |
O1 |
Na2 |
Na3 |
| O1 | | 1.9992 | 1.9992 |
Na2 | 1.9992 | | 3.9984 | Na3 | 1.9992 | 3.9984 | |
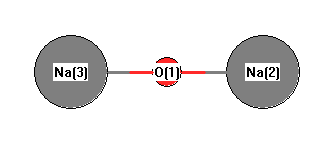 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
O1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.548 |
|
|
|
| 2 |
Na |
0.274 |
|
|
|
| 3 |
Na |
0.274 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.264 |
0.000 |
0.000 |
| y |
0.000 |
-21.264 |
0.000 |
| z |
0.000 |
0.000 |
9.430 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-15.347 |
0.000 |
0.000 |
| y |
0.000 |
-15.347 |
0.000 |
| z |
0.000 |
0.000 |
30.694 |
|
| Polar |
|---|
| 3z2-r2 | 61.389 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.055 |
0.000 |
0.000 |
| y |
0.000 |
14.055 |
0.000 |
| z |
0.000 |
0.000 |
30.931 |
<r2> (average value of r
2) Å
2
| <r2> |
94.822 |
| (<r2>)1/2 |
9.738 |
Jump to
S1C1
Energy calculated at PBEPBE/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -399.475516 |
| Energy at 298.15K | -399.476914 |
| HF Energy | -399.475516 |
| Nuclear repulsion energy | 62.674917 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at PBEPBE/6-31G(2df,p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.149 |
| Na2 |
0.000 |
1.989 |
-0.054 |
| Na3 |
0.000 |
-1.989 |
-0.054 |
Atom - Atom Distances (Å)
| |
O1 |
Na2 |
Na3 |
| O1 | | 1.9995 | 1.9995 |
Na2 | 1.9995 | | 3.9784 | Na3 | 1.9995 | 3.9784 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
O1 |
Na3 |
168.372 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.550 |
|
|
|
| 2 |
Na |
0.275 |
|
|
|
| 3 |
Na |
0.275 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.846 |
0.846 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.281 |
0.000 |
0.000 |
| y |
0.000 |
8.994 |
0.000 |
| z |
0.000 |
0.000 |
-21.318 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-15.118 |
0.000 |
0.000 |
| y |
0.000 |
30.293 |
0.000 |
| z |
0.000 |
0.000 |
-15.175 |
|
| Polar |
|---|
| 3z2-r2 | -30.349 |
|---|
| x2-y2 | -30.274 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.926 |
0.000 |
0.000 |
| y |
0.000 |
30.885 |
0.000 |
| z |
0.000 |
0.000 |
13.769 |
<r2> (average value of r
2) Å
2
| <r2> |
94.287 |
| (<r2>)1/2 |
9.710 |
