Jump to
S1C2
Energy calculated at M06-2X/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -166.471503 |
| Energy at 298.15K | |
| HF Energy | -166.471503 |
| Nuclear repulsion energy | 49.225740 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
4065 |
3872 |
67.42 |
|
|
|
| 2 |
A1 |
767 |
730 |
6.94 |
|
|
|
| 3 |
A1 |
602 |
573 |
152.82 |
|
|
|
| 4 |
A1 |
316 |
301 |
11.81 |
|
|
|
| 5 |
A2 |
81i |
77i |
0.00 |
|
|
|
| 6 |
B1 |
354 |
338 |
59.30 |
|
|
|
| 7 |
B2 |
4062 |
3869 |
174.76 |
|
|
|
| 8 |
B2 |
1586 |
1511 |
347.15 |
|
|
|
| 9 |
B2 |
499 |
475 |
288.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6085.0 cm
-1
Scaled (by 0.9524) Zero Point Vibrational Energy (zpe) 5795.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-31G(2df,p)
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
0.007 |
| O2 |
0.000 |
1.416 |
0.077 |
| O3 |
0.000 |
-1.416 |
0.077 |
| H4 |
0.000 |
2.054 |
-0.629 |
| H5 |
0.000 |
-2.054 |
-0.629 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4182 | 1.4182 | 2.1500 | 2.1500 |
O2 | 1.4182 | | 2.8330 | 0.9512 | 3.5412 | O3 | 1.4182 | 2.8330 | | 3.5412 | 0.9512 | H4 | 2.1500 | 0.9512 | 3.5412 | | 4.1072 | H5 | 2.1500 | 3.5412 | 0.9512 | 4.1072 | |
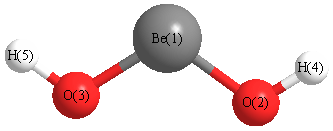 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
129.240 |
|
Be1 |
O3 |
H5 |
129.240 |
| O2 |
Be1 |
O3 |
174.375 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
-0.043 |
|
|
|
| 2 |
O |
-0.305 |
|
|
|
| 3 |
O |
-0.305 |
|
|
|
| 4 |
H |
0.327 |
|
|
|
| 5 |
H |
0.327 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.415 |
2.415 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.411 |
0.000 |
0.000 |
| y |
0.000 |
-14.039 |
0.000 |
| z |
0.000 |
0.000 |
-14.070 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.356 |
0.000 |
0.000 |
| y |
0.000 |
1.202 |
0.000 |
| z |
0.000 |
0.000 |
1.155 |
|
| Polar |
|---|
| 3z2-r2 | 2.309 |
|---|
| x2-y2 | -2.372 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.292 |
0.000 |
0.000 |
| y |
0.000 |
3.504 |
0.000 |
| z |
0.000 |
0.000 |
2.446 |
<r2> (average value of r
2) Å
2
| <r2> |
50.693 |
| (<r2>)1/2 |
7.120 |
Jump to
S1C1
Energy calculated at M06-2X/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -166.472473 |
| Energy at 298.15K | -166.474012 |
| HF Energy | -166.472473 |
| Nuclear repulsion energy | 49.206897 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4036 |
3844 |
31.28 |
|
|
|
| 2 |
A |
762 |
726 |
1.70 |
|
|
|
| 3 |
A |
575 |
548 |
88.58 |
|
|
|
| 4 |
A |
322 |
306 |
22.24 |
|
|
|
| 5 |
A |
237 |
226 |
131.26 |
|
|
|
| 6 |
B |
4034 |
3842 |
194.43 |
|
|
|
| 7 |
B |
1585 |
1510 |
339.85 |
|
|
|
| 8 |
B |
572 |
545 |
333.39 |
|
|
|
| 9 |
B |
310 |
295 |
74.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6216.5 cm
-1
Scaled (by 0.9524) Zero Point Vibrational Energy (zpe) 5920.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-31G(2df,p)
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
-0.010 |
| O2 |
0.000 |
1.419 |
-0.053 |
| O3 |
0.000 |
-1.419 |
-0.053 |
| H4 |
0.566 |
2.006 |
0.440 |
| H5 |
-0.566 |
-2.006 |
0.440 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4197 | 1.4197 | 2.1319 | 2.1319 |
O2 | 1.4197 | | 2.8381 | 0.9523 | 3.5058 | O3 | 1.4197 | 2.8381 | | 3.5058 | 0.9523 | H4 | 2.1319 | 0.9523 | 3.5058 | | 4.1677 | H5 | 2.1319 | 3.5058 | 0.9523 | 4.1677 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
126.869 |
|
Be1 |
O3 |
H5 |
126.869 |
| O2 |
Be1 |
O3 |
176.551 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
Be |
-0.033 |
|
|
|
| 2 |
O |
-0.310 |
|
|
|
| 3 |
O |
-0.310 |
|
|
|
| 4 |
H |
0.326 |
|
|
|
| 5 |
H |
0.326 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.669 |
1.669 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.689 |
4.427 |
0.000 |
| y |
4.427 |
-14.869 |
0.000 |
| z |
0.000 |
0.000 |
-15.293 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.392 |
4.427 |
0.000 |
| y |
4.427 |
0.122 |
0.000 |
| z |
0.000 |
0.000 |
-0.514 |
|
| Polar |
|---|
| 3z2-r2 | -1.028 |
|---|
| x2-y2 | 0.181 |
|---|
| xy | 4.427 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.406 |
0.184 |
0.000 |
| y |
0.184 |
3.479 |
0.000 |
| z |
0.000 |
0.000 |
2.380 |
<r2> (average value of r
2) Å
2
| <r2> |
50.674 |
| (<r2>)1/2 |
7.119 |
