Jump to
S1C2
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -1194.277420 |
| Energy at 298.15K | -1194.279869 |
| HF Energy | -1193.722671 |
| Nuclear repulsion energy | 197.216163 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2755 |
2589 |
0.06 |
|
|
|
| 2 |
A |
908 |
853 |
0.15 |
|
|
|
| 3 |
A |
524 |
492 |
0.43 |
|
|
|
| 4 |
A |
321 |
302 |
24.41 |
|
|
|
| 5 |
A |
214 |
201 |
0.00 |
|
|
|
| 6 |
B |
2754 |
2589 |
0.48 |
|
|
|
| 7 |
B |
895 |
841 |
10.09 |
|
|
|
| 8 |
B |
525 |
493 |
21.79 |
|
|
|
| 9 |
B |
347 |
326 |
18.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4620.7 cm
-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 4343.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.837 |
| S2 |
0.000 |
1.647 |
-0.383 |
| S3 |
0.000 |
-1.647 |
-0.383 |
| H4 |
-1.323 |
1.746 |
-0.566 |
| H5 |
1.323 |
-1.746 |
-0.566 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0500 | 2.0500 | 2.6011 | 2.6011 |
S2 | 2.0500 | | 3.2939 | 1.3390 | 3.6459 | S3 | 2.0500 | 3.2939 | | 3.6459 | 1.3390 | H4 | 2.6011 | 1.3390 | 3.6459 | | 4.3803 | H5 | 2.6011 | 3.6459 | 1.3390 | 4.3803 | |
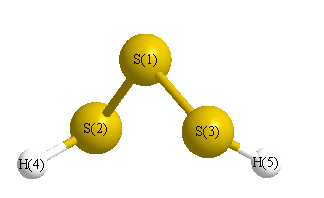 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.069 |
|
S1 |
S3 |
H5 |
98.069 |
| S2 |
S1 |
S3 |
106.909 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -1194.277243 |
| Energy at 298.15K | -1194.279713 |
| HF Energy | -1193.722445 |
| Nuclear repulsion energy | 197.299075 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2750 |
2584 |
0.20 |
|
|
|
| 2 |
A' |
912 |
857 |
3.72 |
|
|
|
| 3 |
A' |
524 |
493 |
0.43 |
|
|
|
| 4 |
A' |
348 |
327 |
18.48 |
|
|
|
| 5 |
A' |
215 |
202 |
0.05 |
|
|
|
| 6 |
A" |
2752 |
2587 |
0.00 |
|
|
|
| 7 |
A" |
898 |
844 |
7.42 |
|
|
|
| 8 |
A" |
526 |
495 |
23.70 |
|
|
|
| 9 |
A" |
334 |
314 |
10.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4629.1 cm
-1
Scaled (by 0.9399) Zero Point Vibrational Energy (zpe) 4350.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.053 |
0.835 |
0.000 |
| S2 |
-0.053 |
-0.386 |
1.646 |
| S3 |
-0.053 |
-0.386 |
-1.646 |
| H4 |
1.273 |
-0.513 |
1.785 |
| H5 |
1.273 |
-0.513 |
-1.785 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0490 | 2.0490 | 2.6004 | 2.6004 |
S2 | 2.0490 | | 3.2910 | 1.3392 | 3.6803 | S3 | 2.0490 | 3.2910 | | 3.6803 | 1.3392 | H4 | 2.6004 | 1.3392 | 3.6803 | | 3.5706 | H5 | 2.6004 | 3.6803 | 1.3392 | 3.5706 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.069 |
|
S1 |
S3 |
H5 |
98.069 |
| S2 |
S1 |
S3 |
106.851 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
