Jump to
S1C2
Energy calculated at mPW1PW91/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -454.722731 |
| Energy at 298.15K | -454.726510 |
| HF Energy | -454.722731 |
| Nuclear repulsion energy | 58.022525 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3568 |
3406 |
8.54 |
|
|
|
| 2 |
A' |
2677 |
2556 |
14.44 |
|
|
|
| 3 |
A' |
1637 |
1563 |
11.90 |
|
|
|
| 4 |
A' |
1058 |
1010 |
8.27 |
|
|
|
| 5 |
A' |
903 |
862 |
34.52 |
|
|
|
| 6 |
A' |
652 |
622 |
89.87 |
|
|
|
| 7 |
A" |
3666 |
3500 |
23.67 |
|
|
|
| 8 |
A" |
1145 |
1093 |
1.35 |
|
|
|
| 9 |
A" |
460 |
439 |
47.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7882.3 cm
-1
Scaled (by 0.9547) Zero Point Vibrational Energy (zpe) 7525.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.015 |
1.099 |
0.000 |
| S2 |
0.015 |
-0.612 |
0.000 |
| H3 |
-1.320 |
-0.790 |
0.000 |
| H4 |
0.492 |
1.440 |
0.823 |
| H5 |
0.492 |
1.440 |
-0.823 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7111 | 2.3129 | 1.0105 | 1.0105 |
S2 | 1.7111 | | 1.3465 | 2.2613 | 2.2613 | H3 | 2.3129 | 1.3465 | | 2.9883 | 2.9883 | H4 | 1.0105 | 2.2613 | 2.9883 | | 1.6463 | H5 | 1.0105 | 2.2613 | 2.9883 | 1.6463 | |
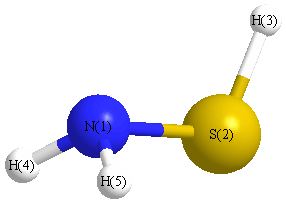 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
97.592 |
|
S2 |
N1 |
H4 |
109.678 |
| S2 |
N1 |
H5 |
109.678 |
|
H4 |
N1 |
H5 |
109.094 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.554 |
|
|
|
| 2 |
S |
-0.181 |
|
|
|
| 3 |
H |
0.168 |
|
|
|
| 4 |
H |
0.284 |
|
|
|
| 5 |
H |
0.284 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.492 |
1.070 |
0.000 |
1.178 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.749 |
3.478 |
0.000 |
| y |
3.478 |
-18.597 |
0.000 |
| z |
0.000 |
0.000 |
-20.003 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.551 |
3.478 |
0.000 |
| y |
3.478 |
0.780 |
0.000 |
| z |
0.000 |
0.000 |
-1.331 |
|
| Polar |
|---|
| 3z2-r2 | -2.661 |
|---|
| x2-y2 | -0.153 |
|---|
| xy | 3.478 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.405 |
0.447 |
0.000 |
| y |
0.447 |
4.425 |
0.000 |
| z |
0.000 |
0.000 |
3.182 |
<r2> (average value of r
2) Å
2
| <r2> |
34.743 |
| (<r2>)1/2 |
5.894 |
Jump to
S1C1
Energy calculated at mPW1PW91/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -454.722311 |
| Energy at 298.15K | -454.726141 |
| HF Energy | -454.722311 |
| Nuclear repulsion energy | 58.288566 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3571 |
3409 |
3.87 |
|
|
|
| 2 |
A' |
2593 |
2475 |
44.86 |
|
|
|
| 3 |
A' |
1624 |
1550 |
12.21 |
|
|
|
| 4 |
A' |
1032 |
985 |
27.80 |
|
|
|
| 5 |
A' |
903 |
862 |
15.25 |
|
|
|
| 6 |
A' |
622 |
593 |
144.17 |
|
|
|
| 7 |
A" |
3679 |
3512 |
22.93 |
|
|
|
| 8 |
A" |
1124 |
1073 |
1.88 |
|
|
|
| 9 |
A" |
558 |
533 |
2.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7851.8 cm
-1
Scaled (by 0.9547) Zero Point Vibrational Energy (zpe) 7496.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.083 |
1.088 |
0.000 |
| S2 |
0.083 |
-0.604 |
0.000 |
| H3 |
-1.236 |
-0.919 |
0.000 |
| H4 |
-0.336 |
1.480 |
0.831 |
| H5 |
-0.336 |
1.480 |
-0.831 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.6918 | 2.4015 | 1.0094 | 1.0094 |
S2 | 1.6918 | | 1.3562 | 2.2822 | 2.2822 | H3 | 2.4015 | 1.3562 | | 2.6937 | 2.6937 | H4 | 1.0094 | 2.2822 | 2.6937 | | 1.6610 | H5 | 1.0094 | 2.2822 | 2.6937 | 1.6610 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
103.428 |
|
S2 |
N1 |
H4 |
112.868 |
| S2 |
N1 |
H5 |
112.868 |
|
H4 |
N1 |
H5 |
110.732 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.540 |
|
|
|
| 2 |
S |
-0.162 |
|
|
|
| 3 |
H |
0.140 |
|
|
|
| 4 |
H |
0.282 |
|
|
|
| 5 |
H |
0.282 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.022 |
1.205 |
0.000 |
2.354 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.617 |
-0.968 |
0.000 |
| y |
-0.968 |
-17.860 |
0.000 |
| z |
0.000 |
0.000 |
-19.922 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.726 |
-0.968 |
0.000 |
| y |
-0.968 |
1.910 |
0.000 |
| z |
0.000 |
0.000 |
-1.184 |
|
| Polar |
|---|
| 3z2-r2 | -2.369 |
|---|
| x2-y2 | -1.757 |
|---|
| xy | -0.968 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.352 |
0.208 |
0.000 |
| y |
0.208 |
4.416 |
0.000 |
| z |
0.000 |
0.000 |
3.172 |
<r2> (average value of r
2) Å
2
| <r2> |
34.587 |
| (<r2>)1/2 |
5.881 |
