Jump to
S1C2
Energy calculated at B3LYPultrafine/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -454.734880 |
| Energy at 298.15K | -454.738625 |
| HF Energy | -454.734880 |
| Nuclear repulsion energy | 57.556981 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYPultrafine/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3514 |
3391 |
5.69 |
|
|
|
| 2 |
A' |
2624 |
2532 |
16.79 |
|
|
|
| 3 |
A' |
1629 |
1572 |
10.67 |
|
|
|
| 4 |
A' |
1042 |
1006 |
8.30 |
|
|
|
| 5 |
A' |
897 |
866 |
41.36 |
|
|
|
| 6 |
A' |
633 |
611 |
69.86 |
|
|
|
| 7 |
A" |
3605 |
3479 |
16.52 |
|
|
|
| 8 |
A" |
1136 |
1096 |
1.19 |
|
|
|
| 9 |
A" |
438 |
422 |
47.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7758.5 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 7487.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYPultrafine/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.014 |
1.112 |
0.000 |
| S2 |
0.014 |
-0.618 |
0.000 |
| H3 |
-1.325 |
-0.793 |
0.000 |
| H4 |
0.502 |
1.445 |
0.823 |
| H5 |
0.502 |
1.445 |
-0.823 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7296 | 2.3286 | 1.0136 | 1.0136 |
S2 | 1.7296 | | 1.3504 | 2.2742 | 2.2742 | H3 | 2.3286 | 1.3504 | | 3.0045 | 3.0045 | H4 | 1.0136 | 2.2742 | 3.0045 | | 1.6469 | H5 | 1.0136 | 2.2742 | 3.0045 | 1.6469 | |
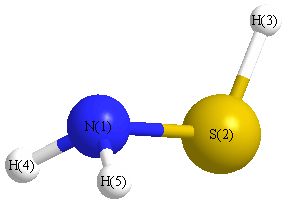 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
97.470 |
|
S2 |
N1 |
H4 |
109.199 |
| S2 |
N1 |
H5 |
109.199 |
|
H4 |
N1 |
H5 |
108.658 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.494 |
|
|
|
| 2 |
S |
-0.179 |
|
|
|
| 3 |
H |
0.153 |
|
|
|
| 4 |
H |
0.260 |
|
|
|
| 5 |
H |
0.260 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.516 |
1.006 |
0.000 |
1.131 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.815 |
3.447 |
0.000 |
| y |
3.447 |
-18.764 |
0.000 |
| z |
0.000 |
0.000 |
-20.136 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.634 |
3.447 |
0.000 |
| y |
3.447 |
0.712 |
0.000 |
| z |
0.000 |
0.000 |
-1.346 |
|
| Polar |
|---|
| 3z2-r2 | -2.693 |
|---|
| x2-y2 | -0.052 |
|---|
| xy | 3.447 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.427 |
0.470 |
0.000 |
| y |
0.470 |
4.510 |
0.000 |
| z |
0.000 |
0.000 |
3.193 |
<r2> (average value of r
2) Å
2
| <r2> |
35.201 |
| (<r2>)1/2 |
5.933 |
Jump to
S1C1
Energy calculated at B3LYPultrafine/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -454.734431 |
| Energy at 298.15K | -454.738238 |
| HF Energy | -454.734431 |
| Nuclear repulsion energy | 57.875002 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYPultrafine/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3517 |
3394 |
2.22 |
|
|
|
| 2 |
A' |
2535 |
2446 |
50.93 |
|
|
|
| 3 |
A' |
1615 |
1559 |
11.87 |
|
|
|
| 4 |
A' |
1023 |
987 |
28.74 |
|
|
|
| 5 |
A' |
882 |
852 |
20.01 |
|
|
|
| 6 |
A' |
608 |
587 |
127.36 |
|
|
|
| 7 |
A" |
3619 |
3493 |
16.26 |
|
|
|
| 8 |
A" |
1115 |
1076 |
1.77 |
|
|
|
| 9 |
A" |
548 |
528 |
2.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7731.3 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 7460.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYPultrafine/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.084 |
1.099 |
0.000 |
| S2 |
0.084 |
-0.609 |
0.000 |
| H3 |
-1.240 |
-0.926 |
0.000 |
| H4 |
-0.344 |
1.487 |
0.831 |
| H5 |
-0.344 |
1.487 |
-0.831 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7072 | 2.4188 | 1.0124 | 1.0124 |
S2 | 1.7072 | | 1.3611 | 2.2950 | 2.2950 | H3 | 2.4188 | 1.3611 | | 2.7053 | 2.7053 | H4 | 1.0124 | 2.2950 | 2.7053 | | 1.6627 | H5 | 1.0124 | 2.2950 | 2.7053 | 1.6627 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
103.485 |
|
S2 |
N1 |
H4 |
112.586 |
| S2 |
N1 |
H5 |
112.586 |
|
H4 |
N1 |
H5 |
110.407 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.480 |
|
|
|
| 2 |
S |
-0.159 |
|
|
|
| 3 |
H |
0.123 |
|
|
|
| 4 |
H |
0.258 |
|
|
|
| 5 |
H |
0.258 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.994 |
1.149 |
0.000 |
2.301 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.697 |
-1.004 |
0.000 |
| y |
-1.004 |
-18.010 |
0.000 |
| z |
0.000 |
0.000 |
-20.049 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.667 |
-1.004 |
0.000 |
| y |
-1.004 |
1.863 |
0.000 |
| z |
0.000 |
0.000 |
-1.195 |
|
| Polar |
|---|
| 3z2-r2 | -2.391 |
|---|
| x2-y2 | -1.687 |
|---|
| xy | -1.004 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.366 |
0.198 |
0.000 |
| y |
0.198 |
4.494 |
0.000 |
| z |
0.000 |
0.000 |
3.183 |
<r2> (average value of r
2) Å
2
| <r2> |
34.998 |
| (<r2>)1/2 |
5.916 |
