Jump to
S1C2
S1C3
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
Geometric Data calculated at CCD/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
Geometric Data calculated at CCD/6-31G(2df,p)
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -209.922293 |
| Energy at 298.15K | -209.931498 |
| HF Energy | -209.130548 |
| Nuclear repulsion energy | 134.844508 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G(2df,p)
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3855 |
3651 |
59.49 |
|
|
|
| 2 |
A |
3659 |
3465 |
3.76 |
|
|
|
| 3 |
A |
3569 |
3381 |
1.16 |
|
|
|
| 4 |
A |
3152 |
2986 |
32.61 |
|
|
|
| 5 |
A |
3142 |
2976 |
26.10 |
|
|
|
| 6 |
A |
3062 |
2901 |
51.89 |
|
|
|
| 7 |
A |
3033 |
2873 |
53.51 |
|
|
|
| 8 |
A |
1694 |
1605 |
30.01 |
|
|
|
| 9 |
A |
1554 |
1472 |
0.21 |
|
|
|
| 10 |
A |
1536 |
1455 |
5.30 |
|
|
|
| 11 |
A |
1490 |
1412 |
74.06 |
|
|
|
| 12 |
A |
1445 |
1369 |
15.95 |
|
|
|
| 13 |
A |
1415 |
1340 |
3.66 |
|
|
|
| 14 |
A |
1360 |
1288 |
2.06 |
|
|
|
| 15 |
A |
1288 |
1220 |
25.84 |
|
|
|
| 16 |
A |
1221 |
1157 |
4.80 |
|
|
|
| 17 |
A |
1156 |
1095 |
58.44 |
|
|
|
| 18 |
A |
1102 |
1044 |
26.19 |
|
|
|
| 19 |
A |
1037 |
983 |
16.70 |
|
|
|
| 20 |
A |
973 |
921 |
58.95 |
|
|
|
| 21 |
A |
906 |
858 |
10.78 |
|
|
|
| 22 |
A |
858 |
813 |
70.13 |
|
|
|
| 23 |
A |
552 |
523 |
84.33 |
|
|
|
| 24 |
A |
528 |
500 |
54.23 |
|
|
|
| 25 |
A |
332 |
315 |
0.63 |
|
|
|
| 26 |
A |
273 |
259 |
13.62 |
|
|
|
| 27 |
A |
197 |
187 |
5.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22194.0 cm
-1
Scaled (by 0.9472) Zero Point Vibrational Energy (zpe) 21022.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.358 |
-0.567 |
0.119 |
| C2 |
-0.634 |
0.648 |
-0.278 |
| C3 |
0.783 |
0.551 |
0.276 |
| O4 |
1.411 |
-0.626 |
-0.173 |
| H5 |
-1.636 |
-0.495 |
1.091 |
| H6 |
-2.213 |
-0.657 |
-0.414 |
| H7 |
-1.108 |
1.581 |
0.052 |
| H8 |
-0.579 |
0.665 |
-1.369 |
| H9 |
1.383 |
1.399 |
-0.061 |
| H10 |
0.745 |
0.588 |
1.376 |
| H11 |
0.729 |
-1.301 |
-0.112 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4684 | 2.4195 | 2.7845 | 1.0135 | 1.0116 | 2.1634 | 2.0823 | 3.3772 | 2.7080 | 2.2237 |
C2 | 1.4684 | | 1.5241 | 2.4110 | 2.0456 | 2.0531 | 1.0978 | 1.0926 | 2.1629 | 2.1539 | 2.3833 | C3 | 2.4195 | 1.5241 | | 1.4070 | 2.7582 | 3.3028 | 2.1646 | 2.1389 | 1.0924 | 1.1008 | 1.8926 | O4 | 2.7845 | 2.4110 | 1.4070 | | 3.3012 | 3.6318 | 3.3561 | 2.6563 | 2.0279 | 2.0769 | 0.9619 | H5 | 1.0135 | 2.0456 | 2.7582 | 3.3012 | | 1.6195 | 2.3811 | 2.9178 | 3.7452 | 2.6312 | 2.7727 | H6 | 1.0116 | 2.0531 | 3.3028 | 3.6318 | 1.6195 | | 2.5397 | 2.3084 | 4.1571 | 3.6744 | 3.0260 | H7 | 2.1634 | 1.0978 | 2.1646 | 3.3561 | 2.3811 | 2.5397 | | 1.7720 | 2.4996 | 2.4841 | 3.4212 | H8 | 2.0823 | 1.0926 | 2.1389 | 2.6563 | 2.9178 | 2.3084 | 1.7720 | | 2.4700 | 3.0485 | 2.6743 | H9 | 3.3772 | 2.1629 | 1.0924 | 2.0279 | 3.7452 | 4.1571 | 2.4996 | 2.4700 | | 1.7685 | 2.7784 | H10 | 2.7080 | 2.1539 | 1.1008 | 2.0769 | 2.6312 | 3.6744 | 2.4841 | 3.0485 | 1.7685 | | 2.4044 | H11 | 2.2237 | 2.3833 | 1.8926 | 0.9619 | 2.7727 | 3.0260 | 3.4212 | 2.6743 | 2.7784 | 2.4044 | |
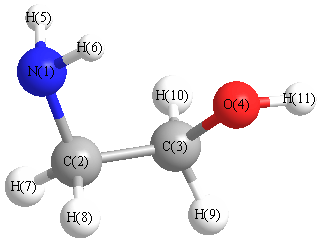 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
107.888 |
|
N1 |
C2 |
H7 |
114.152 |
| N1 |
C2 |
H8 |
107.896 |
|
C2 |
N1 |
H5 |
109.658 |
| C2 |
N1 |
H6 |
110.400 |
|
C2 |
C3 |
O4 |
110.617 |
| C2 |
C3 |
H9 |
110.423 |
|
C2 |
C3 |
H10 |
109.226 |
| C3 |
C2 |
H7 |
110.238 |
|
C3 |
C2 |
H8 |
108.524 |
| C3 |
O4 |
H11 |
104.479 |
|
O4 |
C3 |
H9 |
107.792 |
| O4 |
C3 |
H10 |
111.239 |
|
H5 |
N1 |
H6 |
106.206 |
| H7 |
C2 |
H8 |
107.996 |
|
H9 |
C3 |
H10 |
107.489 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
