return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Furan, 2,3-dihydro-5-methyl-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-269.867349
Energy at 298.15K-269.876606
HF Energy-268.863204
Nuclear repulsion energy238.735264
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3308 3125 1.76      
2 A 3222 3043 6.52      
3 A 3192 3015 23.42      
4 A 3189 3012 6.25      
5 A 3161 2986 15.67      
6 A 3107 2935 12.20      
7 A 3099 2927 40.53      
8 A 3093 2921 30.83      
9 A 1734 1638 35.76      
10 A 1540 1454 0.36      
11 A 1518 1433 0.59      
12 A 1515 1431 5.04      
13 A 1498 1415 6.72      
14 A 1434 1355 22.74      
15 A 1416 1337 3.20      
16 A 1338 1263 11.19      
17 A 1285 1214 39.01      
18 A 1275 1204 14.99      
19 A 1225 1157 4.71      
20 A 1217 1149 30.21      
21 A 1094 1033 0.35      
22 A 1068 1009 7.86      
23 A 1058 999 41.36      
24 A 1040 982 4.94      
25 A 987 932 12.87      
26 A 962 909 9.59      
27 A 929 877 21.05      
28 A 875 826 6.16      
29 A 740 699 24.76      
30 A 672 635 0.22      
31 A 636 601 1.36      
32 A 556 525 1.69      
33 A 329 311 1.97      
34 A 229 216 4.62      
35 A 179 169 3.58      
36 A 160 151 1.04      

Unscaled Zero Point Vibrational Energy (zpe) 26939.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 25444.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.25295 0.10999 0.08068

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.340 -0.811 0.130
H2 -1.933 -1.451 -0.523
H3 -1.545 -1.068 1.174
O4 0.052 -1.089 -0.126
C5 -1.536 0.696 -0.101
H6 -2.237 1.119 0.620
H7 -1.917 0.916 -1.104
C8 -0.115 1.173 0.069
H9 0.206 2.200 0.130
C10 2.181 0.009 0.013
H11 2.531 -0.471 -0.903
H12 2.634 0.997 0.090
H13 2.514 -0.604 0.852
C14 0.699 0.113 0.001

Atom - Atom Distances (Å)
  C1 H2 H3 O4 C5 H6 H7 C8 H9 C10 H11 H12 H13 C14
C11.08921.09471.44221.53692.18472.19982.33213.38483.61674.02074.36553.92642.2425
H21.08921.78152.05622.22322.82932.43773.24594.28134.39774.58595.21704.73083.1061
H31.09471.78152.05922.17632.36043.04382.87903.85224.04874.61374.78564.09852.7945
O41.44222.05622.05922.38933.26732.97602.27653.30282.39942.67063.32592.69301.3711
C51.53692.22322.17632.38931.09131.09521.50832.31343.78144.30654.18494.35912.3125
H62.18472.82932.36043.26731.09131.76512.19312.71624.59555.25224.90115.05963.1655
H72.19982.43773.04382.97601.09521.76512.16462.77064.34284.66364.70525.07662.9515
C82.33213.24592.87902.27651.50832.19312.16461.07812.57463.26332.75473.26891.3386
H93.38484.28133.85223.30282.31342.71622.77061.07812.95173.68882.70983.70312.1486
C103.61674.39774.04872.39943.78144.59554.34282.57462.95171.09201.08901.09171.4849
H114.02074.58594.61372.67064.30655.25224.66363.26333.68881.09201.77501.76022.1243
H124.36555.21704.78563.32594.18494.90114.70522.75472.70981.08901.77501.77702.1284
H133.92644.73084.09852.69304.35915.05965.07663.26893.70311.09171.76021.77702.1288
C142.24253.10612.79451.37112.31253.16552.95151.33862.14861.48492.12432.12842.1288

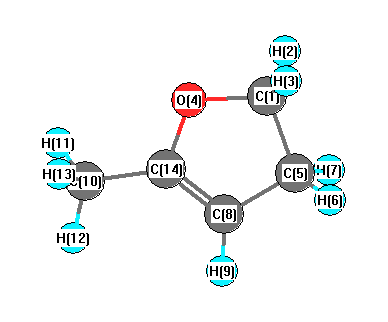
picture of Furan, 2,3-dihydro-5-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O4 C14 105.679 C1 C5 H6 111.323
C1 C5 H7 112.302 C1 C5 C8 99.959
H2 C1 H3 109.324 H2 C1 O4 107.824
H2 C1 C5 114.610 H3 C1 O4 107.740
H3 C1 C5 110.459 O4 C1 C5 106.605
O4 C14 C8 114.298 O4 C14 C10 114.241
C5 C8 H9 126.056 C5 C8 C14 108.493
H6 C5 H7 107.657 H6 C5 C8 114.085
H7 C5 C8 111.505 C8 C14 C10 131.449
H9 C8 C14 125.165 H11 C10 H12 108.949
H11 C10 H13 107.429 H11 C10 C14 110.116
H12 C10 H13 109.157 H12 C10 C14 110.626
H13 C10 C14 110.491
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability