Jump to
S1C2
S1C3
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -691.893067 |
| Energy at 298.15K | -691.892446 |
| HF Energy | -691.449752 |
| Nuclear repulsion energy | 61.068687 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.306 |
| C2 |
0.000 |
0.000 |
-1.279 |
| N3 |
0.000 |
0.000 |
-2.449 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.5853 | 3.7555 |
C2 | 2.5853 | | 1.1701 | N3 | 3.7555 | 1.1701 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -691.895658 |
| Energy at 298.15K | |
| HF Energy | -691.457397 |
| Nuclear repulsion energy | 64.728752 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.204 |
| C2 |
0.000 |
0.000 |
-2.394 |
| N3 |
0.000 |
0.000 |
-1.216 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.5973 | 2.4195 |
C2 | 3.5973 | | 1.1778 | N3 | 2.4195 | 1.1778 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -691.902800 |
| Energy at 298.15K | -691.902712 |
| HF Energy | -691.460633 |
| Nuclear repulsion energy | 69.062762 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCD/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.031 |
0.000 |
| C2 |
0.631 |
-1.574 |
0.000 |
| N3 |
-0.541 |
-1.451 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.6805 | 2.5406 |
C2 | 2.6805 | | 1.1787 | N3 | 2.5406 | 1.1787 | |
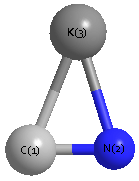 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
70.399 |
|
K1 |
N3 |
C2 |
83.683 |
| C2 |
K1 |
N3 |
25.917 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
