Jump to
S1C2
S1C3
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -692.711312 |
| Energy at 298.15K | -692.710431 |
| HF Energy | -692.711312 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.305 |
| C2 |
0.000 |
0.000 |
-1.280 |
| N3 |
0.000 |
0.000 |
-2.445 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.5846 | 3.7505 |
C2 | 2.5846 | | 1.1659 | N3 | 3.7505 | 1.1659 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.668 |
|
|
|
| 2 |
C |
-0.444 |
|
|
|
| 3 |
N |
-0.225 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
12.082 |
12.082 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.795 |
0.000 |
0.000 |
| y |
0.000 |
-22.795 |
0.000 |
| z |
0.000 |
0.000 |
-29.914 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.559 |
0.000 |
0.000 |
| y |
0.000 |
3.559 |
0.000 |
| z |
0.000 |
0.000 |
-7.118 |
|
| Polar |
|---|
| 3z2-r2 | -14.236 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.314 |
-0.000 |
-0.001 |
| y |
-0.000 |
3.314 |
0.000 |
| z |
-0.001 |
0.000 |
5.837 |
<r2> (average value of r
2) Å
2
| <r2> |
99.763 |
| (<r2>)1/2 |
9.988 |
Jump to
S1C1
S1C3
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -692.715083 |
| Energy at 298.15K | |
| HF Energy | -692.715083 |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.205 |
| C2 |
0.000 |
0.000 |
-2.394 |
| N3 |
0.000 |
0.000 |
-1.218 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.5985 | 2.4229 |
C2 | 3.5985 | | 1.1756 | N3 | 2.4229 | 1.1756 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.702 |
|
|
|
| 2 |
C |
-0.357 |
|
|
|
| 3 |
N |
-0.345 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
11.856 |
11.856 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.784 |
0.000 |
0.000 |
| y |
0.000 |
-22.784 |
0.000 |
| z |
0.000 |
0.000 |
-32.023 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.620 |
0.000 |
0.000 |
| y |
0.000 |
4.620 |
0.000 |
| z |
0.000 |
0.000 |
-9.239 |
|
| Polar |
|---|
| 3z2-r2 | -18.479 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.265 |
-0.001 |
-0.000 |
| y |
-0.001 |
3.264 |
0.000 |
| z |
-0.000 |
0.000 |
5.720 |
<r2> (average value of r
2) Å
2
| <r2> |
88.497 |
| (<r2>)1/2 |
9.407 |
Jump to
S1C1
S1C2
Energy calculated at wB97X-D/6-31G(2df,p)
| | hartrees |
|---|
| Energy at 0K | -692.721074 |
| Energy at 298.15K | -692.721019 |
| HF Energy | -692.721074 |
| Nuclear repulsion energy | 68.800977 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at wB97X-D/6-31G(2df,p)
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.039 |
0.000 |
| C2 |
0.627 |
-1.604 |
0.000 |
| N3 |
-0.538 |
-1.445 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7159 | 2.5419 |
C2 | 2.7159 | | 1.1759 | N3 | 2.5419 | 1.1759 | |
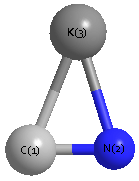 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
68.918 |
|
K1 |
N3 |
C2 |
85.511 |
| C2 |
K1 |
N3 |
25.572 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G(2df,p)
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
K |
0.628 |
|
|
|
| 2 |
C |
-0.319 |
|
|
|
| 3 |
N |
-0.309 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.435 |
9.902 |
0.000 |
9.911 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.070 |
0.877 |
0.000 |
| y |
0.877 |
-25.485 |
0.000 |
| z |
0.000 |
0.000 |
-23.076 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.790 |
0.877 |
0.000 |
| y |
0.877 |
-0.412 |
0.000 |
| z |
0.000 |
0.000 |
3.202 |
|
| Polar |
|---|
| 3z2-r2 | 6.403 |
|---|
| x2-y2 | -1.585 |
|---|
| xy | 0.877 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.373 |
-0.142 |
0.000 |
| y |
-0.142 |
4.306 |
0.000 |
| z |
0.000 |
0.000 |
3.420 |
<r2> (average value of r
2) Å
2
| <r2> |
70.693 |
| (<r2>)1/2 |
8.408 |
