return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Pentadiene, (Z)-)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-194.759663
Energy at 298.15K-194.767103
HF Energy-193.979378
Nuclear repulsion energy155.763588
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3310 3126 6.90      
2 A' 3239 3059 14.83      
3 A' 3223 3044 15.33      
4 A' 3214 3035 2.80      
5 A' 3208 3030 2.69      
6 A' 3196 3019 0.19      
7 A' 3099 2927 14.88      
8 A' 1723 1627 4.02      
9 A' 1656 1564 0.91      
10 A' 1519 1435 6.73      
11 A' 1483 1401 10.44      
12 A' 1434 1354 3.70      
13 A' 1401 1324 0.84      
14 A' 1328 1255 0.84      
15 A' 1298 1226 0.91      
16 A' 1197 1131 1.28      
17 A' 1069 1009 0.02      
18 A' 979 924 9.57      
19 A' 905 855 1.41      
20 A' 612 578 4.68      
21 A' 385 364 3.27      
22 A' 224 211 0.27      
23 A" 3169 2994 10.16      
24 A" 1510 1426 5.52      
25 A" 1067 1007 1.82      
26 A" 1051 993 11.17      
27 A" 999 944 1.93      
28 A" 937 885 33.72      
29 A" 814 769 6.35      
30 A" 650 614 21.33      
31 A" 358 338 0.09      
32 A" 131 124 0.01      
33 A" 116 109 0.54      

Unscaled Zero Point Vibrational Energy (zpe) 25252.1 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 23850.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.53175 0.08840 0.07687

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.388 0.160 0.000
C2 1.099 -0.213 0.000
C3 0.000 0.738 0.000
C4 -1.314 0.449 0.000
C5 -1.943 -0.910 0.000
H6 2.667 1.208 0.000
H7 3.190 -0.565 0.000
H8 0.863 -1.272 0.000
H9 0.290 1.786 0.000
H10 -2.008 1.285 0.000
H11 -1.212 -1.716 0.000
H12 -2.583 -1.036 0.877
H13 -2.583 -1.036 -0.877

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.34182.45663.71284.46091.08391.08182.09202.65414.53754.05885.18755.1875
C21.34181.45352.50213.12072.11612.12051.08462.15683.44952.75603.87353.8735
C32.45661.45351.34532.54772.70803.44602.18741.08712.08142.73673.25403.2540
C43.71282.50211.34531.49744.05254.61682.77492.08771.08652.16742.14122.1412
C54.46093.12072.54771.49745.07305.14462.82913.50022.19551.08851.09321.0932
H61.08392.11612.70804.05255.07301.84843.06642.44674.67594.85695.77645.7764
H71.08182.12053.44604.61685.14461.84842.43223.73365.51774.54955.85855.8585
H82.09201.08462.18742.77492.82913.06642.43223.11103.84452.12133.56393.5639
H92.65412.15681.08712.08773.50022.44673.73363.11102.35193.80994.12114.1211
H104.53753.44952.08141.08652.19554.67595.51773.84452.35193.10472.54642.5464
H114.05882.75602.73672.16741.08854.85694.54952.12133.80993.10471.76461.7646
H125.18753.87353.25402.14121.09325.77645.85853.56394.12112.54641.76461.7541
H135.18753.87353.25402.14121.09325.77645.85853.56394.12112.54641.76461.7541

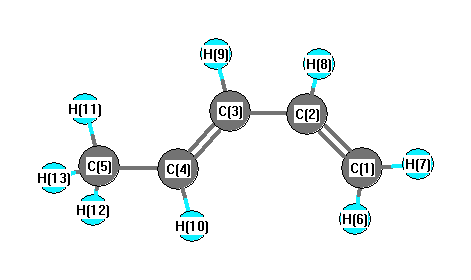
picture of 1,3-Pentadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 122.948 C1 C2 H8 118.741
C2 C1 H6 121.103 C2 C1 H7 121.714
C2 C3 C4 126.712 C2 C3 H9 115.432
C3 C2 H8 118.312 C3 C4 C5 127.246
C3 C4 H10 117.318 C4 C3 H9 117.856
C4 C5 H11 112.935 C4 C5 H12 110.519
C4 C5 H13 110.519 C5 C4 H10 115.437
H6 C1 H7 117.184 H11 C5 H12 107.962
H11 C5 H13 107.962 H12 C5 H13 106.699
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability