return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (cis-2-butenal)

using model chemistry: MP2/6-31G(2df,p)

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at MP2/6-31G(2df,p)
 hartrees
Energy at 0K-230.657426
Energy at 298.15K-230.663264
Nuclear repulsion energy155.428215
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/6-31G(2df,p)
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3234 3054 7.97      
2 A' 3218 3039 3.51      
3 A' 3201 3024 5.12      
4 A' 3097 2925 8.17      
5 A' 2983 2817 134.97      
6 A' 1770 1672 58.10      
7 A' 1709 1614 113.14      
8 A' 1513 1429 19.48      
9 A' 1450 1370 7.73      
10 A' 1427 1347 0.97      
11 A' 1330 1256 1.71      
12 A' 1320 1246 20.01      
13 A' 1155 1091 5.39      
14 A' 1039 981 11.32      
15 A' 903 853 30.36      
16 A' 742 701 34.95      
17 A' 395 373 3.40      
18 A' 205 193 4.69      
19 A" 3174 2998 5.82      
20 A" 1507 1423 6.79      
21 A" 1086 1026 2.46      
22 A" 1046 988 1.63      
23 A" 1029 972 24.11      
24 A" 779 736 0.01      
25 A" 246 233 3.61      
26 A" 208 196 0.49      
27 A" 147 139 5.20      

Unscaled Zero Point Vibrational Energy (zpe) 19954.7 cm-1
Scaled (by 0.9445) Zero Point Vibrational Energy (zpe) 18847.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2/6-31G(2df,p)
ABC
0.63276 0.08782 0.07822

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2/6-31G(2df,p)

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.470 0.555 0.000
C2 0.000 0.703 0.000
C3 0.799 -0.373 0.000
C4 2.290 -0.340 0.000
O5 -2.054 -0.513 0.000
H6 -2.040 1.504 0.000
H7 0.406 1.710 0.000
H8 0.311 -1.345 0.000
H9 2.672 0.682 0.000
H10 2.686 -0.860 0.876
H11 2.686 -0.860 -0.876

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.47772.45183.86551.21781.10722.20302.60504.14424.47684.4768
C21.47771.34062.51652.38762.19191.08522.07212.67213.22863.2286
C32.45181.34061.49162.85683.40392.11961.08772.14942.13632.1363
C43.86552.51651.49164.34804.70682.78402.21981.09021.09311.0931
O51.21782.38762.85684.34802.01783.31592.50764.87504.83274.8327
H61.10722.19193.40394.70682.01782.45483.69474.78365.35645.3564
H72.20301.08522.11962.78403.31592.45483.05652.48853.54513.5451
H82.60502.07211.08772.21982.50763.69473.05653.11132.57702.5770
H94.14422.67212.14941.09024.87504.78362.48853.11131.77311.7731
H104.47683.22862.13631.09314.83275.35643.54512.57701.77311.7525
H114.47683.22862.13631.09314.83275.35643.54512.57701.77311.7525

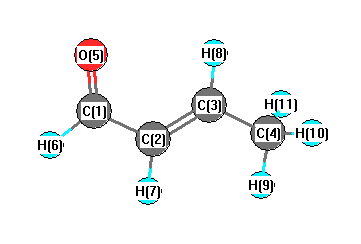
picture of cis-2-butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.832 C1 C2 H7 117.720
C2 C1 O5 124.414 C2 C1 H6 115.229
C2 C3 C4 125.305 C2 C3 H8 116.772
C3 C2 H7 121.447 C3 C4 H9 111.781
C3 C4 H10 110.547 C3 C4 H11 110.547
C4 C3 H8 117.924 O5 C1 H6 120.357
H9 C4 H10 108.608 H9 C4 H11 108.608
H10 C4 H11 106.580
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2/6-31G(2df,p) Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.263      
2 C -0.279      
3 C -0.036      
4 C -0.445      
5 O -0.423      
6 H 0.121      
7 H 0.145      
8 H 0.188      
9 H 0.151      
10 H 0.158      
11 H 0.158      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000